Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -3110.572712 |
| Energy at 298.15K | |
| HF Energy | -3109.964432 |
| Nuclear repulsion energy | 288.484929 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
202 |
202 |
6.23 |
|
|
|
| 2 |
A' |
257 |
257 |
0.67 |
|
|
|
| 3 |
A' |
676 |
676 |
43.10 |
|
|
|
| 4 |
A' |
790 |
790 |
24.50 |
|
|
|
| 5 |
A' |
1093 |
1093 |
1.08 |
|
|
|
| 6 |
A' |
1277 |
1277 |
55.09 |
|
|
|
| 7 |
A' |
1383 |
1383 |
3.44 |
|
|
|
| 8 |
A' |
1507 |
1507 |
4.18 |
|
|
|
| 9 |
A' |
1510 |
1510 |
0.00 |
|
|
|
| 10 |
A' |
3130 |
3130 |
2.11 |
|
|
|
| 11 |
A' |
3138 |
3138 |
13.46 |
|
|
|
| 12 |
A" |
121 |
121 |
5.03 |
|
|
|
| 13 |
A" |
784 |
784 |
2.83 |
|
|
|
| 14 |
A" |
1014 |
1014 |
0.25 |
|
|
|
| 15 |
A" |
1176 |
1176 |
3.93 |
|
|
|
| 16 |
A" |
1327 |
1327 |
0.21 |
|
|
|
| 17 |
A" |
3187 |
3187 |
0.55 |
|
|
|
| 18 |
A" |
3209 |
3209 |
4.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12890.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12890.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.639 |
0.000 |
| C2 |
1.243 |
-0.233 |
0.000 |
| Br3 |
-1.593 |
-0.489 |
0.000 |
| Cl4 |
2.696 |
0.815 |
0.000 |
| H5 |
-0.047 |
1.264 |
0.891 |
| H6 |
-0.047 |
1.264 |
-0.891 |
| H7 |
1.284 |
-0.860 |
0.891 |
| H8 |
1.284 |
-0.860 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5182 | 1.9520 | 2.7020 | 1.0898 | 1.0898 | 2.1654 | 2.1654 |
C2 | 1.5182 | | 2.8474 | 1.7923 | 2.1678 | 2.1678 | 1.0901 | 1.0901 | Br3 | 1.9520 | 2.8474 | | 4.4833 | 2.5018 | 2.5018 | 3.0348 | 3.0348 | Cl4 | 2.7020 | 1.7923 | 4.4833 | | 2.9190 | 2.9190 | 2.3652 | 2.3652 | H5 | 1.0898 | 2.1678 | 2.5018 | 2.9190 | | 1.7830 | 2.5063 | 3.0756 | H6 | 1.0898 | 2.1678 | 2.5018 | 2.9190 | 1.7830 | | 3.0756 | 2.5063 | H7 | 2.1654 | 1.0901 | 3.0348 | 2.3652 | 2.5063 | 3.0756 | | 1.7822 | H8 | 2.1654 | 1.0901 | 3.0348 | 2.3652 | 3.0756 | 2.5063 | 1.7822 | |
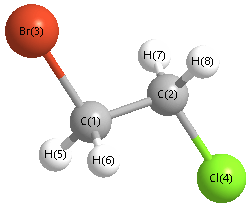 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.133 |
|
C1 |
C2 |
H7 |
111.178 |
| C1 |
C2 |
H8 |
111.178 |
|
C2 |
C1 |
Br3 |
109.642 |
| C2 |
C1 |
Cl4 |
38.805 |
|
C2 |
C1 |
H6 |
111.385 |
| Br3 |
C1 |
H5 |
107.241 |
|
Br3 |
C1 |
H6 |
107.241 |
| Cl4 |
C2 |
H7 |
107.784 |
|
Cl4 |
C2 |
H8 |
107.784 |
| H5 |
C1 |
H6 |
109.774 |
|
H7 |
C2 |
H8 |
109.661 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -3110.569802 |
| Energy at 298.15K | |
| HF Energy | -3109.960979 |
| Nuclear repulsion energy | 304.510462 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3188 |
3188 |
4.73 |
|
|
|
| 2 |
A |
3173 |
3173 |
2.67 |
|
|
|
| 3 |
A |
3121 |
3121 |
18.88 |
|
|
|
| 4 |
A |
3108 |
3108 |
9.20 |
|
|
|
| 5 |
A |
1496 |
1496 |
0.48 |
|
|
|
| 6 |
A |
1487 |
1487 |
7.59 |
|
|
|
| 7 |
A |
1389 |
1389 |
28.09 |
|
|
|
| 8 |
A |
1343 |
1343 |
48.56 |
|
|
|
| 9 |
A |
1249 |
1249 |
4.39 |
|
|
|
| 10 |
A |
1181 |
1181 |
2.95 |
|
|
|
| 11 |
A |
1076 |
1076 |
1.05 |
|
|
|
| 12 |
A |
969 |
969 |
7.27 |
|
|
|
| 13 |
A |
900 |
900 |
17.37 |
|
|
|
| 14 |
A |
712 |
712 |
17.22 |
|
|
|
| 15 |
A |
598 |
598 |
9.46 |
|
|
|
| 16 |
A |
397 |
397 |
5.50 |
|
|
|
| 17 |
A |
251 |
251 |
0.87 |
|
|
|
| 18 |
A |
107 |
107 |
0.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12872.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12872.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.009 |
1.099 |
-0.386 |
| C2 |
1.289 |
0.897 |
0.405 |
| Br3 |
-1.357 |
-0.224 |
0.036 |
| Cl4 |
2.170 |
-0.577 |
-0.087 |
| H5 |
-0.418 |
2.073 |
-0.140 |
| H6 |
0.194 |
1.037 |
-1.458 |
| H7 |
1.953 |
1.745 |
0.221 |
| H8 |
1.084 |
0.821 |
1.474 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5181 | 1.9477 | 2.7502 | 1.0922 | 1.0903 | 2.1366 | 2.1654 |
C2 | 1.5181 | | 2.8972 | 1.7855 | 2.1440 | 2.1666 | 1.0932 | 1.0907 | Br3 | 1.9477 | 2.8972 | | 3.5464 | 2.4882 | 2.4955 | 3.8560 | 3.0191 | Cl4 | 2.7502 | 1.7855 | 3.5464 | | 3.7042 | 2.8967 | 2.3523 | 2.3598 | H5 | 1.0922 | 2.1440 | 2.4882 | 3.7042 | | 1.7849 | 2.4207 | 2.5354 | H6 | 1.0903 | 2.1666 | 2.4955 | 2.8967 | 1.7849 | | 2.5336 | 3.0720 | H7 | 2.1366 | 1.0932 | 3.8560 | 2.3523 | 2.4207 | 2.5336 | | 1.7831 | H8 | 2.1654 | 1.0907 | 3.0191 | 2.3598 | 2.5354 | 3.0720 | 1.7831 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.458 |
|
C1 |
C2 |
H7 |
108.721 |
| C1 |
C2 |
H8 |
111.145 |
|
C2 |
C1 |
Br3 |
112.840 |
| C2 |
C1 |
Cl4 |
36.869 |
|
C2 |
C1 |
H6 |
111.264 |
| Br3 |
C1 |
H5 |
106.447 |
|
Br3 |
C1 |
H6 |
107.051 |
| Cl4 |
C2 |
H7 |
107.138 |
|
Cl4 |
C2 |
H8 |
107.804 |
| H5 |
C1 |
H6 |
109.734 |
|
H7 |
C2 |
H8 |
109.468 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
