Jump to
S1C2
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -208.612422 |
| Energy at 298.15K | -208.618452 |
| HF Energy | -207.942111 |
| Nuclear repulsion energy | 117.258269 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3909 |
3714 |
101.91 |
|
|
|
| 2 |
A |
3192 |
3033 |
10.05 |
|
|
|
| 3 |
A |
3150 |
2993 |
11.37 |
|
|
|
| 4 |
A |
3078 |
2924 |
16.55 |
|
|
|
| 5 |
A |
1703 |
1618 |
0.64 |
|
|
|
| 6 |
A |
1507 |
1432 |
10.17 |
|
|
|
| 7 |
A |
1460 |
1388 |
7.79 |
|
|
|
| 8 |
A |
1412 |
1342 |
25.23 |
|
|
|
| 9 |
A |
1295 |
1231 |
53.91 |
|
|
|
| 10 |
A |
1167 |
1109 |
2.60 |
|
|
|
| 11 |
A |
1037 |
985 |
116.20 |
|
|
|
| 12 |
A |
924 |
878 |
8.01 |
|
|
|
| 13 |
A |
573 |
544 |
14.52 |
|
|
|
| 14 |
A |
326 |
310 |
1.82 |
|
|
|
| 15 |
A |
3154 |
2997 |
14.03 |
|
|
|
| 16 |
A |
1498 |
1423 |
9.05 |
|
|
|
| 17 |
A |
1084 |
1030 |
1.34 |
|
|
|
| 18 |
A |
911 |
866 |
11.06 |
|
|
|
| 19 |
A |
394 |
375 |
137.29 |
|
|
|
| 20 |
A |
285 |
271 |
3.82 |
|
|
|
| 21 |
A |
203 |
193 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16130.1 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 15326.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.310 |
1.290 |
0.000 |
| C2 |
0.000 |
0.570 |
0.000 |
| N3 |
0.010 |
-0.712 |
0.000 |
| O4 |
1.324 |
-1.197 |
0.000 |
| H5 |
1.178 |
-2.146 |
0.000 |
| H6 |
-2.128 |
0.568 |
0.000 |
| H7 |
-1.397 |
1.931 |
0.883 |
| H8 |
-1.397 |
1.931 |
-0.883 |
| H9 |
0.942 |
1.123 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4952 | 2.3989 | 3.6230 | 4.2423 | 1.0913 | 1.0944 | 1.0944 | 2.2581 |
C2 | 1.4952 | | 1.2821 | 2.2079 | 2.9599 | 2.1278 | 2.1411 | 2.1411 | 1.0924 | N3 | 2.3989 | 1.2821 | | 1.4000 | 1.8486 | 2.4922 | 3.1219 | 3.1219 | 2.0583 | O4 | 3.6230 | 2.2079 | 1.4000 | | 0.9596 | 3.8768 | 4.2388 | 4.2388 | 2.3517 | H5 | 4.2423 | 2.9599 | 1.8486 | 0.9596 | | 4.2767 | 4.9017 | 4.9017 | 3.2774 | H6 | 1.0913 | 2.1278 | 2.4922 | 3.8768 | 4.2767 | | 1.7809 | 1.7809 | 3.1195 | H7 | 1.0944 | 2.1411 | 3.1219 | 4.2388 | 4.9017 | 1.7809 | | 1.7667 | 2.6272 | H8 | 1.0944 | 2.1411 | 3.1219 | 4.2388 | 4.9017 | 1.7809 | 1.7667 | | 2.6272 | H9 | 2.2581 | 1.0924 | 2.0583 | 2.3517 | 3.2774 | 3.1195 | 2.6272 | 2.6272 | |
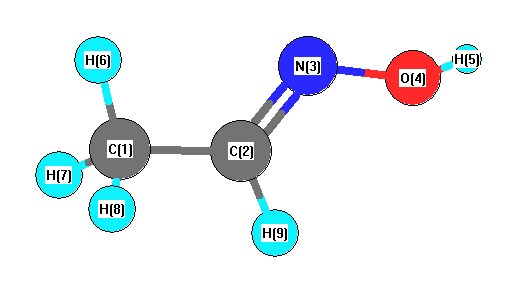 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.285 |
|
C1 |
C2 |
H9 |
120.741 |
| C2 |
C1 |
H6 |
109.717 |
|
C2 |
C1 |
H7 |
110.585 |
| C2 |
C1 |
H8 |
110.585 |
|
C2 |
N3 |
O4 |
110.735 |
| N3 |
C2 |
H9 |
119.974 |
|
N3 |
O4 |
H5 |
101.516 |
| H6 |
C1 |
H7 |
109.134 |
|
H6 |
C1 |
H8 |
109.134 |
| H7 |
C1 |
H8 |
107.639 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -208.611663 |
| Energy at 298.15K | |
| HF Energy | -207.940773 |
| Nuclear repulsion energy | 119.801268 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3912 |
3717 |
103.34 |
|
|
|
| 2 |
A' |
3223 |
3063 |
5.83 |
|
|
|
| 3 |
A' |
3206 |
3047 |
5.61 |
|
|
|
| 4 |
A' |
3085 |
2931 |
12.29 |
|
|
|
| 5 |
A' |
1710 |
1624 |
3.09 |
|
|
|
| 6 |
A' |
1503 |
1428 |
16.25 |
|
|
|
| 7 |
A' |
1422 |
1351 |
19.88 |
|
|
|
| 8 |
A' |
1392 |
1323 |
30.72 |
|
|
|
| 9 |
A' |
1346 |
1279 |
39.22 |
|
|
|
| 10 |
A' |
1163 |
1105 |
7.73 |
|
|
|
| 11 |
A' |
962 |
914 |
115.50 |
|
|
|
| 12 |
A' |
930 |
884 |
0.01 |
|
|
|
| 13 |
A' |
679 |
645 |
12.23 |
|
|
|
| 14 |
A' |
321 |
305 |
1.58 |
|
|
|
| 15 |
A" |
3156 |
2999 |
11.70 |
|
|
|
| 16 |
A" |
1508 |
1433 |
9.85 |
|
|
|
| 17 |
A" |
1070 |
1017 |
1.18 |
|
|
|
| 18 |
A" |
865 |
822 |
13.48 |
|
|
|
| 19 |
A" |
493 |
468 |
37.92 |
|
|
|
| 20 |
A" |
387 |
368 |
91.93 |
|
|
|
| 21 |
A" |
60i |
57i |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16136.1 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 15332.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.448 |
0.482 |
0.000 |
| C2 |
0.000 |
0.866 |
0.000 |
| N3 |
1.022 |
0.085 |
0.000 |
| O4 |
0.650 |
-1.264 |
0.000 |
| H5 |
1.510 |
-1.690 |
0.000 |
| H6 |
-1.566 |
-0.600 |
0.000 |
| H7 |
-1.943 |
0.899 |
0.883 |
| H8 |
-1.943 |
0.899 |
-0.883 |
| H9 |
0.276 |
1.918 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4981 | 2.5018 | 2.7298 | 3.6698 | 1.0883 | 1.0944 | 1.0944 | 2.2439 |
C2 | 1.4981 | | 1.2859 | 2.2268 | 2.9685 | 2.1446 | 2.1343 | 2.1343 | 1.0874 | N3 | 2.5018 | 1.2859 | | 1.3996 | 1.8412 | 2.6767 | 3.1988 | 3.1988 | 1.9781 | O4 | 2.7298 | 2.2268 | 1.3996 | | 0.9594 | 2.3131 | 3.4903 | 3.4903 | 3.2034 | H5 | 3.6698 | 2.9685 | 1.8412 | 0.9594 | | 3.2628 | 4.4050 | 4.4050 | 3.8126 | H6 | 1.0883 | 2.1446 | 2.6767 | 2.3131 | 3.2628 | | 1.7799 | 1.7799 | 3.1193 | H7 | 1.0944 | 2.1343 | 3.1988 | 3.4903 | 4.4050 | 1.7799 | | 1.7651 | 2.5966 | H8 | 1.0944 | 2.1343 | 3.1988 | 3.4903 | 4.4050 | 1.7799 | 1.7651 | | 2.5966 | H9 | 2.2439 | 1.0874 | 1.9781 | 3.2034 | 3.8126 | 3.1193 | 2.5966 | 2.5966 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.786 |
|
C1 |
C2 |
H9 |
119.569 |
| C2 |
C1 |
H6 |
111.034 |
|
C2 |
C1 |
H7 |
109.845 |
| C2 |
C1 |
H8 |
109.845 |
|
C2 |
N3 |
O4 |
111.966 |
| N3 |
C2 |
H9 |
112.644 |
|
N3 |
O4 |
H5 |
100.963 |
| H6 |
C1 |
H7 |
109.270 |
|
H6 |
C1 |
H8 |
109.270 |
| H7 |
C1 |
H8 |
107.503 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
