Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/6-311G**
| | hartrees |
|---|
| Energy at 0K | -209.013269 |
| Energy at 298.15K | |
| HF Energy | -208.769216 |
| Nuclear repulsion energy | 119.820241 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3879 |
3879 |
98.30 |
|
|
|
| 2 |
A' |
3191 |
3191 |
8.52 |
|
|
|
| 3 |
A' |
3174 |
3174 |
5.17 |
|
|
|
| 4 |
A' |
3060 |
3060 |
13.79 |
|
|
|
| 5 |
A' |
1726 |
1726 |
6.83 |
|
|
|
| 6 |
A' |
1495 |
1495 |
17.72 |
|
|
|
| 7 |
A' |
1419 |
1419 |
17.07 |
|
|
|
| 8 |
A' |
1386 |
1386 |
29.61 |
|
|
|
| 9 |
A' |
1349 |
1349 |
44.52 |
|
|
|
| 10 |
A' |
1155 |
1155 |
9.54 |
|
|
|
| 11 |
A' |
940 |
940 |
121.46 |
|
|
|
| 12 |
A' |
919 |
919 |
14.20 |
|
|
|
| 13 |
A' |
679 |
679 |
13.04 |
|
|
|
| 14 |
A' |
317 |
317 |
1.43 |
|
|
|
| 15 |
A" |
3113 |
3113 |
15.10 |
|
|
|
| 16 |
A" |
1504 |
1504 |
10.30 |
|
|
|
| 17 |
A" |
1074 |
1074 |
0.78 |
|
|
|
| 18 |
A" |
869 |
869 |
13.38 |
|
|
|
| 19 |
A" |
503 |
503 |
39.13 |
|
|
|
| 20 |
A" |
389 |
389 |
91.03 |
|
|
|
| 21 |
A" |
18i |
18i |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16061.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16061.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.448 |
0.483 |
0.000 |
| C2 |
0.000 |
0.864 |
0.000 |
| N3 |
1.017 |
0.090 |
0.000 |
| O4 |
0.655 |
-1.267 |
0.000 |
| H5 |
1.513 |
-1.696 |
0.000 |
| H6 |
-1.569 |
-0.597 |
0.000 |
| H7 |
-1.944 |
0.901 |
0.880 |
| H8 |
-1.944 |
0.901 |
-0.880 |
| H9 |
0.270 |
1.916 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4969 | 2.4961 | 2.7355 | 3.6763 | 1.0865 | 1.0933 | 1.0933 | 2.2367 |
C2 | 1.4969 | | 1.2780 | 2.2293 | 2.9738 | 2.1436 | 2.1341 | 2.1341 | 1.0862 | N3 | 2.4961 | 1.2780 | | 1.4053 | 1.8541 | 2.6761 | 3.1937 | 3.1937 | 1.9727 | O4 | 2.7355 | 2.2293 | 1.4053 | | 0.9597 | 2.3226 | 3.4967 | 3.4967 | 3.2064 | H5 | 3.6763 | 2.9738 | 1.8541 | 0.9597 | | 3.2724 | 4.4124 | 4.4124 | 3.8202 | H6 | 1.0865 | 2.1436 | 2.6761 | 2.3226 | 3.2724 | | 1.7769 | 1.7769 | 3.1137 | H7 | 1.0933 | 2.1341 | 3.1937 | 3.4967 | 4.4124 | 1.7769 | | 1.7601 | 2.5895 | H8 | 1.0933 | 2.1341 | 3.1937 | 3.4967 | 4.4124 | 1.7769 | 1.7601 | | 2.5895 | H9 | 2.2367 | 1.0862 | 1.9727 | 3.2064 | 3.8202 | 3.1137 | 2.5895 | 2.5895 | |
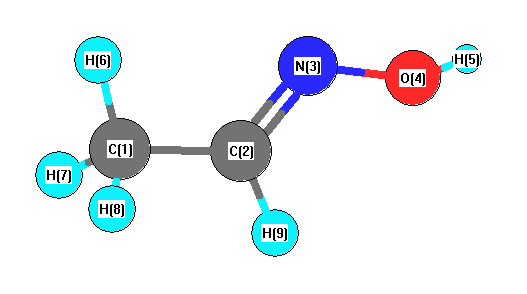 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.022 |
|
C1 |
C2 |
H9 |
119.121 |
| C2 |
C1 |
H6 |
111.150 |
|
C2 |
C1 |
H7 |
109.980 |
| C2 |
C1 |
H8 |
109.980 |
|
C2 |
N3 |
O4 |
112.282 |
| N3 |
C2 |
H9 |
112.857 |
|
N3 |
O4 |
H5 |
101.594 |
| H6 |
C1 |
H7 |
109.211 |
|
H6 |
C1 |
H8 |
109.211 |
| H7 |
C1 |
H8 |
107.218 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
