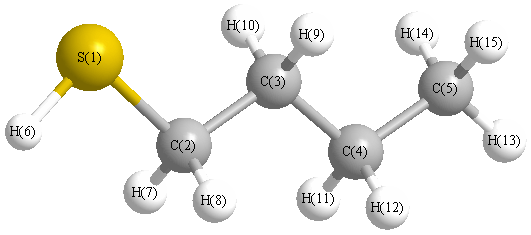
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3127 |
3000 |
34.11 |
|
|
|
| 2 |
A' |
3071 |
2946 |
35.86 |
|
|
|
| 3 |
A' |
3046 |
2923 |
42.02 |
|
|
|
| 4 |
A' |
3043 |
2919 |
16.68 |
|
|
|
| 5 |
A' |
3034 |
2911 |
7.01 |
|
|
|
| 6 |
A' |
2714 |
2604 |
5.93 |
|
|
|
| 7 |
A' |
1506 |
1445 |
8.45 |
|
|
|
| 8 |
A' |
1494 |
1434 |
1.61 |
|
|
|
| 9 |
A' |
1486 |
1425 |
1.48 |
|
|
|
| 10 |
A' |
1481 |
1421 |
1.15 |
|
|
|
| 11 |
A' |
1410 |
1353 |
4.52 |
|
|
|
| 12 |
A' |
1401 |
1345 |
4.80 |
|
|
|
| 13 |
A' |
1339 |
1285 |
12.77 |
|
|
|
| 14 |
A' |
1258 |
1207 |
23.06 |
|
|
|
| 15 |
A' |
1144 |
1098 |
2.60 |
|
|
|
| 16 |
A' |
1092 |
1048 |
0.42 |
|
|
|
| 17 |
A' |
1055 |
1012 |
0.32 |
|
|
|
| 18 |
A' |
936 |
898 |
3.09 |
|
|
|
| 19 |
A' |
861 |
826 |
1.60 |
|
|
|
| 20 |
A' |
763 |
732 |
3.51 |
|
|
|
| 21 |
A' |
392 |
376 |
0.74 |
|
|
|
| 22 |
A' |
323 |
310 |
0.84 |
|
|
|
| 23 |
A' |
153 |
146 |
1.43 |
|
|
|
| 24 |
A" |
3130 |
3002 |
34.47 |
|
|
|
| 25 |
A" |
3120 |
2993 |
36.06 |
|
|
|
| 26 |
A" |
3087 |
2962 |
15.28 |
|
|
|
| 27 |
A" |
3064 |
2939 |
2.08 |
|
|
|
| 28 |
A" |
1495 |
1435 |
9.15 |
|
|
|
| 29 |
A" |
1334 |
1280 |
0.24 |
|
|
|
| 30 |
A" |
1309 |
1256 |
1.07 |
|
|
|
| 31 |
A" |
1229 |
1179 |
1.19 |
|
|
|
| 32 |
A" |
1083 |
1039 |
2.68 |
|
|
|
| 33 |
A" |
929 |
891 |
3.20 |
|
|
|
| 34 |
A" |
795 |
763 |
0.27 |
|
|
|
| 35 |
A" |
744 |
714 |
5.33 |
|
|
|
| 36 |
A" |
248 |
238 |
0.01 |
|
|
|
| 37 |
A" |
176 |
169 |
18.93 |
|
|
|
| 38 |
A" |
111 |
107 |
1.55 |
|
|
|
| 39 |
A" |
94 |
90 |
5.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29037.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27858.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.053 |
|
|
|
| 2 |
C |
-0.409 |
|
|
|
| 3 |
C |
-0.274 |
|
|
|
| 4 |
C |
-0.271 |
|
|
|
| 5 |
C |
-0.334 |
|
|
|
| 6 |
H |
0.083 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
| 15 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.857 |
0.230 |
0.000 |
1.872 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.056 |
2.178 |
0.000 |
| y |
2.178 |
-38.274 |
0.000 |
| z |
0.000 |
0.000 |
-42.367 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.736 |
2.178 |
0.000 |
| y |
2.178 |
4.937 |
0.000 |
| z |
0.000 |
0.000 |
-1.201 |
|
| Polar |
|---|
| 3z2-r2 | -2.403 |
|---|
| x2-y2 | -5.782 |
|---|
| xy | 2.178 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.742 |
-1.541 |
0.000 |
| y |
-1.541 |
11.360 |
0.000 |
| z |
0.000 |
0.000 |
7.518 |
<r2> (average value of r
2) Å
2
| <r2> |
261.602 |
| (<r2>)1/2 |
16.174 |