return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: B2PLYP=FULLultrafine/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULLultrafine/6-311G**
 hartrees
Energy at 0K-213.634579
Energy at 298.15K 
HF Energy-213.345548
Nuclear repulsion energy188.413976
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3512 3512 0.43      
2 A' 3140 3140 48.74      
3 A' 3119 3119 75.43      
4 A' 3064 3064 55.23      
5 A' 3051 3051 12.57      
6 A' 2946 2946 182.70      
7 A' 1545 1545 1.34      
8 A' 1518 1518 2.71      
9 A' 1505 1505 14.66      
10 A' 1443 1443 3.59      
11 A' 1410 1410 1.18      
12 A' 1333 1333 4.48      
13 A' 1247 1247 1.31      
14 A' 1177 1177 11.65      
15 A' 1068 1068 6.58      
16 A' 914 914 7.40      
17 A' 836 836 0.89      
18 A' 790 790 71.98      
19 A' 427 427 0.21      
20 A' 264 264 0.76      
21 A' 186 186 0.68      
22 A' 109 109 1.24      
23 A" 3140 3140 11.80      
24 A" 3119 3119 18.19      
25 A" 3061 3061 2.14      
26 A" 3050 3050 31.49      
27 A" 2942 2942 12.67      
28 A" 1534 1534 5.04      
29 A" 1519 1519 4.57      
30 A" 1513 1513 12.36      
31 A" 1492 1492 15.64      
32 A" 1423 1423 13.99      
33 A" 1371 1371 25.96      
34 A" 1298 1298 4.05      
35 A" 1171 1171 51.01      
36 A" 1125 1125 9.79      
37 A" 1077 1077 1.74      
38 A" 953 953 0.48      
39 A" 819 819 0.30      
40 A" 429 429 0.57      
41 A" 263 263 0.74      
42 A" 108 108 1.21      

Unscaled Zero Point Vibrational Energy (zpe) 33005.3 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 33005.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/6-311G**
ABC
0.59112 0.07004 0.06601

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.280 0.000
C2 0.017 0.520 1.220
C3 0.017 0.520 -1.220
C4 0.017 -0.374 2.454
C5 0.017 -0.374 -2.454
H6 -0.807 -0.874 0.000
H7 -0.835 1.221 1.267
H8 0.924 1.134 1.213
H9 -0.835 1.221 -1.267
H10 0.924 1.134 -1.213
H11 0.074 0.220 3.368
H12 -0.896 -0.974 2.502
H13 0.868 -1.056 2.423
H14 0.074 0.220 -3.368
H15 -0.896 -0.974 -2.502
H16 0.868 -1.056 -2.423

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.45941.45942.45552.45551.01612.14152.07202.14152.07203.40582.75222.68303.40582.75222.6830
C21.45942.44071.52343.78132.02801.10431.09482.72142.66822.16952.16992.15784.59884.11354.0601
C31.45942.44073.78131.52342.02802.72142.66821.10431.09484.59884.11354.06012.16952.16992.1578
C42.45551.52343.78134.90752.63652.16272.15304.13724.06701.09211.09391.09125.85265.07474.9975
C52.45553.78131.52344.90752.63654.13724.06702.16272.15305.85265.07474.99751.09211.09391.0912
H61.01612.02802.02802.63652.63652.44862.91552.44862.91553.64952.50542.95183.64952.50542.9518
H72.14151.10432.72142.16274.13722.44861.76152.53473.04162.49812.51873.06964.82864.36204.6590
H82.07201.09482.66822.15304.06702.91551.76153.04162.42582.49073.06862.50314.74824.64274.2453
H92.14152.72141.10434.13722.16272.44862.53473.04161.76154.82864.36204.65902.49812.51873.0696
H102.07202.66821.09484.06702.15302.91553.04162.42581.76154.74824.64274.24532.49073.06862.5031
H113.40582.16954.59881.09215.85263.64952.49812.49074.82864.74821.76551.77606.73666.06845.9836
H122.75222.16994.11351.09395.07472.50542.51873.06864.36204.64271.76551.76836.06845.00375.2324
H132.68302.15784.06011.09124.99752.95183.06962.50314.65904.24531.77601.76835.98365.23244.8466
H143.40584.59882.16955.85261.09213.64954.82864.74822.49812.49076.73666.06845.98361.76551.7760
H152.75224.11352.16995.07471.09392.50544.36204.64272.51873.06866.06845.00375.23241.76551.7683
H162.68304.06012.15784.99751.09122.95184.65904.24533.06962.50315.98365.23244.84661.77601.7683

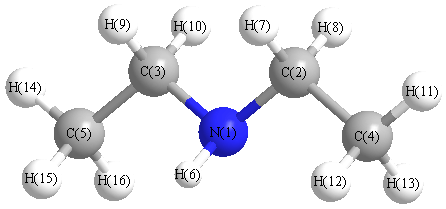
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.804 N1 C2 H7 112.558
N1 C2 H8 107.576 N1 C3 C5 110.804
N1 C3 H9 112.558 N1 C3 H10 107.576
C2 N1 C3 113.481 C2 N1 H6 108.693
C2 C4 H11 111.017 C2 C4 H12 110.939
C2 C4 H13 110.140 C3 N1 H6 108.693
C3 C5 H14 111.017 C3 C5 H15 110.939
C3 C5 H16 110.140 C4 C2 H7 109.753
C4 C2 H8 109.554 C5 C3 H8 150.863
C5 C3 H10 109.554 H7 C2 H8 106.449
H9 C3 H10 106.449 H11 C4 H12 107.727
H11 C4 H13 108.872 H12 C4 H13 108.051
H14 C5 H15 107.727 H14 C5 H16 108.872
H15 C5 H16 108.051
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability