Jump to
S1C2
S1C3
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -209.164534 |
| Energy at 298.15K | -209.170531 |
| HF Energy | -209.164534 |
| Nuclear repulsion energy | 119.569146 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3647 |
3647 |
36.87 |
|
|
|
| 2 |
A' |
3134 |
3134 |
13.08 |
|
|
|
| 3 |
A' |
3044 |
3044 |
43.69 |
|
|
|
| 4 |
A' |
2985 |
2985 |
74.79 |
|
|
|
| 5 |
A' |
1859 |
1859 |
535.60 |
|
|
|
| 6 |
A' |
1545 |
1545 |
30.77 |
|
|
|
| 7 |
A' |
1503 |
1503 |
10.17 |
|
|
|
| 8 |
A' |
1476 |
1476 |
7.10 |
|
|
|
| 9 |
A' |
1411 |
1411 |
16.36 |
|
|
|
| 10 |
A' |
1311 |
1311 |
117.28 |
|
|
|
| 11 |
A' |
1185 |
1185 |
28.09 |
|
|
|
| 12 |
A' |
1024 |
1024 |
49.84 |
|
|
|
| 13 |
A' |
625 |
625 |
15.34 |
|
|
|
| 14 |
A' |
357 |
357 |
8.17 |
|
|
|
| 15 |
A" |
3107 |
3107 |
22.32 |
|
|
|
| 16 |
A" |
1491 |
1491 |
7.54 |
|
|
|
| 17 |
A" |
1150 |
1150 |
0.46 |
|
|
|
| 18 |
A" |
1068 |
1068 |
0.31 |
|
|
|
| 19 |
A" |
618 |
618 |
132.74 |
|
|
|
| 20 |
A" |
194 |
194 |
1.05 |
|
|
|
| 21 |
A" |
58 |
58 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16395.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16395.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.290 |
-0.755 |
0.000 |
| O2 |
1.401 |
-1.221 |
0.000 |
| N3 |
0.000 |
0.572 |
0.000 |
| C4 |
-1.337 |
1.122 |
0.000 |
| H5 |
-0.624 |
-1.379 |
0.000 |
| H6 |
0.797 |
1.192 |
0.000 |
| H7 |
-2.051 |
0.297 |
0.000 |
| H8 |
-1.522 |
1.729 |
0.889 |
| H9 |
-1.522 |
1.729 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2050 | 1.3579 | 2.4839 | 1.1066 | 2.0121 | 2.5659 | 3.2001 | 3.2001 |
O2 | 1.2050 | | 2.2752 | 3.6038 | 2.0312 | 2.4879 | 3.7706 | 4.2467 | 4.2467 | N3 | 1.3579 | 2.2752 | | 1.4461 | 2.0482 | 1.0098 | 2.0691 | 2.1084 | 2.1084 | C4 | 2.4839 | 3.6038 | 1.4461 | | 2.6008 | 2.1351 | 1.0909 | 1.0922 | 1.0922 | H5 | 1.1066 | 2.0312 | 2.0482 | 2.6008 | | 2.9378 | 2.2008 | 3.3549 | 3.3549 | H6 | 2.0121 | 2.4879 | 1.0098 | 2.1351 | 2.9378 | | 2.9849 | 2.5403 | 2.5403 | H7 | 2.5659 | 3.7706 | 2.0691 | 1.0909 | 2.2008 | 2.9849 | | 1.7667 | 1.7667 | H8 | 3.2001 | 4.2467 | 2.1084 | 1.0922 | 3.3549 | 2.5403 | 1.7667 | | 1.7784 | H9 | 3.2001 | 4.2467 | 2.1084 | 1.0922 | 3.3549 | 2.5403 | 1.7667 | 1.7784 | |
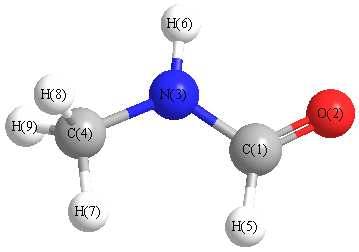 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.682 |
|
C1 |
N3 |
H6 |
115.604 |
| O2 |
C1 |
N3 |
125.075 |
|
O2 |
C1 |
H5 |
122.907 |
| N3 |
C1 |
H5 |
112.018 |
|
N3 |
C4 |
H7 |
108.479 |
| N3 |
C4 |
H8 |
111.565 |
|
N3 |
C4 |
H9 |
111.565 |
| C4 |
N3 |
H6 |
119.714 |
|
H7 |
C4 |
H8 |
108.051 |
| H7 |
C4 |
H9 |
108.051 |
|
H8 |
C4 |
H9 |
109.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.307 |
|
|
|
| 2 |
O |
-0.391 |
|
|
|
| 3 |
N |
-0.447 |
|
|
|
| 4 |
C |
-0.224 |
|
|
|
| 5 |
H |
0.073 |
|
|
|
| 6 |
H |
0.250 |
|
|
|
| 7 |
H |
0.135 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.097 |
2.705 |
0.000 |
4.111 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.386 |
4.024 |
0.000 |
| y |
4.024 |
-24.250 |
0.000 |
| z |
0.000 |
0.000 |
-24.785 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.868 |
4.024 |
0.000 |
| y |
4.024 |
1.336 |
0.000 |
| z |
0.000 |
0.000 |
0.532 |
|
| Polar |
|---|
| 3z2-r2 | 1.065 |
|---|
| x2-y2 | -2.136 |
|---|
| xy | 4.024 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.619 |
-1.040 |
0.000 |
| y |
-1.040 |
5.593 |
0.000 |
| z |
0.000 |
0.000 |
3.276 |
<r2> (average value of r
2) Å
2
| <r2> |
88.656 |
| (<r2>)1/2 |
9.416 |
Jump to
S1C1
S1C3
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -209.164534 |
| Energy at 298.15K | -209.170531 |
| HF Energy | -209.164534 |
| Nuclear repulsion energy | 119.569146 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -209.166828 |
| Energy at 298.15K | -209.172984 |
| HF Energy | -209.166828 |
| Nuclear repulsion energy | 121.818261 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3690 |
3690 |
39.59 |
|
|
|
| 2 |
A |
3174 |
3174 |
0.28 |
|
|
|
| 3 |
A |
3129 |
3129 |
24.53 |
|
|
|
| 4 |
A |
3065 |
3065 |
35.48 |
|
|
|
| 5 |
A |
3006 |
3006 |
97.48 |
|
|
|
| 6 |
A |
1840 |
1840 |
375.60 |
|
|
|
| 7 |
A |
1573 |
1573 |
130.64 |
|
|
|
| 8 |
A |
1514 |
1514 |
9.08 |
|
|
|
| 9 |
A |
1505 |
1505 |
35.52 |
|
|
|
| 10 |
A |
1455 |
1455 |
20.38 |
|
|
|
| 11 |
A |
1431 |
1431 |
3.04 |
|
|
|
| 12 |
A |
1238 |
1238 |
81.50 |
|
|
|
| 13 |
A |
1173 |
1173 |
18.80 |
|
|
|
| 14 |
A |
1159 |
1159 |
0.69 |
|
|
|
| 15 |
A |
1045 |
1045 |
0.33 |
|
|
|
| 16 |
A |
985 |
985 |
25.10 |
|
|
|
| 17 |
A |
783 |
783 |
1.63 |
|
|
|
| 18 |
A |
557 |
557 |
53.29 |
|
|
|
| 19 |
A |
319 |
319 |
13.74 |
|
|
|
| 20 |
A |
285 |
285 |
77.23 |
|
|
|
| 21 |
A |
89 |
89 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16508.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16508.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.864 |
0.433 |
0.003 |
| O2 |
1.352 |
-0.674 |
0.000 |
| N3 |
-0.467 |
0.664 |
-0.020 |
| C4 |
-1.410 |
-0.441 |
0.005 |
| H5 |
1.469 |
1.353 |
0.018 |
| H6 |
-0.797 |
1.611 |
0.059 |
| H7 |
-2.408 |
-0.063 |
-0.210 |
| H8 |
-1.127 |
-1.172 |
-0.753 |
| H9 |
-1.410 |
-0.939 |
0.977 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2105 | 1.3511 | 2.4353 | 1.1016 | 2.0367 | 3.3155 | 2.6663 | 2.8286 |
O2 | 1.2105 | | 2.2592 | 2.7719 | 2.0312 | 3.1380 | 3.8153 | 2.6388 | 2.9420 | N3 | 1.3511 | 2.2592 | | 1.4522 | 2.0555 | 1.0059 | 2.0808 | 2.0840 | 2.1101 | C4 | 2.4353 | 2.7719 | 1.4522 | | 3.3917 | 2.1423 | 1.0886 | 1.0900 | 1.0925 | H5 | 1.1016 | 2.0312 | 2.0555 | 3.3917 | | 2.2804 | 4.1334 | 3.7026 | 3.8030 | H6 | 2.0367 | 3.1380 | 1.0059 | 2.1423 | 2.2804 | | 2.3392 | 2.9179 | 2.7790 | H7 | 3.3155 | 3.8153 | 2.0808 | 1.0886 | 4.1334 | 2.3392 | | 1.7785 | 1.7811 | H8 | 2.6663 | 2.6388 | 2.0840 | 1.0900 | 3.7026 | 2.9179 | 1.7785 | | 1.7680 | H9 | 2.8286 | 2.9420 | 2.1101 | 1.0925 | 3.8030 | 2.7790 | 1.7811 | 1.7680 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.576 |
|
C1 |
N3 |
H6 |
118.829 |
| O2 |
C1 |
N3 |
123.661 |
|
O2 |
C1 |
H5 |
122.855 |
| N3 |
C1 |
H5 |
113.482 |
|
N3 |
C4 |
H7 |
109.123 |
| N3 |
C4 |
H8 |
109.297 |
|
N3 |
C4 |
H9 |
111.253 |
| C4 |
N3 |
H6 |
120.180 |
|
H7 |
C4 |
H8 |
109.443 |
| H7 |
C4 |
H9 |
109.492 |
|
H8 |
C4 |
H9 |
108.210 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.301 |
|
|
|
| 2 |
O |
-0.398 |
|
|
|
| 3 |
N |
-0.455 |
|
|
|
| 4 |
C |
-0.243 |
|
|
|
| 5 |
H |
0.089 |
|
|
|
| 6 |
H |
0.246 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
H |
0.169 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.825 |
2.637 |
0.000 |
3.864 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.426 |
2.771 |
0.000 |
| y |
2.771 |
-22.094 |
0.001 |
| z |
0.000 |
0.001 |
-24.812 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.973 |
2.771 |
0.000 |
| y |
2.771 |
3.025 |
0.001 |
| z |
0.000 |
0.001 |
-1.052 |
|
| Polar |
|---|
| 3z2-r2 | -2.103 |
|---|
| x2-y2 | -3.332 |
|---|
| xy | 2.771 |
|---|
| xz | 0.000 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.949 |
-0.123 |
0.000 |
| y |
-0.123 |
4.989 |
0.000 |
| z |
0.000 |
0.000 |
3.265 |
<r2> (average value of r
2) Å
2
| <r2> |
77.943 |
| (<r2>)1/2 |
8.829 |
