Jump to
S1C2
Energy calculated at QCISD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -1194.199086 |
| Energy at 298.15K | -1194.201437 |
| HF Energy | -1193.762352 |
| Nuclear repulsion energy | 193.679784 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2735 |
2609 |
0.37 |
|
|
|
| 2 |
A |
901 |
860 |
0.90 |
|
|
|
| 3 |
A |
477 |
455 |
0.70 |
|
|
|
| 4 |
A |
307 |
293 |
25.66 |
|
|
|
| 5 |
A |
206 |
197 |
0.02 |
|
|
|
| 6 |
B |
2733 |
2608 |
10.50 |
|
|
|
| 7 |
B |
888 |
847 |
15.83 |
|
|
|
| 8 |
B |
482 |
460 |
18.82 |
|
|
|
| 9 |
B |
341 |
325 |
18.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4534.9 cm
-1
Scaled (by 0.9541) Zero Point Vibrational Energy (zpe) 4326.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.853 |
| S2 |
0.000 |
1.680 |
-0.391 |
| S3 |
0.000 |
-1.680 |
-0.391 |
| H4 |
-1.324 |
1.771 |
-0.577 |
| H5 |
1.324 |
-1.771 |
-0.577 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0904 | 2.0904 | 2.6340 | 2.6340 |
S2 | 2.0904 | | 3.3601 | 1.3405 | 3.7015 | S3 | 2.0904 | 3.3601 | | 3.7015 | 1.3405 | H4 | 2.6340 | 1.3405 | 3.7015 | | 4.4234 | H5 | 2.6340 | 3.7015 | 1.3405 | 4.4234 | |
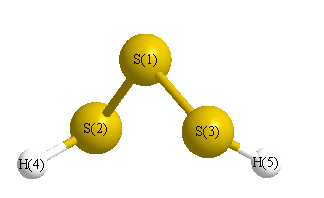 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.908 |
|
S1 |
S3 |
H5 |
97.908 |
| S2 |
S1 |
S3 |
106.967 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -1194.198746 |
| Energy at 298.15K | -1194.201104 |
| HF Energy | -1193.761832 |
| Nuclear repulsion energy | 193.692995 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2729 |
2604 |
15.15 |
|
|
|
| 2 |
A' |
903 |
862 |
8.34 |
|
|
|
| 3 |
A' |
477 |
455 |
0.64 |
|
|
|
| 4 |
A' |
341 |
325 |
19.76 |
|
|
|
| 5 |
A' |
207 |
197 |
0.21 |
|
|
|
| 6 |
A" |
2730 |
2605 |
0.56 |
|
|
|
| 7 |
A" |
894 |
853 |
9.79 |
|
|
|
| 8 |
A" |
483 |
460 |
20.40 |
|
|
|
| 9 |
A" |
311 |
296 |
10.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4536.8 cm
-1
Scaled (by 0.9541) Zero Point Vibrational Energy (zpe) 4328.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.852 |
0.000 |
| S2 |
-0.053 |
-0.393 |
1.679 |
| S3 |
-0.053 |
-0.393 |
-1.679 |
| H4 |
1.275 |
-0.525 |
1.810 |
| H5 |
1.275 |
-0.525 |
-1.810 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0903 | 2.0903 | 2.6332 | 2.6332 |
S2 | 2.0903 | | 3.3585 | 1.3409 | 3.7356 | S3 | 2.0903 | 3.3585 | | 3.7356 | 1.3409 | H4 | 2.6332 | 1.3409 | 3.7356 | | 3.6198 | H5 | 2.6332 | 3.7356 | 1.3409 | 3.6198 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.855 |
|
S1 |
S3 |
H5 |
97.855 |
| S2 |
S1 |
S3 |
106.903 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
