Jump to
S1C2
S1C3
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -210.353673 |
| Energy at 298.15K | -210.362945 |
| HF Energy | -210.353673 |
| Nuclear repulsion energy | 134.986695 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3809 |
3809 |
79.44 |
|
|
|
| 2 |
A |
3627 |
3627 |
6.04 |
|
|
|
| 3 |
A |
3539 |
3539 |
1.53 |
|
|
|
| 4 |
A |
3108 |
3108 |
36.53 |
|
|
|
| 5 |
A |
3099 |
3099 |
25.25 |
|
|
|
| 6 |
A |
3023 |
3023 |
57.72 |
|
|
|
| 7 |
A |
2992 |
2992 |
61.82 |
|
|
|
| 8 |
A |
1660 |
1660 |
36.07 |
|
|
|
| 9 |
A |
1523 |
1523 |
0.73 |
|
|
|
| 10 |
A |
1502 |
1502 |
6.74 |
|
|
|
| 11 |
A |
1460 |
1460 |
78.70 |
|
|
|
| 12 |
A |
1410 |
1410 |
9.39 |
|
|
|
| 13 |
A |
1384 |
1384 |
2.98 |
|
|
|
| 14 |
A |
1332 |
1332 |
2.37 |
|
|
|
| 15 |
A |
1264 |
1264 |
26.22 |
|
|
|
| 16 |
A |
1201 |
1201 |
2.63 |
|
|
|
| 17 |
A |
1146 |
1146 |
77.48 |
|
|
|
| 18 |
A |
1093 |
1093 |
22.98 |
|
|
|
| 19 |
A |
1014 |
1014 |
9.36 |
|
|
|
| 20 |
A |
943 |
943 |
61.41 |
|
|
|
| 21 |
A |
900 |
900 |
10.55 |
|
|
|
| 22 |
A |
836 |
836 |
88.27 |
|
|
|
| 23 |
A |
590 |
590 |
136.01 |
|
|
|
| 24 |
A |
544 |
544 |
12.40 |
|
|
|
| 25 |
A |
340 |
340 |
0.78 |
|
|
|
| 26 |
A |
297 |
297 |
12.27 |
|
|
|
| 27 |
A |
206 |
206 |
5.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21920.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21920.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.352 |
-0.559 |
0.122 |
| C2 |
-0.630 |
0.655 |
-0.275 |
| C3 |
0.788 |
0.553 |
0.271 |
| O4 |
1.396 |
-0.636 |
-0.171 |
| H5 |
-1.646 |
-0.498 |
1.091 |
| H6 |
-2.186 |
-0.691 |
-0.436 |
| H7 |
-1.103 |
1.588 |
0.058 |
| H8 |
-0.577 |
0.670 |
-1.366 |
| H9 |
1.395 |
1.391 |
-0.077 |
| H10 |
0.756 |
0.596 |
1.371 |
| H11 |
0.699 |
-1.302 |
-0.108 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4671 | 2.4163 | 2.7646 | 1.0144 | 1.0121 | 2.1624 | 2.0793 | 3.3747 | 2.7090 | 2.1930 |
C2 | 1.4671 | | 1.5232 | 2.4050 | 2.0553 | 2.0638 | 1.0978 | 1.0925 | 2.1638 | 2.1528 | 2.3712 | C3 | 2.4163 | 1.5232 | | 1.4067 | 2.7746 | 3.3006 | 2.1667 | 2.1350 | 1.0919 | 1.1014 | 1.8952 | O4 | 2.7646 | 2.4050 | 1.4067 | | 3.2961 | 3.5925 | 3.3536 | 2.6511 | 2.0296 | 2.0750 | 0.9661 | H5 | 1.0144 | 2.0553 | 2.7746 | 3.2961 | | 1.6309 | 2.3894 | 2.9222 | 3.7651 | 2.6538 | 2.7534 | H6 | 1.0121 | 2.0638 | 3.3006 | 3.5925 | 1.6309 | | 2.5712 | 2.3034 | 4.1581 | 3.6849 | 2.9668 | H7 | 2.1624 | 1.0978 | 2.1667 | 3.3536 | 2.3894 | 2.5712 | | 1.7742 | 2.5097 | 2.4831 | 3.4097 | H8 | 2.0793 | 1.0925 | 2.1350 | 2.6511 | 2.9222 | 2.3034 | 1.7742 | | 2.4642 | 3.0456 | 2.6640 | H9 | 3.3747 | 2.1638 | 1.0919 | 2.0296 | 3.7651 | 4.1581 | 2.5097 | 2.4642 | | 1.7713 | 2.7818 | H10 | 2.7090 | 2.1528 | 1.1014 | 2.0750 | 2.6538 | 3.6849 | 2.4831 | 3.0456 | 1.7713 | | 2.4070 | H11 | 2.1930 | 2.3712 | 1.8952 | 0.9661 | 2.7534 | 2.9668 | 3.4097 | 2.6640 | 2.7818 | 2.4070 | |
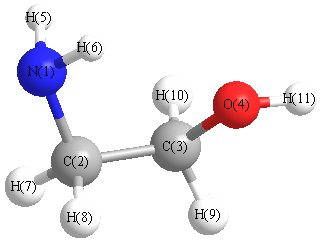 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.796 |
|
N1 |
C2 |
H7 |
114.159 |
| N1 |
C2 |
H8 |
107.755 |
|
C2 |
N1 |
H5 |
110.507 |
| C2 |
N1 |
H6 |
111.379 |
|
C2 |
C3 |
O4 |
110.276 |
| C2 |
C3 |
H9 |
110.590 |
|
C2 |
C3 |
H10 |
109.164 |
| C3 |
C2 |
H7 |
110.466 |
|
C3 |
C2 |
H8 |
108.295 |
| C3 |
O4 |
H11 |
104.477 |
|
O4 |
C3 |
H9 |
107.981 |
| O4 |
C3 |
H10 |
111.070 |
|
H5 |
N1 |
H6 |
107.180 |
| H7 |
C2 |
H8 |
108.197 |
|
H9 |
C3 |
H10 |
107.726 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.545 |
|
|
|
| 2 |
C |
-0.230 |
|
|
|
| 3 |
C |
-0.031 |
|
|
|
| 4 |
O |
-0.427 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
H |
0.231 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
| 11 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.942 |
1.132 |
0.596 |
3.208 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.391 |
2.063 |
0.375 |
| y |
2.063 |
-25.426 |
-0.238 |
| z |
0.375 |
-0.238 |
-24.542 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.407 |
2.063 |
0.375 |
| y |
2.063 |
0.041 |
-0.238 |
| z |
0.375 |
-0.238 |
1.366 |
|
| Polar |
|---|
| 3z2-r2 | 2.732 |
|---|
| x2-y2 | -0.965 |
|---|
| xy | 2.063 |
|---|
| xz | 0.375 |
|---|
| yz | -0.238 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.476 |
0.246 |
0.034 |
| y |
0.246 |
5.230 |
0.040 |
| z |
0.034 |
0.040 |
4.839 |
<r2> (average value of r
2) Å
2
| <r2> |
86.332 |
| (<r2>)1/2 |
9.291 |
