Jump to
S1C2
Vibrational Frequencies calculated at CCSD/6-311G**
Geometric Data calculated at CCSD/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -2611.637567 |
| Energy at 298.15K | |
| HF Energy | -2611.360289 |
| Nuclear repulsion energy | 80.434597 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3200 |
3200 |
6.14 |
|
|
|
| 2 |
A1 |
1426 |
1426 |
23.99 |
|
|
|
| 3 |
A1 |
715 |
715 |
15.67 |
|
|
|
| 4 |
B1 |
160i |
160i |
76.46 |
|
|
|
| 5 |
B2 |
3351 |
3351 |
0.00 |
|
|
|
| 6 |
B2 |
952 |
952 |
1.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4741.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4741.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.490 |
| Br2 |
0.000 |
0.000 |
0.370 |
| H3 |
0.000 |
0.954 |
-1.997 |
| H4 |
0.000 |
-0.954 |
-1.997 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8595 | 1.0801 | 1.0801 |
Br2 | 1.8595 | | 2.5516 | 2.5516 | H3 | 1.0801 | 2.5516 | | 1.9073 | H4 | 1.0801 | 2.5516 | 1.9073 | |
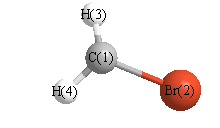 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
118.001 |
|
Br2 |
C1 |
H4 |
118.001 |
| H3 |
C1 |
H4 |
123.998 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
