Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -247.220936 |
| Energy at 298.15K | -247.226600 |
| HF Energy | -247.220936 |
| Nuclear repulsion energy | 157.731201 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3156 |
3156 |
11.09 |
|
|
|
| 2 |
A' |
3032 |
3032 |
25.64 |
|
|
|
| 3 |
A' |
3024 |
3024 |
34.64 |
|
|
|
| 4 |
A' |
2438 |
2438 |
0.49 |
|
|
|
| 5 |
A' |
1527 |
1527 |
7.79 |
|
|
|
| 6 |
A' |
1505 |
1505 |
9.11 |
|
|
|
| 7 |
A' |
1490 |
1490 |
0.23 |
|
|
|
| 8 |
A' |
1430 |
1430 |
34.63 |
|
|
|
| 9 |
A' |
1252 |
1252 |
80.19 |
|
|
|
| 10 |
A' |
1198 |
1198 |
100.44 |
|
|
|
| 11 |
A' |
1036 |
1036 |
36.81 |
|
|
|
| 12 |
A' |
969 |
969 |
7.27 |
|
|
|
| 13 |
A' |
561 |
561 |
1.56 |
|
|
|
| 14 |
A' |
392 |
392 |
2.03 |
|
|
|
| 15 |
A' |
185 |
185 |
2.16 |
|
|
|
| 16 |
A" |
3085 |
3085 |
37.68 |
|
|
|
| 17 |
A" |
3063 |
3063 |
23.75 |
|
|
|
| 18 |
A" |
1498 |
1498 |
11.03 |
|
|
|
| 19 |
A" |
1284 |
1284 |
3.79 |
|
|
|
| 20 |
A" |
1194 |
1194 |
3.03 |
|
|
|
| 21 |
A" |
1040 |
1040 |
2.09 |
|
|
|
| 22 |
A" |
391 |
391 |
1.58 |
|
|
|
| 23 |
A" |
235 |
235 |
4.67 |
|
|
|
| 24 |
A" |
97 |
97 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17540.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17540.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.122 |
-0.350 |
0.000 |
| O2 |
-0.734 |
-0.606 |
0.000 |
| C3 |
0.000 |
0.585 |
0.000 |
| C4 |
1.432 |
0.267 |
0.000 |
| N5 |
2.558 |
0.047 |
0.000 |
| H6 |
-2.622 |
-1.316 |
0.000 |
| H7 |
-2.420 |
0.213 |
0.893 |
| H8 |
-2.420 |
0.213 |
-0.893 |
| H9 |
-0.218 |
1.194 |
-0.888 |
| H10 |
-0.218 |
1.194 |
0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4111 | 2.3186 | 3.6074 | 4.6966 | 1.0877 | 1.0970 | 1.0970 | 2.6069 | 2.6069 |
O2 | 1.4111 | | 1.3988 | 2.3357 | 3.3561 | 2.0172 | 2.0764 | 2.0764 | 2.0719 | 2.0719 | C3 | 2.3186 | 1.3988 | | 1.4673 | 2.6139 | 3.2386 | 2.6066 | 2.6066 | 1.0986 | 1.0986 | C4 | 3.6074 | 2.3357 | 1.4673 | | 1.1468 | 4.3526 | 3.9554 | 3.9554 | 2.0908 | 2.0908 | N5 | 4.6966 | 3.3561 | 2.6139 | 1.1468 | | 5.3564 | 5.0606 | 5.0606 | 3.1319 | 3.1319 | H6 | 1.0877 | 2.0172 | 3.2386 | 4.3526 | 5.3564 | | 1.7817 | 1.7817 | 3.5868 | 3.5868 | H7 | 1.0970 | 2.0764 | 2.6066 | 3.9554 | 5.0606 | 1.7817 | | 1.7862 | 2.9977 | 2.4109 | H8 | 1.0970 | 2.0764 | 2.6066 | 3.9554 | 5.0606 | 1.7817 | 1.7862 | | 2.4109 | 2.9977 | H9 | 2.6069 | 2.0719 | 1.0986 | 2.0908 | 3.1319 | 3.5868 | 2.9977 | 2.4109 | | 1.7769 | H10 | 2.6069 | 2.0719 | 1.0986 | 2.0908 | 3.1319 | 3.5868 | 2.4109 | 2.9977 | 1.7769 | |
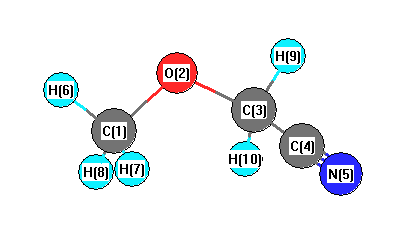 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.210 |
|
O2 |
C1 |
H6 |
106.945 |
| O2 |
C1 |
H7 |
111.142 |
|
O2 |
C1 |
H8 |
111.142 |
| O2 |
C3 |
C4 |
109.138 |
|
O2 |
C3 |
H9 |
111.552 |
| O2 |
C3 |
H10 |
111.552 |
|
C3 |
C4 |
N5 |
178.537 |
| C4 |
C3 |
H9 |
108.279 |
|
C4 |
C3 |
H10 |
108.279 |
| H6 |
C1 |
H7 |
109.283 |
|
H6 |
C1 |
H8 |
109.283 |
| H7 |
C1 |
H8 |
108.999 |
|
H9 |
C3 |
H10 |
107.932 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.188 |
|
|
|
| 2 |
O |
-0.355 |
|
|
|
| 3 |
C |
0.002 |
|
|
|
| 4 |
C |
0.068 |
|
|
|
| 5 |
N |
-0.263 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
| 9 |
H |
0.164 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.426 |
1.857 |
0.000 |
4.800 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.935 |
0.709 |
0.000 |
| y |
0.709 |
-29.379 |
0.000 |
| z |
0.000 |
0.000 |
-29.271 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.610 |
0.709 |
0.000 |
| y |
0.709 |
3.223 |
0.000 |
| z |
0.000 |
0.000 |
3.386 |
|
| Polar |
|---|
| 3z2-r2 | 6.773 |
|---|
| x2-y2 | -6.555 |
|---|
| xy | 0.709 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.076 |
0.004 |
0.000 |
| y |
0.004 |
4.792 |
0.000 |
| z |
0.000 |
0.000 |
4.494 |
<r2> (average value of r
2) Å
2
| <r2> |
141.827 |
| (<r2>)1/2 |
11.909 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -247.224123 |
| Energy at 298.15K | -247.229892 |
| HF Energy | -247.224123 |
| Nuclear repulsion energy | 160.786214 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3173 |
12.78 |
|
|
|
| 2 |
A |
3141 |
3141 |
2.26 |
|
|
|
| 3 |
A |
3097 |
3097 |
26.45 |
|
|
|
| 4 |
A |
3044 |
3044 |
22.99 |
|
|
|
| 5 |
A |
3026 |
3026 |
32.18 |
|
|
|
| 6 |
A |
2415 |
2415 |
0.15 |
|
|
|
| 7 |
A |
1519 |
1519 |
11.91 |
|
|
|
| 8 |
A |
1505 |
1505 |
7.12 |
|
|
|
| 9 |
A |
1498 |
1498 |
5.52 |
|
|
|
| 10 |
A |
1481 |
1481 |
3.52 |
|
|
|
| 11 |
A |
1398 |
1398 |
15.48 |
|
|
|
| 12 |
A |
1326 |
1326 |
9.38 |
|
|
|
| 13 |
A |
1246 |
1246 |
85.77 |
|
|
|
| 14 |
A |
1200 |
1200 |
42.01 |
|
|
|
| 15 |
A |
1189 |
1189 |
27.57 |
|
|
|
| 16 |
A |
1044 |
1044 |
9.56 |
|
|
|
| 17 |
A |
981 |
981 |
31.79 |
|
|
|
| 18 |
A |
915 |
915 |
21.71 |
|
|
|
| 19 |
A |
617 |
617 |
1.65 |
|
|
|
| 20 |
A |
405 |
405 |
3.21 |
|
|
|
| 21 |
A |
376 |
376 |
1.08 |
|
|
|
| 22 |
A |
265 |
265 |
10.58 |
|
|
|
| 23 |
A |
188 |
188 |
4.31 |
|
|
|
| 24 |
A |
125 |
125 |
8.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17587.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17587.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.536 |
-0.780 |
0.135 |
| O2 |
1.081 |
0.435 |
-0.429 |
| C3 |
-0.055 |
0.937 |
0.206 |
| C4 |
-1.234 |
0.059 |
0.033 |
| N5 |
-2.136 |
-0.639 |
-0.096 |
| H6 |
2.439 |
-1.057 |
-0.404 |
| H7 |
0.788 |
-1.573 |
0.022 |
| H8 |
1.769 |
-0.653 |
1.199 |
| H9 |
-0.278 |
1.907 |
-0.239 |
| H10 |
0.106 |
1.073 |
1.283 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4152 | 2.3417 | 2.8963 | 3.6820 | 1.0875 | 1.0954 | 1.0968 | 3.2635 | 2.6073 |
O2 | 1.4152 | | 1.3947 | 2.3911 | 3.4084 | 2.0177 | 2.0788 | 2.0757 | 2.0123 | 2.0716 | C3 | 2.3417 | 1.3947 | | 1.4807 | 2.6282 | 3.2505 | 2.6537 | 2.6155 | 1.0900 | 1.0983 | C4 | 2.8963 | 2.3911 | 1.4807 | | 1.1477 | 3.8641 | 2.5986 | 3.2997 | 2.0984 | 2.0950 | N5 | 3.6820 | 3.4084 | 2.6282 | 1.1477 | | 4.6048 | 3.0719 | 4.1146 | 3.1552 | 3.1404 | H6 | 1.0875 | 2.0177 | 3.2505 | 3.8641 | 4.6048 | | 1.7816 | 1.7834 | 4.0241 | 3.5813 | H7 | 1.0954 | 2.0788 | 2.6537 | 2.5986 | 3.0719 | 1.7816 | | 1.7874 | 3.6485 | 3.0094 | H8 | 1.0968 | 2.0757 | 2.6155 | 3.2997 | 4.1146 | 1.7834 | 1.7874 | | 3.5790 | 2.3982 | H9 | 3.2635 | 2.0123 | 1.0900 | 2.0984 | 3.1552 | 4.0241 | 3.6485 | 3.5790 | | 1.7777 | H10 | 2.6073 | 2.0716 | 1.0983 | 2.0950 | 3.1404 | 3.5813 | 3.0094 | 2.3982 | 1.7777 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.898 |
|
O2 |
C1 |
H6 |
106.722 |
| O2 |
C1 |
H7 |
111.156 |
|
O2 |
C1 |
H8 |
110.811 |
| O2 |
C3 |
C4 |
112.487 |
|
O2 |
C3 |
H9 |
107.535 |
| O2 |
C3 |
H10 |
111.847 |
|
C3 |
C4 |
N5 |
178.901 |
| C4 |
C3 |
H9 |
108.475 |
|
C4 |
C3 |
H10 |
107.732 |
| H6 |
C1 |
H7 |
109.401 |
|
H6 |
C1 |
H8 |
109.461 |
| H7 |
C1 |
H8 |
109.242 |
|
H9 |
C3 |
H10 |
108.661 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.194 |
|
|
|
| 2 |
O |
-0.350 |
|
|
|
| 3 |
C |
-0.038 |
|
|
|
| 4 |
C |
0.064 |
|
|
|
| 5 |
N |
-0.266 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.187 |
|
|
|
| 10 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.475 |
1.217 |
1.452 |
3.117 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.282 |
-5.610 |
0.701 |
| y |
-5.610 |
-27.958 |
0.632 |
| z |
0.701 |
0.632 |
-29.852 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.377 |
-5.610 |
0.701 |
| y |
-5.610 |
5.109 |
0.632 |
| z |
0.701 |
0.632 |
2.268 |
|
| Polar |
|---|
| 3z2-r2 | 4.536 |
|---|
| x2-y2 | -8.324 |
|---|
| xy | -5.610 |
|---|
| xz | 0.701 |
|---|
| yz | 0.632 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.844 |
0.420 |
0.170 |
| y |
0.420 |
5.654 |
0.211 |
| z |
0.170 |
0.211 |
4.525 |
<r2> (average value of r
2) Å
2
| <r2> |
121.322 |
| (<r2>)1/2 |
11.015 |
