Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/6-311G**
| | hartrees |
|---|
| Energy at 0K | -247.117833 |
| Energy at 298.15K | |
| HF Energy | -246.827948 |
| Nuclear repulsion energy | 157.116855 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3169 |
3169 |
17.80 |
|
|
|
| 2 |
A' |
3010 |
3010 |
57.98 |
|
|
|
| 3 |
A' |
3002 |
3002 |
22.76 |
|
|
|
| 4 |
A' |
2306 |
2306 |
1.43 |
|
|
|
| 5 |
A' |
1533 |
1533 |
5.29 |
|
|
|
| 6 |
A' |
1513 |
1513 |
8.02 |
|
|
|
| 7 |
A' |
1493 |
1493 |
0.23 |
|
|
|
| 8 |
A' |
1429 |
1429 |
38.35 |
|
|
|
| 9 |
A' |
1236 |
1236 |
40.50 |
|
|
|
| 10 |
A' |
1164 |
1164 |
140.97 |
|
|
|
| 11 |
A' |
1000 |
1000 |
35.32 |
|
|
|
| 12 |
A' |
958 |
958 |
6.01 |
|
|
|
| 13 |
A' |
548 |
548 |
1.41 |
|
|
|
| 14 |
A' |
379 |
379 |
1.80 |
|
|
|
| 15 |
A' |
181 |
181 |
2.15 |
|
|
|
| 16 |
A" |
3064 |
3064 |
52.01 |
|
|
|
| 17 |
A" |
3033 |
3033 |
33.60 |
|
|
|
| 18 |
A" |
1501 |
1501 |
8.93 |
|
|
|
| 19 |
A" |
1274 |
1274 |
3.97 |
|
|
|
| 20 |
A" |
1191 |
1191 |
3.87 |
|
|
|
| 21 |
A" |
1042 |
1042 |
2.28 |
|
|
|
| 22 |
A" |
373 |
373 |
1.34 |
|
|
|
| 23 |
A" |
226 |
226 |
4.30 |
|
|
|
| 24 |
A" |
85 |
85 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17354.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17354.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.129 |
-0.361 |
0.000 |
| O2 |
-0.734 |
-0.609 |
0.000 |
| C3 |
0.000 |
0.590 |
0.000 |
| C4 |
1.431 |
0.275 |
0.000 |
| N5 |
2.570 |
0.055 |
0.000 |
| H6 |
-2.619 |
-1.332 |
0.000 |
| H7 |
-2.435 |
0.199 |
0.893 |
| H8 |
-2.435 |
0.199 |
-0.893 |
| H9 |
-0.221 |
1.199 |
-0.887 |
| H10 |
-0.221 |
1.199 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4168 | 2.3313 | 3.6166 | 4.7170 | 1.0876 | 1.0972 | 1.0972 | 2.6195 | 2.6195 |
O2 | 1.4168 | | 1.4058 | 2.3391 | 3.3701 | 2.0182 | 2.0842 | 2.0842 | 2.0791 | 2.0791 | C3 | 2.3313 | 1.4058 | | 1.4657 | 2.6246 | 3.2478 | 2.6230 | 2.6230 | 1.0991 | 1.0991 | C4 | 3.6166 | 2.3391 | 1.4657 | | 1.1592 | 4.3570 | 3.9691 | 3.9691 | 2.0910 | 2.0910 | N5 | 4.7170 | 3.3701 | 2.6246 | 1.1592 | | 5.3706 | 5.0860 | 5.0860 | 3.1437 | 3.1437 | H6 | 1.0876 | 2.0182 | 3.2478 | 4.3570 | 5.3706 | | 1.7813 | 1.7813 | 3.5977 | 3.5977 | H7 | 1.0972 | 2.0842 | 2.6230 | 3.9691 | 5.0860 | 1.7813 | | 1.7857 | 3.0123 | 2.4300 | H8 | 1.0972 | 2.0842 | 2.6230 | 3.9691 | 5.0860 | 1.7813 | 1.7857 | | 2.4300 | 3.0123 | H9 | 2.6195 | 2.0791 | 1.0991 | 2.0910 | 3.1437 | 3.5977 | 3.0123 | 2.4300 | | 1.7748 | H10 | 2.6195 | 2.0791 | 1.0991 | 2.0910 | 3.1437 | 3.5977 | 2.4300 | 3.0123 | 1.7748 | |
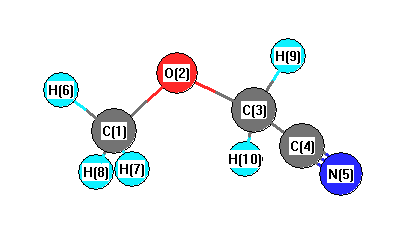 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.368 |
|
O2 |
C1 |
H6 |
106.651 |
| O2 |
C1 |
H7 |
111.367 |
|
O2 |
C1 |
H8 |
111.367 |
| O2 |
C3 |
C4 |
109.077 |
|
O2 |
C3 |
H9 |
111.610 |
| O2 |
C3 |
H10 |
111.610 |
|
C3 |
C4 |
N5 |
178.498 |
| C4 |
C3 |
H9 |
108.382 |
|
C4 |
C3 |
H10 |
108.382 |
| H6 |
C1 |
H7 |
109.236 |
|
H6 |
C1 |
H8 |
109.236 |
| H7 |
C1 |
H8 |
108.928 |
|
H9 |
C3 |
H10 |
107.676 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/6-311G**
| | hartrees |
|---|
| Energy at 0K | -247.120246 |
| Energy at 298.15K | |
| HF Energy | -246.829982 |
| Nuclear repulsion energy | 159.903001 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3170 |
18.71 |
|
|
|
| 2 |
A |
3134 |
3134 |
5.08 |
|
|
|
| 3 |
A |
3084 |
3084 |
38.70 |
|
|
|
| 4 |
A |
3024 |
3024 |
39.08 |
|
|
|
| 5 |
A |
3019 |
3019 |
35.82 |
|
|
|
| 6 |
A |
2287 |
2287 |
1.67 |
|
|
|
| 7 |
A |
1524 |
1524 |
10.25 |
|
|
|
| 8 |
A |
1506 |
1506 |
5.91 |
|
|
|
| 9 |
A |
1499 |
1499 |
5.18 |
|
|
|
| 10 |
A |
1487 |
1487 |
1.91 |
|
|
|
| 11 |
A |
1403 |
1403 |
16.10 |
|
|
|
| 12 |
A |
1325 |
1325 |
7.35 |
|
|
|
| 13 |
A |
1229 |
1229 |
33.16 |
|
|
|
| 14 |
A |
1193 |
1193 |
6.75 |
|
|
|
| 15 |
A |
1160 |
1160 |
114.69 |
|
|
|
| 16 |
A |
1039 |
1039 |
7.43 |
|
|
|
| 17 |
A |
948 |
948 |
33.62 |
|
|
|
| 18 |
A |
903 |
903 |
19.46 |
|
|
|
| 19 |
A |
601 |
601 |
1.75 |
|
|
|
| 20 |
A |
394 |
394 |
2.58 |
|
|
|
| 21 |
A |
361 |
361 |
1.06 |
|
|
|
| 22 |
A |
257 |
257 |
10.52 |
|
|
|
| 23 |
A |
178 |
178 |
3.45 |
|
|
|
| 24 |
A |
117 |
117 |
8.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17419.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17419.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.569 |
-0.768 |
0.138 |
| O2 |
1.077 |
0.433 |
-0.441 |
| C3 |
-0.063 |
0.928 |
0.210 |
| C4 |
-1.247 |
0.058 |
0.039 |
| N5 |
-2.161 |
-0.643 |
-0.098 |
| H6 |
2.472 |
-1.027 |
-0.411 |
| H7 |
0.844 |
-1.584 |
0.046 |
| H8 |
1.815 |
-0.622 |
1.196 |
| H9 |
-0.282 |
1.902 |
-0.226 |
| H10 |
0.108 |
1.061 |
1.287 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4212 | 2.3549 | 2.9358 | 3.7388 | 1.0875 | 1.0950 | 1.0969 | 3.2696 | 2.6080 |
O2 | 1.4212 | | 1.4028 | 2.4015 | 3.4284 | 2.0194 | 2.0880 | 2.0833 | 2.0128 | 2.0782 | C3 | 2.3549 | 1.4028 | | 1.4785 | 2.6384 | 3.2606 | 2.6758 | 2.6277 | 1.0900 | 1.0984 | C4 | 2.9358 | 2.4015 | 1.4785 | | 1.1601 | 3.8993 | 2.6585 | 3.3435 | 2.0978 | 2.0968 | N5 | 3.7388 | 3.4284 | 2.6384 | 1.1601 | | 4.6586 | 3.1518 | 4.1813 | 3.1660 | 3.1566 | H6 | 1.0875 | 2.0194 | 3.2606 | 3.8993 | 4.6586 | | 1.7802 | 1.7824 | 4.0245 | 3.5817 | H7 | 1.0950 | 2.0880 | 2.6758 | 2.6585 | 3.1518 | 1.7802 | | 1.7864 | 3.6739 | 3.0126 | H8 | 1.0969 | 2.0833 | 2.6277 | 3.3435 | 4.1813 | 1.7824 | 1.7864 | | 3.5773 | 2.3994 | H9 | 3.2696 | 2.0128 | 1.0900 | 2.0978 | 3.1660 | 4.0245 | 3.6739 | 3.5773 | | 1.7750 | H10 | 2.6080 | 2.0782 | 1.0984 | 2.0968 | 3.1566 | 3.5817 | 3.0126 | 2.3994 | 1.7750 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.997 |
|
O2 |
C1 |
H6 |
106.446 |
| O2 |
C1 |
H7 |
111.500 |
|
O2 |
C1 |
H8 |
110.993 |
| O2 |
C3 |
C4 |
112.887 |
|
O2 |
C3 |
H9 |
107.023 |
| O2 |
C3 |
H10 |
111.802 |
|
C3 |
C4 |
N5 |
178.821 |
| C4 |
C3 |
H9 |
108.582 |
|
C4 |
C3 |
H10 |
108.007 |
| H6 |
C1 |
H7 |
109.303 |
|
H6 |
C1 |
H8 |
109.366 |
| H7 |
C1 |
H8 |
109.174 |
|
H9 |
C3 |
H10 |
108.412 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
