Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -133.919607 |
| Energy at 298.15K | -133.924984 |
| HF Energy | -133.919607 |
| Nuclear repulsion energy | 71.018048 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3455 |
3455 |
1.57 |
|
|
|
| 2 |
A' |
3131 |
3131 |
18.47 |
|
|
|
| 3 |
A' |
3084 |
3084 |
23.70 |
|
|
|
| 4 |
A' |
3041 |
3041 |
5.59 |
|
|
|
| 5 |
A' |
1775 |
1775 |
74.54 |
|
|
|
| 6 |
A' |
1481 |
1481 |
29.27 |
|
|
|
| 7 |
A' |
1442 |
1442 |
22.27 |
|
|
|
| 8 |
A' |
1397 |
1397 |
9.26 |
|
|
|
| 9 |
A' |
1297 |
1297 |
54.08 |
|
|
|
| 10 |
A' |
1075 |
1075 |
18.05 |
|
|
|
| 11 |
A' |
917 |
917 |
3.92 |
|
|
|
| 12 |
A' |
499 |
499 |
7.24 |
|
|
|
| 13 |
A" |
3115 |
3115 |
7.32 |
|
|
|
| 14 |
A" |
1472 |
1472 |
11.87 |
|
|
|
| 15 |
A" |
1164 |
1164 |
51.65 |
|
|
|
| 16 |
A" |
1080 |
1080 |
9.12 |
|
|
|
| 17 |
A" |
691 |
691 |
7.10 |
|
|
|
| 18 |
A" |
123 |
123 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15120.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15120.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.015 |
-0.625 |
0.000 |
| C2 |
0.000 |
0.484 |
0.000 |
| N3 |
1.260 |
0.380 |
0.000 |
| H4 |
-0.531 |
-1.604 |
0.000 |
| H5 |
-1.660 |
-0.548 |
0.879 |
| H6 |
-1.660 |
-0.548 |
-0.879 |
| H7 |
-0.409 |
1.499 |
0.000 |
| H8 |
1.529 |
-0.608 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5029 | 2.4865 | 1.0915 | 1.0931 | 1.0931 | 2.2086 | 2.5436 |
C2 | 1.5029 | | 1.2640 | 2.1538 | 2.1427 | 2.1427 | 1.0944 | 1.8789 | N3 | 2.4865 | 1.2640 | | 2.6723 | 3.1868 | 3.1868 | 2.0091 | 1.0242 | H4 | 1.0915 | 2.1538 | 2.6723 | | 1.7780 | 1.7780 | 3.1048 | 2.2878 | H5 | 1.0931 | 2.1427 | 3.1868 | 1.7780 | | 1.7584 | 2.5545 | 3.3080 | H6 | 1.0931 | 2.1427 | 3.1868 | 1.7780 | 1.7584 | | 2.5545 | 3.3080 | H7 | 2.2086 | 1.0944 | 2.0091 | 3.1048 | 2.5545 | 2.5545 | | 2.8628 | H8 | 2.5436 | 1.8789 | 1.0242 | 2.2878 | 3.3080 | 3.3080 | 2.8628 | |
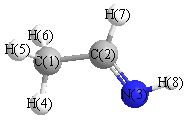 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.750 |
|
C1 |
C2 |
H7 |
115.596 |
| C2 |
C1 |
H4 |
111.239 |
|
C2 |
C1 |
H5 |
110.250 |
| C2 |
C1 |
H6 |
110.250 |
|
C2 |
N3 |
H8 |
109.948 |
| N3 |
C2 |
H7 |
116.654 |
|
H4 |
C1 |
H5 |
108.954 |
| H4 |
C1 |
H6 |
108.954 |
|
H5 |
C1 |
H6 |
107.087 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.407 |
|
|
|
| 2 |
C |
0.005 |
|
|
|
| 3 |
N |
-0.349 |
|
|
|
| 4 |
H |
0.135 |
|
|
|
| 5 |
H |
0.156 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
| 8 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.021 |
-1.440 |
0.000 |
2.482 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.307 |
-3.233 |
0.000 |
| y |
-3.233 |
-18.982 |
0.000 |
| z |
0.000 |
0.000 |
-19.809 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.912 |
-3.233 |
0.000 |
| y |
-3.233 |
1.576 |
0.000 |
| z |
0.000 |
0.000 |
0.336 |
|
| Polar |
|---|
| 3z2-r2 | 0.673 |
|---|
| x2-y2 | -2.325 |
|---|
| xy | -3.233 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.657 |
-0.041 |
0.000 |
| y |
-0.041 |
4.294 |
0.000 |
| z |
0.000 |
0.000 |
3.072 |
<r2> (average value of r
2) Å
2
| <r2> |
50.185 |
| (<r2>)1/2 |
7.084 |
