return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: CCSD/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at CCSD/6-311G**
 hartrees
Energy at 0K-118.144648
Energy at 298.15K 
HF Energy-117.663129
Nuclear repulsion energy75.625025
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3261 3261 17.26      
2 A 3157 3157 15.93      
3 A 3130 3130 33.90      
4 A 3125 3125 43.36      
5 A 3066 3066 20.40      
6 A 3052 3052 25.88      
7 A 2988 2988 35.29      
8 A 1527 1527 3.00      
9 A 1520 1520 5.42      
10 A 1499 1499 0.69      
11 A 1489 1489 1.10      
12 A 1439 1439 1.83      
13 A 1385 1385 2.12      
14 A 1295 1295 0.14      
15 A 1195 1195 0.42      
16 A 1106 1106 0.06      
17 A 1074 1074 1.07      
18 A 933 933 0.84      
19 A 900 900 0.07      
20 A 769 769 0.55      
21 A 465 465 39.71      
22 A 367 367 4.58      
23 A 259 259 0.11      
24 A 140 140 0.44      

Unscaled Zero Point Vibrational Energy (zpe) 19572.1 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19572.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD/6-311G**
ABC
1.08289 0.29832 0.25882

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD/6-311G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.303 -0.297 -0.040
C2 0.076 0.561 0.058
C3 -1.225 -0.248 -0.041
H4 -2.103 0.404 0.034
H5 -1.275 -0.783 -0.996
H6 -1.281 -0.989 0.765
H7 0.080 1.114 1.014
H8 0.099 1.327 -0.729
H9 2.261 0.131 -0.321
H10 1.297 -1.303 0.372

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.49982.52823.47742.79192.79392.14332.13511.08601.0872
C21.49981.53492.18422.17732.17821.10421.09842.25882.2500
C32.52821.53491.09581.09591.09662.16112.16923.51742.7652
H43.47742.18421.09581.77611.77512.49572.50584.38633.8194
H52.79192.17731.09591.77611.77313.07782.53173.71362.9596
H62.79392.17821.09661.77511.77312.51803.08243.87062.6274
H72.14331.10422.16112.49573.07782.51801.75542.73912.7807
H82.13511.09842.16922.50582.53173.08241.75542.50413.0922
H91.08602.25883.51744.38633.71363.87062.73912.50411.8608
H101.08722.25002.76523.81942.95962.62742.78073.09221.8608

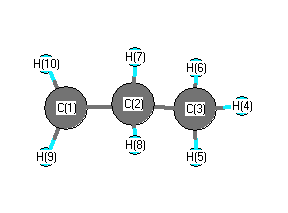
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.834 C1 C2 H7 109.859
C1 C2 H8 109.553 C2 C1 H9 120.909
C2 C1 H10 120.009 C2 C3 H4 111.160
C2 C3 H5 110.607 C2 C3 H6 110.633
C3 C2 H7 108.851 C3 C2 H8 109.814
H4 C3 H5 108.266 H4 C3 H6 108.124
H5 C3 H6 107.938 H7 C2 H8 105.680
H9 C1 H10 117.803
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability