Jump to
S2C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -77.058103 |
| Energy at 298.15K | -77.057851 |
| HF Energy | -76.784154 |
| Nuclear repulsion energy | 23.692008 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3153 |
3153 |
31.22 |
|
|
|
| 2 |
A1 |
1679 |
1679 |
76.22 |
|
|
|
| 3 |
A1 |
1243 |
1243 |
16.81 |
|
|
|
| 4 |
B1 |
768 |
768 |
84.22 |
|
|
|
| 5 |
B2 |
3247 |
3247 |
10.41 |
|
|
|
| 6 |
B2 |
386 |
386 |
7.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5238.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5238.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.484 |
| C2 |
0.000 |
0.000 |
0.827 |
| H3 |
0.000 |
0.942 |
-1.031 |
| H4 |
0.000 |
-0.942 |
-1.031 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3110 | 1.0889 | 1.0889 |
C2 | 1.3110 | | 2.0830 | 2.0830 | H3 | 1.0889 | 2.0830 | | 1.8831 | H4 | 1.0889 | 2.0830 | 1.8831 | |
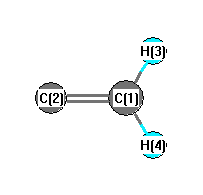 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.155 |
|
C2 |
C1 |
H4 |
120.155 |
| H3 |
C1 |
H4 |
119.689 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -76.990431 |
| Energy at 298.15K | -76.990508 |
| Nuclear repulsion energy | 23.435427 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3086 |
3086 |
4.72 |
|
|
|
| 2 |
A1 |
1565 |
1565 |
4.17 |
|
|
|
| 3 |
A1 |
1423 |
1423 |
2.92 |
|
|
|
| 4 |
B1 |
802 |
802 |
16.94 |
|
|
|
| 5 |
B2 |
3168 |
3168 |
7.17 |
|
|
|
| 6 |
B2 |
1009 |
1009 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5526.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5526.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.489 |
| C2 |
0.000 |
0.000 |
0.840 |
| H3 |
0.000 |
0.936 |
-1.052 |
| H4 |
0.000 |
-0.936 |
-1.052 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3294 | 1.0920 | 1.0920 |
C2 | 1.3294 | | 2.1111 | 2.1111 | H3 | 1.0920 | 2.1111 | | 1.8715 | H4 | 1.0920 | 2.1111 | 1.8715 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
