Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-311G**
| | hartrees |
|---|
| Energy at 0K | -100.273362 |
| Energy at 298.15K | -100.272427 |
| HF Energy | -100.273362 |
| Nuclear repulsion energy | 27.743685 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-311G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.058 |
| C2 |
0.000 |
0.000 |
-0.151 |
| N3 |
0.000 |
0.000 |
1.011 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9079 | 3.0697 |
C2 | 1.9079 | | 1.1617 | N3 | 3.0697 | 1.1617 | |
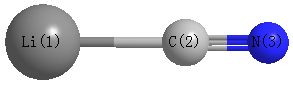 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.594 |
|
|
|
| 2 |
C |
-0.284 |
|
|
|
| 3 |
N |
-0.310 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-8.918 |
8.918 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.191 |
0.000 |
0.000 |
| y |
0.000 |
-14.191 |
0.000 |
| z |
0.000 |
0.000 |
1.124 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.657 |
0.000 |
0.000 |
| y |
0.000 |
-7.657 |
0.000 |
| z |
0.000 |
0.000 |
15.315 |
|
| Polar |
|---|
| 3z2-r2 | 30.629 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.304 |
0.000 |
0.000 |
| y |
0.000 |
2.304 |
0.000 |
| z |
0.000 |
0.000 |
4.215 |
<r2> (average value of r
2) Å
2
| <r2> |
25.680 |
| (<r2>)1/2 |
5.068 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-311G**
| | hartrees |
|---|
| Energy at 0K | -100.277638 |
| Energy at 298.15K | -100.276890 |
| HF Energy | -100.277638 |
| Nuclear repulsion energy | 29.417810 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.435 |
-0.581 |
0.000 |
| C2 |
-0.717 |
-0.367 |
0.000 |
| N3 |
0.000 |
0.564 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1625 | 1.8352 |
C2 | 2.1625 | | 1.1750 | N3 | 1.8352 | 1.1750 | |
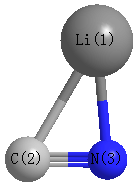 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
58.050 |
|
Li1 |
N3 |
C2 |
89.041 |
| C2 |
Li1 |
N3 |
32.908 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.491 |
|
|
|
| 2 |
C |
-0.116 |
|
|
|
| 3 |
N |
-0.376 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.440 |
-2.433 |
0.000 |
6.884 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-6.127 |
-5.854 |
0.000 |
| y |
-5.854 |
-15.095 |
0.000 |
| z |
0.000 |
0.000 |
-14.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.618 |
-5.854 |
0.000 |
| y |
-5.854 |
-4.833 |
0.000 |
| z |
0.000 |
0.000 |
-3.785 |
|
| Polar |
|---|
| 3z2-r2 | -7.570 |
|---|
| x2-y2 | 8.968 |
|---|
| xy | -5.854 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.361 |
0.350 |
0.000 |
| y |
0.350 |
3.276 |
0.000 |
| z |
0.000 |
0.000 |
2.595 |
<r2> (average value of r
2) Å
2
| <r2> |
20.618 |
| (<r2>)1/2 |
4.541 |
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-311G**
| | hartrees |
|---|
| Energy at 0K | -100.278108 |
| Energy at 298.15K | -100.277021 |
| HF Energy | -100.278108 |
| Nuclear repulsion energy | 28.468911 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
2166 |
2081 |
178.15 |
|
|
|
| 2 |
Σ |
728 |
700 |
155.92 |
|
|
|
| 3 |
Π |
110 |
106 |
29.73 |
|
|
|
| 3 |
Π |
110 |
106 |
29.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1557.8 cm
-1
Scaled (by 0.9604) Zero Point Vibrational Energy (zpe) 1496.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.873 |
| C2 |
0.000 |
0.000 |
-1.065 |
| N3 |
0.000 |
0.000 |
0.110 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9374 | 1.7628 |
C2 | 2.9374 | | 1.1746 | N3 | 1.7628 | 1.1746 | |
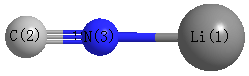 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.595 |
|
|
|
| 2 |
C |
-0.134 |
|
|
|
| 3 |
N |
-0.461 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
8.727 |
8.727 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.171 |
0.000 |
0.000 |
| y |
0.000 |
-14.171 |
0.000 |
| z |
0.000 |
0.000 |
-2.464 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.854 |
0.000 |
0.000 |
| y |
0.000 |
-5.854 |
0.000 |
| z |
0.000 |
0.000 |
11.707 |
|
| Polar |
|---|
| 3z2-r2 | 23.415 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.336 |
0.000 |
0.000 |
| y |
0.000 |
2.336 |
0.000 |
| z |
0.000 |
0.000 |
4.406 |
<r2> (average value of r
2) Å
2
| <r2> |
23.820 |
| (<r2>)1/2 |
4.881 |
