Jump to
S1C2
S2C1
S2C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -131.396212 |
| Energy at 298.15K | |
| HF Energy | -131.396212 |
| Nuclear repulsion energy | 47.450625 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.004 |
-1.233 |
0.000 |
| C2 |
0.000 |
0.099 |
0.000 |
| N3 |
-0.092 |
1.273 |
0.000 |
| H4 |
0.617 |
-2.112 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3323 | 2.5081 | 1.0717 |
C2 | 1.3323 | | 1.1776 | 2.2959 | N3 | 2.5081 | 1.1776 | | 3.4588 | H4 | 1.0717 | 2.2959 | 3.4588 | |
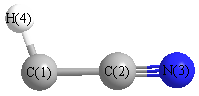 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
175.711 |
|
C2 |
C1 |
H4 |
145.297 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -131.394758 |
| Energy at 298.15K | |
| HF Energy | -131.394758 |
| Nuclear repulsion energy | 47.582762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3445 |
3445 |
75.24 |
|
|
|
| 2 |
Σ |
1726 |
1726 |
40.36 |
|
|
|
| 3 |
Σ |
1243 |
1243 |
16.17 |
|
|
|
| 4 |
Π |
459 |
459 |
0.46 |
|
|
|
| 4 |
Π |
459 |
459 |
0.46 |
|
|
|
| 5 |
Π |
357i |
357i |
44.56 |
|
|
|
| 5 |
Π |
357i |
357i |
44.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3309.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3309.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.209 |
| C2 |
0.000 |
0.000 |
0.091 |
| N3 |
0.000 |
0.000 |
1.283 |
| H4 |
0.000 |
0.000 |
-2.274 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2998 | 2.4920 | 1.0650 |
C2 | 1.2998 | | 1.1922 | 2.3648 | N3 | 2.4920 | 1.1922 | | 3.5570 | H4 | 1.0650 | 2.3648 | 3.5570 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.009 |
|
|
|
| 2 |
C |
0.043 |
|
|
|
| 3 |
N |
-0.252 |
|
|
|
| 4 |
H |
0.199 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.448 |
3.448 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.344 |
0.000 |
0.000 |
| y |
0.000 |
-17.344 |
0.000 |
| z |
0.000 |
0.000 |
-15.450 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.947 |
0.000 |
0.000 |
| y |
0.000 |
-0.947 |
0.000 |
| z |
0.000 |
0.000 |
1.895 |
|
| Polar |
|---|
| 3z2-r2 | 3.789 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.899 |
0.000 |
0.000 |
| y |
0.000 |
1.899 |
0.000 |
| z |
0.000 |
0.000 |
6.374 |
<r2> (average value of r
2) Å
2
| <r2> |
35.952 |
| (<r2>)1/2 |
5.996 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -131.370504 |
| Energy at 298.15K | -131.369981 |
| HF Energy | -131.370504 |
| Nuclear repulsion energy | 47.125956 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3100 |
3100 |
5.12 |
|
|
|
| 2 |
A' |
2183 |
2183 |
26.20 |
|
|
|
| 3 |
A' |
1071 |
1071 |
63.86 |
|
|
|
| 4 |
A' |
980 |
980 |
14.77 |
|
|
|
| 5 |
A' |
454 |
454 |
23.00 |
|
|
|
| 6 |
A" |
334 |
334 |
9.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4061.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4061.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
-1.290 |
0.000 |
| C2 |
0.000 |
0.099 |
0.000 |
| N3 |
-0.110 |
1.260 |
0.000 |
| H4 |
0.993 |
-1.674 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3893 | 2.5503 | 1.0990 |
C2 | 1.3893 | | 1.1657 | 2.0322 | N3 | 2.5503 | 1.1657 | | 3.1343 | H4 | 1.0990 | 2.0322 | 3.1343 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.026 |
|
|
|
| 2 |
C |
0.082 |
|
|
|
| 3 |
N |
-0.231 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.687 |
-1.604 |
0.000 |
2.328 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.392 |
-2.815 |
0.000 |
| y |
-2.815 |
-20.803 |
0.000 |
| z |
0.000 |
0.000 |
-15.937 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.978 |
-2.815 |
0.000 |
| y |
-2.815 |
-4.138 |
0.000 |
| z |
0.000 |
0.000 |
3.161 |
|
| Polar |
|---|
| 3z2-r2 | 6.321 |
|---|
| x2-y2 | 3.410 |
|---|
| xy | -2.815 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.488 |
-0.347 |
0.000 |
| y |
-0.347 |
6.168 |
0.000 |
| z |
0.000 |
0.000 |
2.397 |
<r2> (average value of r
2) Å
2
| <r2> |
36.297 |
| (<r2>)1/2 |
6.025 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -131.370504 |
| Energy at 298.15K | -131.369981 |
| HF Energy | -131.370504 |
| Nuclear repulsion energy | 47.125956 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
