return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (3-Methylisoxazole)

using model chemistry: MP2/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-311G**
 hartrees
Energy at 0K-284.629868
Energy at 298.15K-284.636316
HF Energy-283.694358
Nuclear repulsion energy223.202930
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3319 3154 0.05      
2 A 3297 3133 0.13      
3 A 3190 3031 5.20      
4 A 3166 3008 11.44      
5 A 3085 2931 15.72      
6 A 1593 1514 19.45      
7 A 1518 1443 13.29      
8 A 1502 1427 13.70      
9 A 1502 1427 8.28      
10 A 1438 1366 15.39      
11 A 1418 1348 0.81      
12 A 1298 1233 0.20      
13 A 1183 1124 17.92      
14 A 1085 1031 11.07      
15 A 1073 1020 0.66      
16 A 1034 982 2.05      
17 A 996 947 12.98      
18 A 984 935 4.50      
19 A 920 875 9.09      
20 A 851 808 0.68      
21 A 779 740 49.37      
22 A 666 633 4.58      
23 A 642 610 0.01      
24 A 619 588 3.20      
25 A 333 317 5.78      
26 A 264 251 5.32      
27 A 120 114 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 18938.0 cm-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 17994.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-311G**
ABC
0.30533 0.11741 0.08617

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-311G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.133 0.020 0.000
H2 -2.527 0.526 -0.886
H3 -2.483 -1.014 0.000
H4 -2.527 0.526 0.886
C5 1.488 0.573 0.000
O6 1.371 -0.769 -0.000
N7 0.035 -1.106 -0.000
C8 -0.637 0.039 -0.000
C9 0.251 1.150 -0.000
H10 0.010 2.202 0.000
H11 2.498 0.954 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 C9 H10 H11
C11.09351.09161.09353.66303.59142.44311.49582.63833.05764.7244
H21.09351.77681.77154.11194.20163.16432.14312.98193.16655.1206
H31.09161.77681.77684.27663.86192.52042.12503.48704.06855.3561
H41.09351.77151.77684.11184.20153.16422.14312.98193.16645.1205
C53.66304.11194.27664.11181.34672.21972.19141.36512.19991.0797
O63.59144.20163.86194.20151.34671.37722.16432.22173.26752.0592
N72.44313.16432.52043.16422.21971.37721.32742.26603.30743.2107
C81.49582.14312.12502.14312.19142.16431.32741.42282.25753.2664
C92.63832.98193.48702.98191.36512.22172.26601.42281.07902.2555
H103.05763.16654.06853.16642.19993.26753.30742.25751.07902.7837
H114.72445.12065.35615.12051.07972.05923.21073.26642.25552.7837

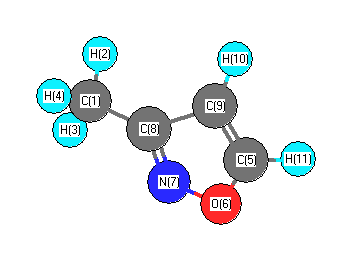
picture of 3-Methylisoxazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 N7 119.732 C1 C8 C9 129.353
H2 C1 H3 108.812 H2 C1 H4 108.200
H2 C1 C8 110.768 H3 C1 H4 108.809
H3 C1 C8 109.437 H4 C1 C8 110.764
C5 O6 N7 109.155 C5 C9 C8 103.617
C5 C9 H10 127.963 O6 C5 C9 110.023
O6 C5 H11 115.698 O6 N7 C8 106.290
N7 C8 C9 110.915 C8 C9 H10 128.420
C9 C5 H11 134.279
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability