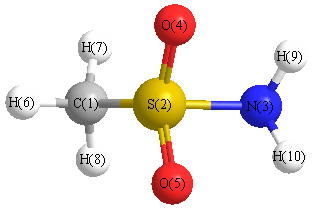
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3556 |
3412 |
45.47 |
|
|
|
| 2 |
A' |
3196 |
3066 |
0.03 |
|
|
|
| 3 |
A' |
3088 |
2962 |
0.05 |
|
|
|
| 4 |
A' |
1599 |
1534 |
49.41 |
|
|
|
| 5 |
A' |
1449 |
1390 |
8.33 |
|
|
|
| 6 |
A' |
1359 |
1304 |
22.64 |
|
|
|
| 7 |
A' |
1160 |
1113 |
195.23 |
|
|
|
| 8 |
A' |
995 |
954 |
32.46 |
|
|
|
| 9 |
A' |
889 |
852 |
126.63 |
|
|
|
| 10 |
A' |
738 |
708 |
12.38 |
|
|
|
| 11 |
A' |
660 |
633 |
236.37 |
|
|
|
| 12 |
A' |
502 |
482 |
51.28 |
|
|
|
| 13 |
A' |
476 |
457 |
10.39 |
|
|
|
| 14 |
A' |
298 |
286 |
4.96 |
|
|
|
| 15 |
A" |
3666 |
3517 |
53.96 |
|
|
|
| 16 |
A" |
3205 |
3075 |
0.14 |
|
|
|
| 17 |
A" |
1448 |
1389 |
1.02 |
|
|
|
| 18 |
A" |
1380 |
1324 |
263.89 |
|
|
|
| 19 |
A" |
1094 |
1049 |
1.27 |
|
|
|
| 20 |
A" |
975 |
936 |
1.49 |
|
|
|
| 21 |
A" |
401 |
385 |
0.04 |
|
|
|
| 22 |
A" |
331 |
318 |
1.93 |
|
|
|
| 23 |
A" |
228 |
219 |
0.14 |
|
|
|
| 24 |
A" |
165 |
158 |
40.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16428.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15761.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.483 |
|
|
|
| 2 |
S |
0.994 |
|
|
|
| 3 |
N |
-0.602 |
|
|
|
| 4 |
O |
-0.505 |
|
|
|
| 5 |
O |
-0.505 |
|
|
|
| 6 |
H |
0.191 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.178 |
|
|
|
| 9 |
H |
0.276 |
|
|
|
| 10 |
H |
0.276 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.759 |
3.237 |
0.000 |
3.684 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.408 |
5.761 |
0.000 |
| y |
5.761 |
-36.675 |
0.000 |
| z |
0.000 |
0.000 |
-41.260 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.560 |
5.761 |
0.000 |
| y |
5.761 |
0.158 |
0.000 |
| z |
0.000 |
0.000 |
-6.718 |
|
| Polar |
|---|
| 3z2-r2 | -13.437 |
|---|
| x2-y2 | 4.268 |
|---|
| xy | 5.761 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.248 |
0.297 |
0.000 |
| y |
0.297 |
5.910 |
0.000 |
| z |
0.000 |
0.000 |
6.071 |
<r2> (average value of r
2) Å
2
| <r2> |
120.352 |
| (<r2>)1/2 |
10.970 |