return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H11NO (Diethylhydroxylamine)

using model chemistry: MP2/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-311G**
 hartrees
Energy at 0K-288.192406
Energy at 298.15K-288.205988
HF Energy-287.193965
Nuclear repulsion energy264.548632
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3661 3478 0.91      
2 A 3197 3038 30.96      
3 A 3178 3020 28.90      
4 A 3130 2974 30.77      
5 A 3089 2935 2.66      
6 A 2961 2813 141.16      
7 A 1534 1458 12.01      
8 A 1512 1436 1.23      
9 A 1505 1430 20.20      
10 A 1451 1379 31.80      
11 A 1434 1362 36.38      
12 A 1404 1334 0.18      
13 A 1321 1255 2.81      
14 A 1209 1149 0.65      
15 A 1188 1129 11.72      
16 A 1112 1056 2.84      
17 A 971 922 10.98      
18 A 844 802 1.17      
19 A 800 760 18.04      
20 A 482 458 4.06      
21 A 435 413 0.09      
22 A 224 213 0.11      
23 A 210 200 1.96      
24 A 115 109 0.41      
25 A 3197 3037 0.15      
26 A 3178 3020 12.36      
27 A 3119 2964 0.30      
28 A 3088 2934 39.27      
29 A 2951 2804 3.26      
30 A 1531 1455 0.53      
31 A 1506 1431 0.80      
32 A 1498 1424 1.69      
33 A 1427 1356 29.61      
34 A 1397 1327 15.54      
35 A 1316 1250 0.81      
36 A 1227 1166 1.76      
37 A 1118 1063 2.80      
38 A 1095 1040 16.59      
39 A 975 926 0.17      
40 A 823 782 0.02      
41 A 570 542 39.99      
42 A 446 424 88.40      
43 A 353 335 4.64      
44 A 260 247 0.00      
45 A 133 126 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 34085.2 cm-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 32387.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-311G**
ABC
0.25936 0.06780 0.05947

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-311G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.183 -0.167 0.000
O2 1.076 -0.828 0.000
H3 1.735 -0.111 0.000
C4 -0.261 -0.211 2.462
C5 -0.261 -0.211 -2.462
C6 -0.261 0.650 1.207
C7 -0.261 0.650 -1.207
H8 0.664 -0.784 2.533
H9 0.664 -0.784 -2.533
H10 0.579 1.373 -1.234
H11 0.579 1.373 1.234
H12 -1.190 1.225 -1.139
H13 -1.190 1.225 1.139
H14 -1.098 -0.912 2.426
H15 -1.098 -0.912 -2.426
H16 -0.358 0.416 3.353
H17 -0.358 0.416 -3.353

Atom - Atom Distances (Å)
  N1 O2 H3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
N11.42301.91882.46342.46341.45961.45962.74132.74132.11542.11542.06102.06102.69762.69763.40773.4077
O21.42300.97332.86872.86872.33042.33042.56642.56642.57212.57213.26353.26353.25903.25903.85343.8534
H31.91880.97333.17043.17042.45322.45322.83072.83072.24962.24963.41133.41133.81453.81453.98753.9875
C42.46342.86873.17044.92361.52153.76861.09075.11184.10732.17323.98632.16211.09255.00841.09385.8493
C52.46342.86873.17044.92363.76861.52155.11181.09072.17324.10732.16213.98635.00841.09255.84931.0938
C61.45962.33042.45321.52153.76862.41442.16104.11102.68061.10872.58831.09502.15064.04232.16074.5673
C71.45962.33042.45323.76861.52152.41444.11102.16101.10872.68061.09502.58834.04232.15064.56732.1607
H82.74132.56642.83071.09075.11182.16104.11105.06564.34132.51984.57803.06891.77045.26431.77726.0933
H92.74132.56642.83075.11181.09074.11102.16105.06562.51984.34133.06894.57805.26431.77046.09331.7772
H102.11542.57212.24964.10732.17322.68061.10874.34132.51982.46711.77802.96344.62883.07504.77822.5074
H112.11542.57212.24962.17324.10731.10872.68062.51984.34132.46712.96341.77803.07504.62882.50744.7782
H122.06103.26353.41133.98632.16212.58831.09504.57803.06891.77802.96342.27814.15732.49584.63952.4997
H132.06103.26353.41132.16213.98631.09502.58833.06894.57802.96341.77802.27812.49584.15732.49974.6395
H142.69763.25903.81451.09255.00842.15064.04231.77045.26434.62883.07504.15732.49584.85221.78015.9756
H152.69763.25903.81455.00841.09254.04232.15065.26431.77043.07504.62882.49584.15734.85225.97561.7801
H163.40773.85343.98751.09385.84932.16074.56731.77726.09334.77822.50744.63952.49971.78015.97566.7060
H173.40773.85343.98755.84931.09384.56732.16076.09331.77722.50744.77822.49974.63955.97561.78016.7060

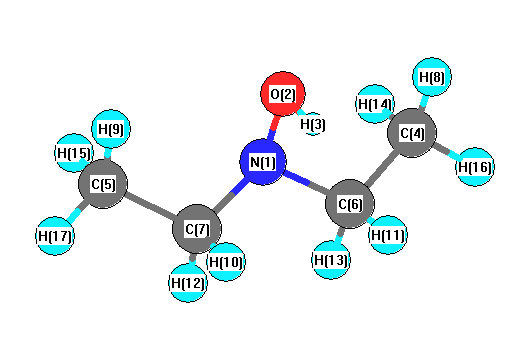
picture of Diethylhydroxylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 H3 104.847 N1 C6 C4 111.432
N1 C6 H11 110.157 N1 C6 H13 106.692
N1 C7 C5 111.432 N1 C7 H10 110.157
N1 C7 H12 106.692 O2 N1 C6 107.880
O2 N1 C7 107.880 C4 C6 H11 110.455
C4 C6 H13 110.386 C5 C7 H10 110.455
C5 C7 H12 110.386 C6 N1 C7 111.598
C6 C4 H8 110.552 C6 C4 H14 109.630
C6 C4 H16 110.349 C7 C5 H9 110.552
C7 C5 H15 109.630 C7 C5 H17 110.349
H8 C4 H14 108.367 H8 C4 H16 108.885
H9 C5 H15 108.367 H9 C5 H17 108.885
H10 C7 H12 107.580 H11 C6 H13 107.580
H14 C4 H16 109.014 H15 C5 H17 109.014
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability