Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -376.809588 |
| Energy at 298.15K | |
| HF Energy | -375.892349 |
| Nuclear repulsion energy | 191.074454 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3170 |
34.23 |
|
|
|
| 2 |
A |
3146 |
3146 |
21.95 |
|
|
|
| 3 |
A |
3103 |
3103 |
17.85 |
|
|
|
| 4 |
A |
1528 |
1528 |
1.97 |
|
|
|
| 5 |
A |
1508 |
1508 |
19.21 |
|
|
|
| 6 |
A |
1455 |
1455 |
28.62 |
|
|
|
| 7 |
A |
1382 |
1382 |
29.24 |
|
|
|
| 8 |
A |
1299 |
1299 |
13.30 |
|
|
|
| 9 |
A |
1205 |
1205 |
103.86 |
|
|
|
| 10 |
A |
1174 |
1174 |
60.13 |
|
|
|
| 11 |
A |
1160 |
1160 |
145.31 |
|
|
|
| 12 |
A |
1132 |
1132 |
48.51 |
|
|
|
| 13 |
A |
937 |
937 |
35.89 |
|
|
|
| 14 |
A |
592 |
592 |
5.21 |
|
|
|
| 15 |
A |
488 |
488 |
20.94 |
|
|
|
| 16 |
A |
438 |
438 |
5.78 |
|
|
|
| 17 |
A |
250 |
250 |
8.80 |
|
|
|
| 18 |
A |
126 |
126 |
8.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12046.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12046.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.772 |
-0.594 |
-0.282 |
| C2 |
0.469 |
0.021 |
0.328 |
| F3 |
-1.875 |
0.108 |
0.150 |
| F4 |
1.533 |
-0.755 |
-0.008 |
| F5 |
0.672 |
1.259 |
-0.182 |
| H6 |
-0.704 |
-0.533 |
-1.372 |
| H7 |
-0.858 |
-1.636 |
0.038 |
| H8 |
0.415 |
0.100 |
1.416 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5137 | 1.3769 | 2.3269 | 2.3513 | 1.0936 | 1.0935 | 2.1851 |
C2 | 1.5137 | | 2.3525 | 1.3590 | 1.3538 | 2.1382 | 2.1426 | 1.0926 | F3 | 1.3769 | 2.3525 | | 3.5191 | 2.8146 | 2.0247 | 2.0219 | 2.6164 | F4 | 2.3269 | 1.3590 | 3.5191 | | 2.1970 | 2.6291 | 2.5485 | 2.0027 | F5 | 2.3513 | 1.3538 | 2.8146 | 2.1970 | | 2.5535 | 3.2816 | 1.9904 | H6 | 1.0936 | 2.1382 | 2.0247 | 2.6291 | 2.5535 | | 1.7969 | 3.0701 | H7 | 1.0935 | 2.1426 | 2.0219 | 2.5485 | 3.2816 | 1.7969 | | 2.5559 | H8 | 2.1851 | 1.0926 | 2.6164 | 2.0027 | 1.9904 | 3.0701 | 2.5559 | |
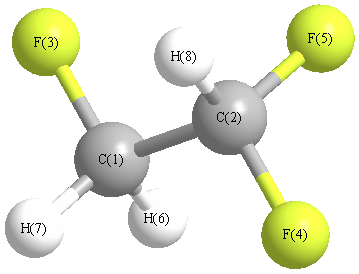 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.074 |
|
C1 |
C2 |
F5 |
110.048 |
| C1 |
C2 |
H8 |
112.952 |
|
C2 |
C1 |
F3 |
108.852 |
| C2 |
C1 |
H6 |
109.125 |
|
C2 |
C1 |
H7 |
109.480 |
| F3 |
C1 |
H6 |
109.542 |
|
F3 |
C1 |
H7 |
109.323 |
| F4 |
C2 |
F5 |
108.162 |
|
F4 |
C2 |
H8 |
109.068 |
| F5 |
C2 |
H8 |
108.428 |
|
H6 |
C1 |
H7 |
110.493 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -376.806983 |
| Energy at 298.15K | |
| HF Energy | -375.889452 |
| Nuclear repulsion energy | 193.881436 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3120 |
3120 |
57.54 |
|
|
|
| 2 |
A' |
3092 |
3092 |
10.69 |
|
|
|
| 3 |
A' |
1526 |
1526 |
7.56 |
|
|
|
| 4 |
A' |
1485 |
1485 |
30.51 |
|
|
|
| 5 |
A' |
1449 |
1449 |
27.16 |
|
|
|
| 6 |
A' |
1224 |
1224 |
115.75 |
|
|
|
| 7 |
A' |
1142 |
1142 |
18.32 |
|
|
|
| 8 |
A' |
903 |
903 |
39.21 |
|
|
|
| 9 |
A' |
776 |
776 |
54.42 |
|
|
|
| 10 |
A' |
529 |
529 |
14.06 |
|
|
|
| 11 |
A' |
241 |
241 |
2.29 |
|
|
|
| 12 |
A" |
3152 |
3152 |
22.98 |
|
|
|
| 13 |
A" |
1450 |
1450 |
37.74 |
|
|
|
| 14 |
A" |
1317 |
1317 |
19.53 |
|
|
|
| 15 |
A" |
1192 |
1192 |
103.98 |
|
|
|
| 16 |
A" |
988 |
988 |
58.85 |
|
|
|
| 17 |
A" |
377 |
377 |
0.06 |
|
|
|
| 18 |
A" |
121 |
121 |
2.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12041.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12041.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.740 |
-0.832 |
0.000 |
| C2 |
0.355 |
0.631 |
0.000 |
| F3 |
-0.388 |
-1.614 |
0.000 |
| F4 |
-0.388 |
0.919 |
1.096 |
| F5 |
-0.388 |
0.919 |
-1.096 |
| H6 |
1.328 |
-1.047 |
-0.898 |
| H7 |
1.328 |
-1.047 |
0.898 |
| H8 |
1.240 |
1.277 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5128 | 1.3728 | 2.3535 | 2.3535 | 1.0946 | 1.0946 | 2.1669 |
C2 | 1.5128 | | 2.3654 | 1.3550 | 1.3550 | 2.1378 | 2.1378 | 1.0953 | F3 | 1.3728 | 2.3654 | | 2.7606 | 2.7606 | 2.0177 | 2.0177 | 3.3179 | F4 | 2.3535 | 1.3550 | 2.7606 | | 2.1924 | 3.2842 | 2.6171 | 1.9946 | F5 | 2.3535 | 1.3550 | 2.7606 | 2.1924 | | 2.6171 | 3.2842 | 1.9946 | H6 | 1.0946 | 2.1378 | 2.0177 | 3.2842 | 2.6171 | | 1.7958 | 2.4929 | H7 | 1.0946 | 2.1378 | 2.0177 | 2.6171 | 3.2842 | 1.7958 | | 2.4929 | H8 | 2.1669 | 1.0953 | 3.3179 | 1.9946 | 1.9946 | 2.4929 | 2.4929 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.180 |
|
C1 |
C2 |
F5 |
110.180 |
| C1 |
C2 |
H8 |
111.361 |
|
C2 |
C1 |
F3 |
110.018 |
| C2 |
C1 |
H6 |
109.089 |
|
C2 |
C1 |
H7 |
109.089 |
| F3 |
C1 |
H6 |
109.204 |
|
F3 |
C1 |
H7 |
109.204 |
| F4 |
C2 |
F5 |
107.995 |
|
F4 |
C2 |
H8 |
108.518 |
| F5 |
C2 |
H8 |
108.518 |
|
H6 |
C1 |
H7 |
110.227 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
