Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -188.011738 |
| Energy at 298.15K | -188.015916 |
| HF Energy | -188.011738 |
| Nuclear repulsion energy | 102.137675 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3670 |
3670 |
28.93 |
|
|
|
| 2 |
A |
3579 |
3579 |
3.96 |
|
|
|
| 3 |
A |
2419 |
2419 |
0.18 |
|
|
|
| 4 |
A |
1654 |
1654 |
11.64 |
|
|
|
| 5 |
A |
1204 |
1204 |
0.10 |
|
|
|
| 6 |
A |
836 |
836 |
2.25 |
|
|
|
| 7 |
A |
559 |
559 |
143.88 |
|
|
|
| 8 |
A |
452 |
452 |
16.12 |
|
|
|
| 9 |
A |
423 |
423 |
22.70 |
|
|
|
| 10 |
A |
225 |
225 |
22.22 |
|
|
|
| 11 |
B |
3670 |
3670 |
28.23 |
|
|
|
| 12 |
B |
3581 |
3581 |
18.58 |
|
|
|
| 13 |
B |
1656 |
1656 |
43.19 |
|
|
|
| 14 |
B |
1385 |
1385 |
121.75 |
|
|
|
| 15 |
B |
1204 |
1204 |
0.13 |
|
|
|
| 16 |
B |
612 |
612 |
333.22 |
|
|
|
| 17 |
B |
457 |
457 |
31.53 |
|
|
|
| 18 |
B |
225 |
225 |
25.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13904.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13904.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.002 |
0.602 |
0.048 |
| C2 |
-0.002 |
-0.602 |
0.048 |
| N3 |
-0.002 |
1.952 |
-0.080 |
| N4 |
0.002 |
-1.952 |
-0.080 |
| H5 |
-0.340 |
2.458 |
0.726 |
| H6 |
0.849 |
2.350 |
-0.453 |
| H7 |
0.340 |
-2.458 |
0.726 |
| H8 |
-0.849 |
-2.350 |
-0.453 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2041 | 1.3561 | 2.5574 | 2.0053 | 2.0053 | 3.1524 | 3.1124 |
C2 | 1.2041 | | 2.5574 | 1.3561 | 3.1524 | 3.1124 | 2.0053 | 2.0053 | N3 | 1.3561 | 2.5574 | | 3.9042 | 1.0102 | 1.0104 | 4.4962 | 4.3999 | N4 | 2.5574 | 1.3561 | 3.9042 | | 4.4962 | 4.3999 | 1.0102 | 1.0104 | H5 | 2.0053 | 3.1524 | 1.0102 | 4.4962 | | 1.6777 | 4.9627 | 4.9760 | H6 | 2.0053 | 3.1124 | 1.0104 | 4.3999 | 1.6777 | | 4.9760 | 4.9964 | H7 | 3.1524 | 2.0053 | 4.4962 | 1.0102 | 4.9627 | 4.9760 | | 1.6777 | H8 | 3.1124 | 2.0053 | 4.3999 | 1.0104 | 4.9760 | 4.9964 | 1.6777 | |
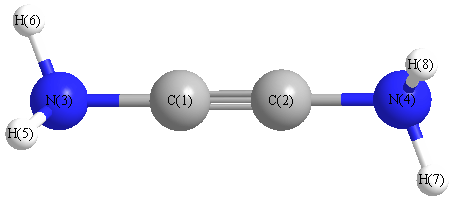 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
174.556 |
|
C1 |
N3 |
H5 |
115.092 |
| C1 |
N3 |
H6 |
115.071 |
|
C2 |
C1 |
N3 |
174.556 |
| C2 |
N4 |
H7 |
115.092 |
|
C2 |
N4 |
H8 |
115.071 |
| H5 |
N3 |
H6 |
112.266 |
|
H7 |
N4 |
H8 |
112.266 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.133 |
|
|
|
| 2 |
C |
0.133 |
|
|
|
| 3 |
N |
-0.615 |
|
|
|
| 4 |
N |
-0.615 |
|
|
|
| 5 |
H |
0.240 |
|
|
|
| 6 |
H |
0.242 |
|
|
|
| 7 |
H |
0.240 |
|
|
|
| 8 |
H |
0.242 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.515 |
1.515 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.594 |
4.213 |
0.000 |
| y |
4.213 |
-13.740 |
0.000 |
| z |
0.000 |
0.000 |
-23.748 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.850 |
4.213 |
0.000 |
| y |
4.213 |
9.931 |
0.000 |
| z |
0.000 |
0.000 |
-5.081 |
|
| Polar |
|---|
| 3z2-r2 | -10.162 |
|---|
| x2-y2 | -9.854 |
|---|
| xy | 4.213 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.126 |
0.230 |
0.000 |
| y |
0.230 |
9.000 |
0.000 |
| z |
0.000 |
0.000 |
3.117 |
<r2> (average value of r
2) Å
2
| <r2> |
96.780 |
| (<r2>)1/2 |
9.838 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -188.009421 |
| Energy at 298.15K | |
| HF Energy | -188.009421 |
| Nuclear repulsion energy | 102.491129 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3639 |
3639 |
0.00 |
|
|
|
| 2 |
A1 |
2392 |
2392 |
0.00 |
|
|
|
| 3 |
A1 |
1637 |
1637 |
0.00 |
|
|
|
| 4 |
A1 |
860 |
860 |
0.00 |
|
|
|
| 5 |
B1 |
524 |
524 |
0.00 |
|
|
|
| 6 |
B2 |
3644 |
3644 |
41.62 |
|
|
|
| 7 |
B2 |
1649 |
1649 |
77.89 |
|
|
|
| 8 |
B2 |
1444 |
1444 |
152.02 |
|
|
|
| 9 |
E |
3751 |
3751 |
46.46 |
|
|
|
| 9 |
E |
3751 |
3751 |
46.46 |
|
|
|
| 10 |
E |
1146 |
1146 |
2.02 |
|
|
|
| 10 |
E |
1146 |
1146 |
2.02 |
|
|
|
| 11 |
E |
473 |
473 |
0.17 |
|
|
|
| 11 |
E |
473 |
473 |
0.17 |
|
|
|
| 12 |
E |
207 |
207 |
32.20 |
|
|
|
| 12 |
E |
207 |
207 |
32.20 |
|
|
|
| 13 |
E |
401i |
401i |
263.65 |
|
|
|
| 13 |
E |
401i |
401i |
263.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13070.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13070.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.604 |
| C2 |
0.000 |
0.000 |
-0.604 |
| N3 |
0.000 |
0.000 |
1.940 |
| N4 |
0.000 |
0.000 |
-1.940 |
| H5 |
0.000 |
0.860 |
2.459 |
| H6 |
0.000 |
-0.860 |
2.459 |
| H7 |
0.860 |
0.000 |
-2.459 |
| H8 |
-0.860 |
0.000 |
-2.459 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2085 | 1.3358 | 2.5443 | 2.0444 | 2.0444 | 3.1816 | 3.1816 |
C2 | 1.2085 | | 2.5443 | 1.3358 | 3.1816 | 3.1816 | 2.0444 | 2.0444 | N3 | 1.3358 | 2.5443 | | 3.8800 | 1.0043 | 1.0043 | 4.4823 | 4.4823 | N4 | 2.5443 | 1.3358 | 3.8800 | | 4.4823 | 4.4823 | 1.0043 | 1.0043 | H5 | 2.0444 | 3.1816 | 1.0043 | 4.4823 | | 1.7197 | 5.0661 | 5.0661 | H6 | 2.0444 | 3.1816 | 1.0043 | 4.4823 | 1.7197 | | 5.0661 | 5.0661 | H7 | 3.1816 | 2.0444 | 4.4823 | 1.0043 | 5.0661 | 5.0661 | | 1.7197 | H8 | 3.1816 | 2.0444 | 4.4823 | 1.0043 | 5.0661 | 5.0661 | 1.7197 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.113 |
| C1 |
N3 |
H6 |
121.113 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.113 |
|
C2 |
N4 |
H8 |
121.113 |
| H5 |
N3 |
H6 |
117.774 |
|
H7 |
N4 |
H8 |
117.774 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.162 |
|
|
|
| 2 |
C |
0.162 |
|
|
|
| 3 |
N |
-0.658 |
|
|
|
| 4 |
N |
-0.658 |
|
|
|
| 5 |
H |
0.248 |
|
|
|
| 6 |
H |
0.248 |
|
|
|
| 7 |
H |
0.248 |
|
|
|
| 8 |
H |
0.248 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.827 |
0.000 |
0.000 |
| y |
0.000 |
-23.827 |
0.000 |
| z |
0.000 |
0.000 |
-10.783 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.522 |
0.000 |
0.000 |
| y |
0.000 |
-6.522 |
0.000 |
| z |
0.000 |
0.000 |
13.044 |
|
| Polar |
|---|
| 3z2-r2 | 26.087 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.992 |
0.000 |
0.000 |
| y |
0.000 |
2.992 |
0.000 |
| z |
0.000 |
0.000 |
9.249 |
<r2> (average value of r
2) Å
2
| <r2> |
96.382 |
| (<r2>)1/2 |
9.817 |
