Jump to
S1C2
Energy calculated at TPSSh/6-311G**
| | hartrees |
|---|
| Energy at 0K | -188.093912 |
| Energy at 298.15K | -188.097901 |
| HF Energy | -188.093912 |
| Nuclear repulsion energy | 101.945280 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3580 |
3580 |
11.70 |
|
|
|
| 2 |
A |
3499 |
3499 |
1.83 |
|
|
|
| 3 |
A |
2355 |
2355 |
0.36 |
|
|
|
| 4 |
A |
1660 |
1660 |
9.73 |
|
|
|
| 5 |
A |
1200 |
1200 |
0.03 |
|
|
|
| 6 |
A |
825 |
825 |
2.79 |
|
|
|
| 7 |
A |
551 |
551 |
142.11 |
|
|
|
| 8 |
A |
425 |
425 |
15.63 |
|
|
|
| 9 |
A |
411 |
411 |
10.35 |
|
|
|
| 10 |
A |
188 |
188 |
23.43 |
|
|
|
| 11 |
B |
3579 |
3579 |
10.95 |
|
|
|
| 12 |
B |
3503 |
3503 |
2.83 |
|
|
|
| 13 |
B |
1660 |
1660 |
21.04 |
|
|
|
| 14 |
B |
1376 |
1376 |
107.63 |
|
|
|
| 15 |
B |
1200 |
1200 |
0.08 |
|
|
|
| 16 |
B |
613 |
613 |
335.95 |
|
|
|
| 17 |
B |
413 |
413 |
16.66 |
|
|
|
| 18 |
B |
189 |
189 |
25.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13613.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13613.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.002 |
0.605 |
0.059 |
| C2 |
-0.002 |
-0.605 |
0.059 |
| N3 |
-0.002 |
1.953 |
-0.087 |
| N4 |
0.002 |
-1.953 |
-0.087 |
| H5 |
-0.339 |
2.473 |
0.714 |
| H6 |
0.851 |
2.349 |
-0.462 |
| H7 |
0.339 |
-2.473 |
0.714 |
| H8 |
-0.851 |
-2.349 |
-0.462 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2106 | 1.3559 | 2.5628 | 2.0083 | 2.0086 | 3.1651 | 3.1192 |
C2 | 1.2106 | | 2.5628 | 1.3559 | 3.1651 | 3.1192 | 2.0083 | 2.0086 | N3 | 1.3559 | 2.5628 | | 3.9067 | 1.0125 | 1.0127 | 4.5110 | 4.4018 | N4 | 2.5628 | 1.3559 | 3.9067 | | 4.5110 | 4.4018 | 1.0125 | 1.0127 | H5 | 2.0083 | 3.1651 | 1.0125 | 4.5110 | | 1.6780 | 4.9920 | 4.9901 | H6 | 2.0086 | 3.1192 | 1.0127 | 4.4018 | 1.6780 | | 4.9901 | 4.9975 | H7 | 3.1651 | 2.0083 | 4.5110 | 1.0125 | 4.9920 | 4.9901 | | 1.6780 | H8 | 3.1192 | 2.0086 | 4.4018 | 1.0127 | 4.9901 | 4.9975 | 1.6780 | |
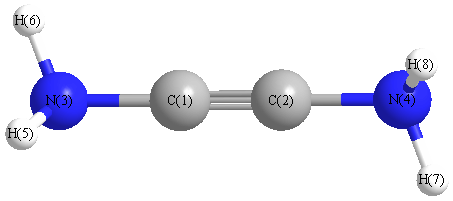 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
173.818 |
|
C1 |
N3 |
H5 |
115.217 |
| C1 |
N3 |
H6 |
115.227 |
|
C2 |
C1 |
N3 |
173.818 |
| C2 |
N4 |
H7 |
115.217 |
|
C2 |
N4 |
H8 |
115.227 |
| H5 |
N3 |
H6 |
111.907 |
|
H7 |
N4 |
H8 |
111.907 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.142 |
|
|
|
| 2 |
C |
0.142 |
|
|
|
| 3 |
N |
-0.580 |
|
|
|
| 4 |
N |
-0.580 |
|
|
|
| 5 |
H |
0.218 |
|
|
|
| 6 |
H |
0.220 |
|
|
|
| 7 |
H |
0.218 |
|
|
|
| 8 |
H |
0.220 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.443 |
1.443 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.180 |
0.248 |
0.000 |
| y |
0.248 |
9.498 |
0.000 |
| z |
0.000 |
0.000 |
3.171 |
<r2> (average value of r
2) Å
2
| <r2> |
96.971 |
| (<r2>)1/2 |
9.847 |
Jump to
S1C1
Energy calculated at TPSSh/6-311G**
| | hartrees |
|---|
| Energy at 0K | -188.091552 |
| Energy at 298.15K | |
| HF Energy | -188.091552 |
| Nuclear repulsion energy | 102.324840 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3560 |
3560 |
0.00 |
|
|
|
| 2 |
A1 |
2331 |
2331 |
0.00 |
|
|
|
| 3 |
A1 |
1646 |
1646 |
0.00 |
|
|
|
| 4 |
A1 |
846 |
846 |
0.00 |
|
|
|
| 5 |
B1 |
551 |
551 |
0.00 |
|
|
|
| 6 |
B2 |
3567 |
3567 |
5.50 |
|
|
|
| 7 |
B2 |
1656 |
1656 |
34.74 |
|
|
|
| 8 |
B2 |
1434 |
1434 |
144.02 |
|
|
|
| 9 |
E |
3657 |
3657 |
22.71 |
|
|
|
| 9 |
E |
3657 |
3657 |
22.71 |
|
|
|
| 10 |
E |
1142 |
1142 |
1.57 |
|
|
|
| 10 |
E |
1142 |
1142 |
1.57 |
|
|
|
| 11 |
E |
426 |
426 |
0.79 |
|
|
|
| 11 |
E |
426 |
426 |
0.79 |
|
|
|
| 12 |
E |
150 |
150 |
34.77 |
|
|
|
| 12 |
E |
150 |
150 |
34.77 |
|
|
|
| 13 |
E |
403i |
403i |
243.05 |
|
|
|
| 13 |
E |
403i |
403i |
243.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12767.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12767.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-311G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.608 |
| C2 |
0.000 |
0.000 |
-0.608 |
| N3 |
0.000 |
0.000 |
1.942 |
| N4 |
0.000 |
0.000 |
-1.942 |
| H5 |
0.000 |
0.859 |
2.467 |
| H6 |
0.000 |
-0.859 |
2.467 |
| H7 |
0.859 |
0.000 |
-2.467 |
| H8 |
-0.859 |
0.000 |
-2.467 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2162 | 1.3337 | 2.5498 | 2.0479 | 2.0479 | 3.1929 | 3.1929 |
C2 | 1.2162 | | 2.5498 | 1.3337 | 3.1929 | 3.1929 | 2.0479 | 2.0479 | N3 | 1.3337 | 2.5498 | | 3.8835 | 1.0072 | 1.0072 | 4.4917 | 4.4917 | N4 | 2.5498 | 1.3337 | 3.8835 | | 4.4917 | 4.4917 | 1.0072 | 1.0072 | H5 | 2.0479 | 3.1929 | 1.0072 | 4.4917 | | 1.7188 | 5.0815 | 5.0815 | H6 | 2.0479 | 3.1929 | 1.0072 | 4.4917 | 1.7188 | | 5.0815 | 5.0815 | H7 | 3.1929 | 2.0479 | 4.4917 | 1.0072 | 5.0815 | 5.0815 | | 1.7188 | H8 | 3.1929 | 2.0479 | 4.4917 | 1.0072 | 5.0815 | 5.0815 | 1.7188 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.432 |
| C1 |
N3 |
H6 |
121.432 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.432 |
|
C2 |
N4 |
H8 |
121.432 |
| H5 |
N3 |
H6 |
117.135 |
|
H7 |
N4 |
H8 |
117.135 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.170 |
|
|
|
| 2 |
C |
0.170 |
|
|
|
| 3 |
N |
-0.619 |
|
|
|
| 4 |
N |
-0.619 |
|
|
|
| 5 |
H |
0.225 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
H |
0.225 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.032 |
0.000 |
0.000 |
| y |
0.000 |
3.032 |
0.000 |
| z |
0.000 |
0.000 |
9.747 |
<r2> (average value of r
2) Å
2
| <r2> |
96.560 |
| (<r2>)1/2 |
9.827 |
