Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -245.119501 |
| Energy at 298.15K | |
| HF Energy | -245.119501 |
| Nuclear repulsion energy | 124.027947 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3883 |
3883 |
106.81 |
|
|
|
| 2 |
A' |
3811 |
3811 |
86.00 |
|
|
|
| 3 |
A' |
3672 |
3672 |
73.20 |
|
|
|
| 4 |
A' |
1898 |
1898 |
579.74 |
|
|
|
| 5 |
A' |
1623 |
1623 |
180.46 |
|
|
|
| 6 |
A' |
1457 |
1457 |
135.20 |
|
|
|
| 7 |
A' |
1253 |
1253 |
221.57 |
|
|
|
| 8 |
A' |
1089 |
1089 |
29.93 |
|
|
|
| 9 |
A' |
987 |
987 |
35.58 |
|
|
|
| 10 |
A' |
604 |
604 |
39.15 |
|
|
|
| 11 |
A' |
496 |
496 |
5.75 |
|
|
|
| 12 |
A" |
803 |
803 |
35.15 |
|
|
|
| 13 |
A" |
583 |
583 |
55.78 |
|
|
|
| 14 |
A" |
465 |
465 |
80.34 |
|
|
|
| 15 |
A" |
135i |
135i |
250.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11244.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11244.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.131 |
0.000 |
| O2 |
-0.064 |
1.332 |
0.000 |
| N3 |
1.136 |
-0.605 |
0.000 |
| O4 |
-1.088 |
-0.674 |
0.000 |
| H5 |
2.016 |
-0.123 |
0.000 |
| H6 |
1.101 |
-1.609 |
0.000 |
| H7 |
-1.851 |
-0.085 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2030 | 1.3541 | 1.3538 | 2.0322 | 2.0588 | 1.8633 |
O2 | 1.2030 | | 2.2796 | 2.2528 | 2.5391 | 3.1634 | 2.2805 | N3 | 1.3541 | 2.2796 | | 2.2255 | 1.0034 | 1.0037 | 3.0319 | O4 | 1.3538 | 2.2528 | 2.2255 | | 3.1529 | 2.3802 | 0.9636 | H5 | 2.0322 | 2.5391 | 1.0034 | 3.1529 | | 1.7446 | 3.8671 | H6 | 2.0588 | 3.1634 | 1.0037 | 2.3802 | 1.7446 | | 3.3216 | H7 | 1.8633 | 2.2805 | 3.0319 | 0.9636 | 3.8671 | 3.3216 | |
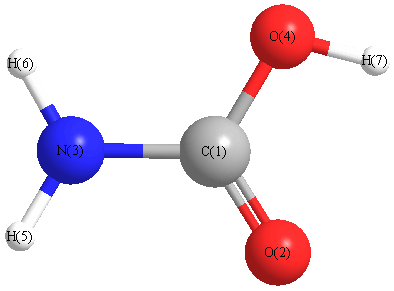 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.323 |
|
C1 |
N3 |
H6 |
120.946 |
| C1 |
O4 |
H7 |
105.803 |
|
O2 |
C1 |
N3 |
126.018 |
| O2 |
C1 |
O4 |
123.445 |
|
N3 |
C1 |
O4 |
110.538 |
| H5 |
N3 |
H6 |
120.731 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.534 |
|
|
|
| 2 |
O |
-0.418 |
|
|
|
| 3 |
N |
-0.538 |
|
|
|
| 4 |
O |
-0.394 |
|
|
|
| 5 |
H |
0.268 |
|
|
|
| 6 |
H |
0.265 |
|
|
|
| 7 |
H |
0.284 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.780 |
-2.287 |
0.000 |
2.416 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.253 |
-2.447 |
0.000 |
| y |
-2.447 |
-26.072 |
0.000 |
| z |
0.000 |
0.000 |
-23.757 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.661 |
-2.447 |
0.000 |
| y |
-2.447 |
-6.567 |
0.000 |
| z |
0.000 |
0.000 |
-3.094 |
|
| Polar |
|---|
| 3z2-r2 | -6.188 |
|---|
| x2-y2 | 10.819 |
|---|
| xy | -2.447 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.102 |
-0.319 |
0.000 |
| y |
-0.319 |
4.056 |
0.000 |
| z |
0.000 |
0.000 |
2.028 |
<r2> (average value of r
2) Å
2
| <r2> |
63.914 |
| (<r2>)1/2 |
7.995 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -245.119529 |
| Energy at 298.15K | -245.124171 |
| HF Energy | -245.119529 |
| Nuclear repulsion energy | 124.019471 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3885 |
3885 |
106.07 |
|
|
|
| 2 |
A |
3798 |
3798 |
80.80 |
|
|
|
| 3 |
A |
3662 |
3662 |
68.39 |
|
|
|
| 4 |
A |
1899 |
1899 |
570.10 |
|
|
|
| 5 |
A |
1624 |
1624 |
171.44 |
|
|
|
| 6 |
A |
1457 |
1457 |
136.37 |
|
|
|
| 7 |
A |
1258 |
1258 |
215.55 |
|
|
|
| 8 |
A |
1093 |
1093 |
37.83 |
|
|
|
| 9 |
A |
989 |
989 |
34.28 |
|
|
|
| 10 |
A |
802 |
802 |
40.48 |
|
|
|
| 11 |
A |
607 |
607 |
41.24 |
|
|
|
| 12 |
A |
583 |
583 |
62.37 |
|
|
|
| 13 |
A |
499 |
499 |
21.67 |
|
|
|
| 14 |
A |
473 |
473 |
56.59 |
|
|
|
| 15 |
A |
187 |
187 |
257.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11407.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11407.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.041 |
0.124 |
-0.001 |
| O2 |
-0.468 |
1.249 |
0.004 |
| N3 |
1.269 |
-0.230 |
-0.032 |
| O4 |
-0.830 |
-0.974 |
0.002 |
| H5 |
1.948 |
0.500 |
0.085 |
| H6 |
1.535 |
-1.189 |
0.101 |
| H7 |
-1.736 |
-0.646 |
0.004 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2027 | 1.3565 | 1.3530 | 2.0258 | 2.0540 | 1.8626 |
O2 | 1.2027 | | 2.2807 | 2.2523 | 2.5305 | 3.1565 | 2.2804 | N3 | 1.3565 | 2.2807 | | 2.2272 | 1.0042 | 1.0044 | 3.0338 | O4 | 1.3530 | 2.2523 | 2.2272 | | 3.1465 | 2.3775 | 0.9636 | H5 | 2.0258 | 2.5305 | 1.0042 | 3.1465 | | 1.7387 | 3.8596 | H6 | 2.0540 | 3.1565 | 1.0044 | 2.3775 | 1.7387 | | 3.3178 | H7 | 1.8626 | 2.2804 | 3.0338 | 0.9636 | 3.8596 | 3.3178 | |
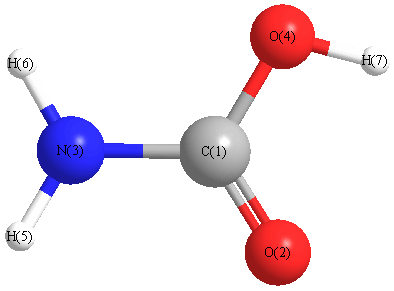 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.441 |
|
C1 |
N3 |
H6 |
120.177 |
| C1 |
O4 |
H7 |
105.812 |
|
O2 |
C1 |
N3 |
125.933 |
| O2 |
C1 |
O4 |
123.490 |
|
N3 |
C1 |
O4 |
110.569 |
| H5 |
N3 |
H6 |
119.912 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.528 |
|
|
|
| 2 |
O |
-0.417 |
|
|
|
| 3 |
N |
-0.531 |
|
|
|
| 4 |
O |
-0.392 |
|
|
|
| 5 |
H |
0.265 |
|
|
|
| 6 |
H |
0.262 |
|
|
|
| 7 |
H |
0.284 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.376 |
-1.917 |
0.402 |
2.393 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.986 |
1.190 |
0.789 |
| y |
1.190 |
-26.505 |
-0.210 |
| z |
0.789 |
-0.210 |
-23.722 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.128 |
1.190 |
0.789 |
| y |
1.190 |
-7.151 |
-0.210 |
| z |
0.789 |
-0.210 |
-2.977 |
|
| Polar |
|---|
| 3z2-r2 | -5.953 |
|---|
| x2-y2 | 11.519 |
|---|
| xy | 1.190 |
|---|
| xz | 0.789 |
|---|
| yz | -0.210 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.284 |
-0.240 |
0.006 |
| y |
-0.240 |
3.890 |
0.000 |
| z |
0.006 |
0.000 |
2.039 |
<r2> (average value of r
2) Å
2
| <r2> |
63.919 |
| (<r2>)1/2 |
7.995 |
