Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -133.264952 |
| Energy at 298.15K | -133.269136 |
| HF Energy | -133.264952 |
| Nuclear repulsion energy | 65.057978 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3470 |
3470 |
1.64 |
|
|
|
| 2 |
A' |
3285 |
3285 |
1.11 |
|
|
|
| 3 |
A' |
3171 |
3171 |
2.08 |
|
|
|
| 4 |
A' |
3077 |
3077 |
36.48 |
|
|
|
| 5 |
A' |
1515 |
1515 |
10.28 |
|
|
|
| 6 |
A' |
1485 |
1485 |
3.26 |
|
|
|
| 7 |
A' |
1389 |
1389 |
40.46 |
|
|
|
| 8 |
A' |
1264 |
1264 |
10.68 |
|
|
|
| 9 |
A' |
1099 |
1099 |
25.69 |
|
|
|
| 10 |
A' |
1000 |
1000 |
6.82 |
|
|
|
| 11 |
A' |
497 |
497 |
13.05 |
|
|
|
| 12 |
A" |
1076 |
1076 |
0.03 |
|
|
|
| 13 |
A" |
808 |
808 |
8.34 |
|
|
|
| 14 |
A" |
696 |
696 |
116.00 |
|
|
|
| 15 |
A" |
507 |
507 |
3.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12169.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12169.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| C2 |
1.146 |
-0.394 |
0.000 |
| N3 |
-1.186 |
-0.118 |
0.000 |
| H4 |
0.155 |
1.503 |
0.000 |
| H5 |
2.142 |
0.028 |
0.000 |
| H6 |
1.021 |
-1.468 |
0.000 |
| H7 |
-1.897 |
0.614 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4054 | 1.3018 | 1.0944 | 2.1773 | 2.1462 | 1.9069 |
C2 | 1.4054 | | 2.3483 | 2.1401 | 1.0809 | 1.0820 | 3.2058 | N3 | 1.3018 | 2.3483 | | 2.1033 | 3.3307 | 2.5871 | 1.0207 | H4 | 1.0944 | 2.1401 | 2.1033 | | 2.4748 | 3.0949 | 2.2359 | H5 | 2.1773 | 1.0809 | 3.3307 | 2.4748 | | 1.8692 | 4.0811 | H6 | 2.1462 | 1.0820 | 2.5871 | 3.0949 | 1.8692 | | 3.5849 | H7 | 1.9069 | 3.2058 | 1.0207 | 2.2359 | 4.0811 | 3.5849 | |
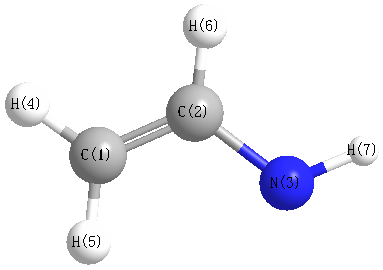 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.714 |
|
C1 |
C2 |
H6 |
118.700 |
| C1 |
N3 |
H7 |
109.789 |
|
C2 |
C1 |
N3 |
120.268 |
| C2 |
C1 |
H4 |
117.220 |
|
N3 |
C1 |
H4 |
122.512 |
| H5 |
C2 |
H6 |
119.586 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.028 |
|
|
|
| 2 |
C |
-0.234 |
|
|
|
| 3 |
N |
-0.353 |
|
|
|
| 4 |
H |
0.116 |
|
|
|
| 5 |
H |
0.150 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.187 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.166 |
1.605 |
0.000 |
1.984 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.962 |
-2.803 |
0.000 |
| y |
-2.803 |
-17.606 |
0.000 |
| z |
0.000 |
0.000 |
-20.310 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.996 |
-2.803 |
0.000 |
| y |
-2.803 |
1.030 |
0.000 |
| z |
0.000 |
0.000 |
-3.026 |
|
| Polar |
|---|
| 3z2-r2 | -6.052 |
|---|
| x2-y2 | 0.644 |
|---|
| xy | -2.803 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.544 |
-0.440 |
0.000 |
| y |
-0.440 |
4.187 |
0.000 |
| z |
0.000 |
0.000 |
2.319 |
<r2> (average value of r
2) Å
2
| <r2> |
45.279 |
| (<r2>)1/2 |
6.729 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -133.264097 |
| Energy at 298.15K | -133.268277 |
| HF Energy | -133.264097 |
| Nuclear repulsion energy | 65.054541 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3430 |
3430 |
2.77 |
|
|
|
| 2 |
A' |
3269 |
3269 |
3.59 |
|
|
|
| 3 |
A' |
3161 |
3161 |
1.81 |
|
|
|
| 4 |
A' |
3126 |
3126 |
17.38 |
|
|
|
| 5 |
A' |
1502 |
1502 |
7.94 |
|
|
|
| 6 |
A' |
1471 |
1471 |
2.53 |
|
|
|
| 7 |
A' |
1404 |
1404 |
7.47 |
|
|
|
| 8 |
A' |
1260 |
1260 |
52.94 |
|
|
|
| 9 |
A' |
1122 |
1122 |
21.38 |
|
|
|
| 10 |
A' |
989 |
989 |
0.32 |
|
|
|
| 11 |
A' |
505 |
505 |
8.26 |
|
|
|
| 12 |
A" |
1101 |
1101 |
56.24 |
|
|
|
| 13 |
A" |
819 |
819 |
68.14 |
|
|
|
| 14 |
A" |
702 |
702 |
1.95 |
|
|
|
| 15 |
A" |
481 |
481 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12170.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12170.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.122 |
-0.398 |
0.000 |
| N3 |
-1.249 |
0.083 |
0.000 |
| H4 |
0.184 |
1.525 |
0.000 |
| H5 |
2.128 |
-0.001 |
0.000 |
| H6 |
0.996 |
-1.475 |
0.000 |
| H7 |
-1.298 |
-0.941 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4066 | 1.3017 | 1.0907 | 2.1752 | 2.1675 | 1.9028 |
C2 | 1.4066 | | 2.4191 | 2.1401 | 1.0814 | 1.0839 | 2.4804 | N3 | 1.3017 | 2.4191 | | 2.0327 | 3.3777 | 2.7325 | 1.0250 | H4 | 1.0907 | 2.1401 | 2.0327 | | 2.4719 | 3.1082 | 2.8771 | H5 | 2.1752 | 1.0814 | 3.3777 | 2.4719 | | 1.8579 | 3.5527 | H6 | 2.1675 | 1.0839 | 2.7325 | 3.1082 | 1.8579 | | 2.3558 | H7 | 1.9028 | 2.4804 | 1.0250 | 2.8771 | 3.5527 | 2.3558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.373 |
|
C1 |
C2 |
H6 |
120.433 |
| C1 |
N3 |
H7 |
109.153 |
|
C2 |
C1 |
N3 |
126.522 |
| C2 |
C1 |
H4 |
117.404 |
|
N3 |
C1 |
H4 |
116.074 |
| H5 |
C2 |
H6 |
118.195 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.019 |
|
|
|
| 2 |
C |
-0.263 |
|
|
|
| 3 |
N |
-0.338 |
|
|
|
| 4 |
H |
0.142 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.244 |
-1.116 |
0.000 |
2.506 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.805 |
2.168 |
0.000 |
| y |
2.168 |
-16.312 |
0.000 |
| z |
0.000 |
0.000 |
-20.310 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.494 |
2.168 |
0.000 |
| y |
2.168 |
4.246 |
0.000 |
| z |
0.000 |
0.000 |
-1.752 |
|
| Polar |
|---|
| 3z2-r2 | -3.504 |
|---|
| x2-y2 | -4.494 |
|---|
| xy | 2.168 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.274 |
-0.151 |
0.000 |
| y |
-0.151 |
4.317 |
0.000 |
| z |
0.000 |
0.000 |
2.329 |
<r2> (average value of r
2) Å
2
| <r2> |
45.266 |
| (<r2>)1/2 |
6.728 |
