Jump to
S1C2
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -322.847639 |
| Energy at 298.15K | |
| HF Energy | -321.818568 |
| Nuclear repulsion energy | 234.270288 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3173 |
3015 |
24.12 |
|
|
|
| 2 |
A' |
3097 |
2943 |
25.95 |
|
|
|
| 3 |
A' |
3077 |
2923 |
38.13 |
|
|
|
| 4 |
A' |
3074 |
2921 |
8.06 |
|
|
|
| 5 |
A' |
1656 |
1574 |
126.30 |
|
|
|
| 6 |
A' |
1544 |
1467 |
16.45 |
|
|
|
| 7 |
A' |
1528 |
1452 |
3.15 |
|
|
|
| 8 |
A' |
1514 |
1439 |
0.93 |
|
|
|
| 9 |
A' |
1443 |
1371 |
22.01 |
|
|
|
| 10 |
A' |
1433 |
1361 |
6.16 |
|
|
|
| 11 |
A' |
1345 |
1278 |
7.36 |
|
|
|
| 12 |
A' |
1182 |
1123 |
6.77 |
|
|
|
| 13 |
A' |
1110 |
1054 |
38.25 |
|
|
|
| 14 |
A' |
1075 |
1021 |
7.37 |
|
|
|
| 15 |
A' |
943 |
896 |
85.48 |
|
|
|
| 16 |
A' |
845 |
803 |
493.30 |
|
|
|
| 17 |
A' |
663 |
630 |
90.92 |
|
|
|
| 18 |
A' |
394 |
374 |
0.57 |
|
|
|
| 19 |
A' |
353 |
336 |
4.06 |
|
|
|
| 20 |
A' |
156 |
148 |
0.38 |
|
|
|
| 21 |
A" |
3169 |
3011 |
57.67 |
|
|
|
| 22 |
A" |
3149 |
2992 |
2.64 |
|
|
|
| 23 |
A" |
3130 |
2974 |
9.27 |
|
|
|
| 24 |
A" |
1522 |
1446 |
7.89 |
|
|
|
| 25 |
A" |
1336 |
1270 |
0.10 |
|
|
|
| 26 |
A" |
1292 |
1228 |
0.40 |
|
|
|
| 27 |
A" |
1208 |
1148 |
1.23 |
|
|
|
| 28 |
A" |
910 |
865 |
1.25 |
|
|
|
| 29 |
A" |
780 |
741 |
1.31 |
|
|
|
| 30 |
A" |
242 |
230 |
0.03 |
|
|
|
| 31 |
A" |
211 |
201 |
0.92 |
|
|
|
| 32 |
A" |
98 |
94 |
1.32 |
|
|
|
| 33 |
A" |
56i |
53i |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23297.4 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 22137.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.906 |
2.212 |
0.000 |
| C2 |
-1.508 |
0.735 |
0.000 |
| C3 |
0.000 |
0.573 |
0.000 |
| O4 |
0.278 |
-0.837 |
0.000 |
| N5 |
1.675 |
-1.032 |
0.000 |
| O6 |
1.939 |
-2.187 |
0.000 |
| H7 |
-2.993 |
2.323 |
0.000 |
| H8 |
-1.515 |
2.723 |
0.885 |
| H9 |
-1.515 |
2.723 |
-0.885 |
| H10 |
-1.915 |
0.231 |
-0.882 |
| H11 |
-1.915 |
0.231 |
0.882 |
| H12 |
0.439 |
1.034 |
0.891 |
| H13 |
0.439 |
1.034 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5291 | 2.5135 | 3.7503 | 4.8315 | 5.8420 | 1.0927 | 1.0943 | 1.0943 | 2.1683 | 2.1683 | 2.7716 | 2.7716 |
C2 | 1.5291 | | 1.5167 | 2.3795 | 3.6405 | 4.5188 | 2.1738 | 2.1756 | 2.1756 | 1.0945 | 1.0945 | 2.1625 | 2.1625 | C3 | 2.5135 | 1.5167 | | 1.4374 | 2.3197 | 3.3728 | 3.4669 | 2.7748 | 2.7748 | 2.1361 | 2.1361 | 1.0956 | 1.0956 | O4 | 3.7503 | 2.3795 | 1.4374 | | 1.4104 | 2.1401 | 4.5480 | 4.0830 | 4.0830 | 2.5938 | 2.5938 | 2.0792 | 2.0792 | N5 | 4.8315 | 3.6405 | 2.3197 | 1.4104 | | 1.1845 | 5.7483 | 5.0056 | 5.0056 | 3.9065 | 3.9065 | 2.5672 | 2.5672 | O6 | 5.8420 | 4.5188 | 3.3728 | 2.1401 | 1.1845 | | 6.6827 | 6.0676 | 6.0676 | 4.6343 | 4.6343 | 3.6630 | 3.6630 | H7 | 1.0927 | 2.1738 | 3.4669 | 4.5480 | 5.7483 | 6.6827 | | 1.7686 | 1.7686 | 2.5130 | 2.5130 | 3.7731 | 3.7731 | H8 | 1.0943 | 2.1756 | 2.7748 | 4.0830 | 5.0056 | 6.0676 | 1.7686 | | 1.7704 | 3.0809 | 2.5239 | 2.5828 | 3.1348 | H9 | 1.0943 | 2.1756 | 2.7748 | 4.0830 | 5.0056 | 6.0676 | 1.7686 | 1.7704 | | 2.5239 | 3.0809 | 3.1348 | 2.5828 | H10 | 2.1683 | 1.0945 | 2.1361 | 2.5938 | 3.9065 | 4.6343 | 2.5130 | 3.0809 | 2.5239 | | 1.7634 | 3.0552 | 2.4881 | H11 | 2.1683 | 1.0945 | 2.1361 | 2.5938 | 3.9065 | 4.6343 | 2.5130 | 2.5239 | 3.0809 | 1.7634 | | 2.4881 | 3.0552 | H12 | 2.7716 | 2.1625 | 1.0956 | 2.0792 | 2.5672 | 3.6630 | 3.7731 | 2.5828 | 3.1348 | 3.0552 | 2.4881 | | 1.7828 | H13 | 2.7716 | 2.1625 | 1.0956 | 2.0792 | 2.5672 | 3.6630 | 3.7731 | 3.1348 | 2.5828 | 2.4881 | 3.0552 | 1.7828 | |
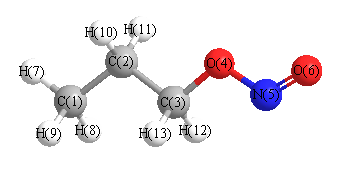 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.226 |
|
C1 |
C2 |
H10 |
110.380 |
| C1 |
C2 |
H11 |
110.380 |
|
C2 |
C1 |
H7 |
110.923 |
| C2 |
C1 |
H8 |
110.970 |
|
C2 |
C1 |
H9 |
110.970 |
| C2 |
C3 |
O4 |
107.286 |
|
C2 |
C3 |
H12 |
110.718 |
| C2 |
C3 |
H13 |
110.718 |
|
C3 |
C2 |
H10 |
108.706 |
| C3 |
C2 |
H11 |
108.706 |
|
C3 |
O4 |
N5 |
109.088 |
| O4 |
C3 |
H12 |
109.601 |
|
O4 |
C3 |
H13 |
109.601 |
| O4 |
N5 |
O6 |
110.829 |
|
H7 |
C1 |
H8 |
107.933 |
| H7 |
C1 |
H9 |
107.933 |
|
H8 |
C1 |
H9 |
107.978 |
| H10 |
C2 |
H11 |
107.335 |
|
H12 |
C3 |
H13 |
108.896 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-311G**
| | hartrees |
|---|
| Energy at 0K | -322.849986 |
| Energy at 298.15K | -322.859070 |
| HF Energy | -321.817738 |
| Nuclear repulsion energy | 241.719771 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3020 |
23.61 |
|
|
|
| 2 |
A |
3166 |
3009 |
33.63 |
|
|
|
| 3 |
A |
3151 |
2994 |
22.43 |
|
|
|
| 4 |
A |
3134 |
2978 |
11.25 |
|
|
|
| 5 |
A |
3085 |
2931 |
39.10 |
|
|
|
| 6 |
A |
3080 |
2927 |
11.06 |
|
|
|
| 7 |
A |
3078 |
2925 |
17.91 |
|
|
|
| 8 |
A |
1665 |
1582 |
149.40 |
|
|
|
| 9 |
A |
1530 |
1454 |
8.61 |
|
|
|
| 10 |
A |
1521 |
1445 |
7.44 |
|
|
|
| 11 |
A |
1510 |
1435 |
2.87 |
|
|
|
| 12 |
A |
1498 |
1423 |
2.54 |
|
|
|
| 13 |
A |
1438 |
1366 |
8.78 |
|
|
|
| 14 |
A |
1424 |
1353 |
11.40 |
|
|
|
| 15 |
A |
1394 |
1325 |
1.07 |
|
|
|
| 16 |
A |
1322 |
1256 |
5.00 |
|
|
|
| 17 |
A |
1311 |
1246 |
8.16 |
|
|
|
| 18 |
A |
1205 |
1145 |
12.25 |
|
|
|
| 19 |
A |
1138 |
1081 |
11.13 |
|
|
|
| 20 |
A |
1117 |
1061 |
31.70 |
|
|
|
| 21 |
A |
1024 |
973 |
10.66 |
|
|
|
| 22 |
A |
943 |
896 |
62.36 |
|
|
|
| 23 |
A |
902 |
857 |
1.78 |
|
|
|
| 24 |
A |
832 |
791 |
333.42 |
|
|
|
| 25 |
A |
788 |
748 |
113.35 |
|
|
|
| 26 |
A |
607 |
577 |
85.01 |
|
|
|
| 27 |
A |
456 |
433 |
19.39 |
|
|
|
| 28 |
A |
391 |
372 |
3.45 |
|
|
|
| 29 |
A |
283 |
269 |
2.06 |
|
|
|
| 30 |
A |
258 |
245 |
1.20 |
|
|
|
| 31 |
A |
184 |
175 |
0.43 |
|
|
|
| 32 |
A |
114 |
108 |
0.95 |
|
|
|
| 33 |
A |
56 |
53 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23390.6 cm
-1
Scaled (by 0.9502) Zero Point Vibrational Energy (zpe) 22225.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.161 |
-0.931 |
0.183 |
| C2 |
-1.758 |
0.458 |
-0.310 |
| C3 |
-0.409 |
0.900 |
0.222 |
| O4 |
0.561 |
-0.027 |
-0.299 |
| N5 |
1.781 |
0.210 |
0.316 |
| O6 |
2.571 |
-0.554 |
-0.141 |
| H7 |
-3.143 |
-1.214 |
-0.206 |
| H8 |
-2.211 |
-0.953 |
1.276 |
| H9 |
-1.433 |
-1.678 |
-0.140 |
| H10 |
-1.728 |
0.480 |
-1.404 |
| H11 |
-2.496 |
1.204 |
0.008 |
| H12 |
-0.154 |
1.910 |
-0.113 |
| H13 |
-0.385 |
0.863 |
1.316 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5277 | 2.5341 | 2.9084 | 4.1055 | 4.7576 | 1.0934 | 1.0948 | 1.0920 | 2.1674 | 2.1683 | 3.4910 | 2.7672 |
C2 | 1.5277 | | 1.5159 | 2.3700 | 3.6025 | 4.4489 | 2.1733 | 2.1707 | 2.1676 | 1.0951 | 1.0963 | 2.1729 | 2.1666 | C3 | 2.5341 | 1.5159 | | 1.4395 | 2.2978 | 3.3350 | 3.4819 | 2.7913 | 2.7974 | 2.1349 | 2.1198 | 1.0945 | 1.0953 | O4 | 2.9084 | 2.3700 | 1.4395 | | 1.3861 | 2.0832 | 3.8904 | 3.3200 | 2.5939 | 2.5920 | 3.3104 | 2.0740 | 2.0727 | N5 | 4.1055 | 3.6025 | 2.2978 | 1.3861 | | 1.1896 | 5.1516 | 4.2669 | 3.7552 | 3.9167 | 4.4016 | 2.6113 | 2.4730 | O6 | 4.7576 | 4.4489 | 3.3350 | 2.0832 | 1.1896 | | 5.7516 | 5.0029 | 4.1587 | 4.5980 | 5.3650 | 3.6738 | 3.5864 | H7 | 1.0934 | 2.1733 | 3.4819 | 3.8904 | 5.1516 | 5.7516 | | 1.7707 | 1.7724 | 2.5114 | 2.5126 | 4.3246 | 3.7735 | H8 | 1.0948 | 2.1707 | 2.7913 | 3.3200 | 4.2669 | 5.0029 | 1.7707 | | 1.7711 | 3.0778 | 2.5186 | 3.7892 | 2.5758 | H9 | 1.0920 | 2.1676 | 2.7974 | 2.5939 | 3.7552 | 4.1587 | 1.7724 | 1.7711 | | 2.5185 | 3.0757 | 3.8098 | 3.1109 | H10 | 2.1674 | 1.0951 | 2.1349 | 2.5920 | 3.9167 | 4.5980 | 2.5114 | 3.0778 | 2.5185 | | 1.7634 | 2.4880 | 3.0577 | H11 | 2.1683 | 1.0963 | 2.1198 | 3.3104 | 4.4016 | 5.3650 | 2.5126 | 2.5186 | 3.0757 | 1.7634 | | 2.4490 | 2.5069 | H12 | 3.4910 | 2.1729 | 1.0945 | 2.0740 | 2.6113 | 3.6738 | 4.3246 | 3.7892 | 3.8098 | 2.4880 | 2.4490 | | 1.7862 | H13 | 2.7672 | 2.1666 | 1.0953 | 2.0727 | 2.4730 | 3.5864 | 3.7735 | 2.5758 | 3.1109 | 3.0577 | 2.5069 | 1.7862 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.728 |
|
C1 |
C2 |
H10 |
110.363 |
| C1 |
C2 |
H11 |
110.361 |
|
C2 |
C1 |
H7 |
110.936 |
| C2 |
C1 |
H8 |
110.642 |
|
C2 |
C1 |
H9 |
110.560 |
| C2 |
C3 |
O4 |
106.595 |
|
C2 |
C3 |
H12 |
111.676 |
| C2 |
C3 |
H13 |
111.121 |
|
C3 |
C2 |
H10 |
108.632 |
| C3 |
C2 |
H11 |
107.393 |
|
C3 |
O4 |
N5 |
108.809 |
| O4 |
C3 |
H12 |
109.104 |
|
O4 |
C3 |
H13 |
108.950 |
| O4 |
N5 |
O6 |
107.709 |
|
H7 |
C1 |
H8 |
108.038 |
| H7 |
C1 |
H9 |
108.393 |
|
H8 |
C1 |
H9 |
108.174 |
| H10 |
C2 |
H11 |
107.162 |
|
H12 |
C3 |
H13 |
109.311 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
