Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -93.938240 |
| Energy at 298.15K | -93.939544 |
| HF Energy | -93.938240 |
| Nuclear repulsion energy | 28.291234 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3341 |
3341 |
4.92 |
|
|
|
| 2 |
A' |
3022 |
3022 |
33.45 |
|
|
|
| 3 |
A' |
1886 |
1886 |
37.82 |
|
|
|
| 4 |
A' |
1048 |
1048 |
130.39 |
|
|
|
| 5 |
A' |
859 |
859 |
115.47 |
|
|
|
| 6 |
A" |
933 |
933 |
3.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5544.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5544.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.112 |
0.633 |
0.000 |
| N2 |
0.112 |
-0.587 |
0.000 |
| H3 |
-0.673 |
1.400 |
0.000 |
| H4 |
-0.782 |
-1.092 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2205 | 1.0972 | 1.9432 |
N2 | 1.2205 | | 2.1368 | 1.0269 | H3 | 1.0972 | 2.1368 | | 2.4947 | H4 | 1.9432 | 1.0269 | 2.4947 | |
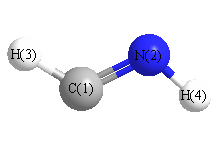 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
119.440 |
|
H3 |
C1 |
N2 |
134.353 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.001 |
|
|
|
| 2 |
N |
-0.351 |
|
|
|
| 3 |
H |
0.137 |
|
|
|
| 4 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.268 |
0.517 |
0.000 |
2.326 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.053 |
0.682 |
0.000 |
| y |
0.682 |
-11.216 |
0.000 |
| z |
0.000 |
0.000 |
-12.658 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.116 |
0.682 |
0.000 |
| y |
0.682 |
1.139 |
0.000 |
| z |
0.000 |
0.000 |
-1.023 |
|
| Polar |
|---|
| 3z2-r2 | -2.047 |
|---|
| x2-y2 | -0.837 |
|---|
| xy | 0.682 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.132 |
0.116 |
0.000 |
| y |
0.116 |
3.896 |
0.000 |
| z |
0.000 |
0.000 |
1.430 |
<r2> (average value of r
2) Å
2
| <r2> |
16.680 |
| (<r2>)1/2 |
4.084 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -93.945497 |
| Energy at 298.15K | -93.946841 |
| HF Energy | -93.945497 |
| Nuclear repulsion energy | 28.241374 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3450 |
3450 |
13.71 |
|
|
|
| 2 |
A' |
3052 |
3052 |
20.35 |
|
|
|
| 3 |
A' |
1854 |
1854 |
35.96 |
|
|
|
| 4 |
A' |
1202 |
1202 |
8.05 |
|
|
|
| 5 |
A' |
899 |
899 |
253.76 |
|
|
|
| 6 |
A" |
992 |
992 |
124.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5724.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5724.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.645 |
0.000 |
| N2 |
0.000 |
-0.583 |
0.000 |
| H3 |
0.900 |
1.272 |
0.000 |
| H4 |
-0.901 |
-1.064 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2278 | 1.0969 | 1.9320 |
N2 | 1.2278 | | 2.0615 | 1.0216 | H3 | 1.0969 | 2.0615 | | 2.9497 | H4 | 1.9320 | 1.0216 | 2.9497 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
118.085 |
|
H3 |
C1 |
N2 |
124.848 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.039 |
|
|
|
| 2 |
N |
-0.344 |
|
|
|
| 3 |
H |
0.150 |
|
|
|
| 4 |
H |
0.233 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.501 |
0.222 |
0.000 |
0.548 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.211 |
3.201 |
0.000 |
| y |
3.201 |
-11.477 |
0.000 |
| z |
0.000 |
0.000 |
-12.692 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.873 |
3.201 |
0.000 |
| y |
3.201 |
0.475 |
0.000 |
| z |
0.000 |
0.000 |
-1.348 |
|
| Polar |
|---|
| 3z2-r2 | -2.697 |
|---|
| x2-y2 | 0.266 |
|---|
| xy | 3.201 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.344 |
0.344 |
0.000 |
| y |
0.344 |
3.806 |
0.000 |
| z |
0.000 |
0.000 |
1.506 |
<r2> (average value of r
2) Å
2
| <r2> |
16.612 |
| (<r2>)1/2 |
4.076 |
