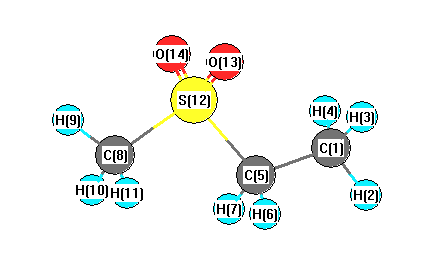
Vibrational Frequencies calculated at wB97X-D/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3198 |
3059 |
1.40 |
|
|
|
| 2 |
A |
3191 |
3053 |
0.62 |
|
|
|
| 3 |
A |
3162 |
3025 |
8.51 |
|
|
|
| 4 |
A |
3146 |
3009 |
3.75 |
|
|
|
| 5 |
A |
3136 |
2999 |
14.13 |
|
|
|
| 6 |
A |
3089 |
2955 |
3.79 |
|
|
|
| 7 |
A |
3084 |
2949 |
1.73 |
|
|
|
| 8 |
A |
3065 |
2932 |
16.06 |
|
|
|
| 9 |
A |
1512 |
1447 |
9.17 |
|
|
|
| 10 |
A |
1504 |
1439 |
11.88 |
|
|
|
| 11 |
A |
1468 |
1404 |
11.10 |
|
|
|
| 12 |
A |
1459 |
1395 |
5.95 |
|
|
|
| 13 |
A |
1451 |
1388 |
5.00 |
|
|
|
| 14 |
A |
1424 |
1362 |
1.79 |
|
|
|
| 15 |
A |
1366 |
1307 |
100.20 |
|
|
|
| 16 |
A |
1359 |
1300 |
139.40 |
|
|
|
| 17 |
A |
1319 |
1262 |
28.00 |
|
|
|
| 18 |
A |
1271 |
1216 |
7.46 |
|
|
|
| 19 |
A |
1160 |
1110 |
176.31 |
|
|
|
| 20 |
A |
1092 |
1045 |
1.50 |
|
|
|
| 21 |
A |
1076 |
1030 |
7.09 |
|
|
|
| 22 |
A |
1003 |
959 |
3.51 |
|
|
|
| 23 |
A |
979 |
936 |
17.30 |
|
|
|
| 24 |
A |
973 |
931 |
22.61 |
|
|
|
| 25 |
A |
812 |
776 |
52.11 |
|
|
|
| 26 |
A |
738 |
705 |
29.12 |
|
|
|
| 27 |
A |
658 |
630 |
13.61 |
|
|
|
| 28 |
A |
502 |
480 |
24.80 |
|
|
|
| 29 |
A |
454 |
435 |
36.08 |
|
|
|
| 30 |
A |
406 |
389 |
7.75 |
|
|
|
| 31 |
A |
327 |
313 |
0.95 |
|
|
|
| 32 |
A |
290 |
278 |
1.61 |
|
|
|
| 33 |
A |
248 |
238 |
2.84 |
|
|
|
| 34 |
A |
221 |
211 |
0.74 |
|
|
|
| 35 |
A |
172 |
164 |
0.93 |
|
|
|
| 36 |
A |
104 |
100 |
2.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25209.2 cm
-1
Scaled (by 0.9565) Zero Point Vibrational Energy (zpe) 24112.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.339 |
|
|
|
| 2 |
H |
0.166 |
|
|
|
| 3 |
H |
0.139 |
|
|
|
| 4 |
H |
0.124 |
|
|
|
| 5 |
C |
-0.449 |
|
|
|
| 6 |
H |
0.205 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
C |
-0.502 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
S |
0.948 |
|
|
|
| 13 |
O |
-0.509 |
|
|
|
| 14 |
O |
-0.505 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.332 |
3.011 |
-2.322 |
5.055 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.579 |
-5.563 |
-1.396 |
| y |
-5.563 |
-41.928 |
-0.553 |
| z |
-1.396 |
-0.553 |
-48.725 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.747 |
-5.563 |
-1.396 |
| y |
-5.563 |
4.724 |
-0.553 |
| z |
-1.396 |
-0.553 |
-5.471 |
|
| Polar |
|---|
| 3z2-r2 | -10.943 |
|---|
| x2-y2 | -2.652 |
|---|
| xy | -5.563 |
|---|
| xz | -1.396 |
|---|
| yz | -0.553 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.143 |
-0.169 |
0.059 |
| y |
-0.169 |
8.257 |
0.040 |
| z |
0.059 |
0.040 |
7.835 |
<r2> (average value of r
2) Å
2
| <r2> |
190.760 |
| (<r2>)1/2 |
13.812 |