Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -190.190520 |
| Energy at 298.15K | |
| HF Energy | -189.583237 |
| Nuclear repulsion energy | 88.197493 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3120 |
3120 |
26.04 |
|
|
|
| 2 |
A1 |
2338 |
2338 |
876.51 |
|
|
|
| 3 |
A1 |
1798 |
1798 |
32.92 |
|
|
|
| 4 |
A1 |
1501 |
1501 |
4.33 |
|
|
|
| 5 |
A1 |
922 |
922 |
3.03 |
|
|
|
| 6 |
B1 |
1005 |
1005 |
22.08 |
|
|
|
| 7 |
B1 |
650 |
650 |
26.31 |
|
|
|
| 8 |
B1 |
213 |
213 |
0.76 |
|
|
|
| 9 |
B2 |
3197 |
3197 |
10.04 |
|
|
|
| 10 |
B2 |
1076 |
1076 |
2.79 |
|
|
|
| 11 |
B2 |
466 |
466 |
15.38 |
|
|
|
| 12 |
B2 |
139i |
139i |
19.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8073.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8073.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.881 |
| C2 |
0.000 |
0.000 |
-0.557 |
| C3 |
0.000 |
0.000 |
0.725 |
| O4 |
0.000 |
0.000 |
1.899 |
| H5 |
0.000 |
0.927 |
-2.455 |
| H6 |
0.000 |
-0.927 |
-2.455 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3236 | 2.6059 | 3.7796 | 1.0906 | 1.0906 |
C2 | 1.3236 | | 1.2824 | 2.4560 | 2.1123 | 2.1123 | C3 | 2.6059 | 1.2824 | | 1.1736 | 3.3128 | 3.3128 | O4 | 3.7796 | 2.4560 | 1.1736 | | 4.4517 | 4.4517 | H5 | 1.0906 | 2.1123 | 3.3128 | 4.4517 | | 1.8541 | H6 | 1.0906 | 2.1123 | 3.3128 | 4.4517 | 1.8541 | |
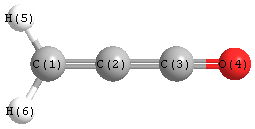 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.787 |
| C2 |
C1 |
H6 |
121.787 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.426 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -190.191819 |
| Energy at 298.15K | |
| HF Energy | -189.582170 |
| Nuclear repulsion energy | 88.464470 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3210 |
3210 |
9.15 |
|
|
|
| 2 |
A' |
3127 |
3127 |
24.39 |
|
|
|
| 3 |
A' |
2233 |
2233 |
919.34 |
|
|
|
| 4 |
A' |
1761 |
1761 |
8.06 |
|
|
|
| 5 |
A' |
1504 |
1504 |
1.91 |
|
|
|
| 6 |
A' |
1082 |
1082 |
26.90 |
|
|
|
| 7 |
A' |
943 |
943 |
1.42 |
|
|
|
| 8 |
A' |
506 |
506 |
13.35 |
|
|
|
| 9 |
A' |
164 |
164 |
25.11 |
|
|
|
| 10 |
A" |
1007 |
1007 |
24.07 |
|
|
|
| 11 |
A" |
715 |
715 |
15.64 |
|
|
|
| 12 |
A" |
270 |
270 |
2.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8261.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8261.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.054 |
-1.485 |
0.000 |
| C2 |
0.000 |
-0.667 |
0.000 |
| C3 |
-0.359 |
0.596 |
0.000 |
| O4 |
-0.897 |
1.629 |
0.000 |
| H5 |
2.084 |
-1.125 |
0.000 |
| H6 |
0.921 |
-2.565 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3334 | 2.5143 | 3.6739 | 1.0909 | 1.0888 |
C2 | 1.3334 | | 1.3130 | 2.4651 | 2.1332 | 2.1096 | C3 | 2.5143 | 1.3130 | | 1.1649 | 2.9873 | 3.4102 | O4 | 3.6739 | 2.4651 | 1.1649 | | 4.0577 | 4.5711 | H5 | 1.0909 | 2.1332 | 2.9873 | 4.0577 | | 1.8510 | H6 | 1.0888 | 2.1096 | 3.4102 | 4.5711 | 1.8510 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
143.642 |
|
C2 |
C1 |
H5 |
122.952 |
| C2 |
C1 |
H6 |
120.799 |
|
C2 |
C3 |
O4 |
168.324 |
| H5 |
C1 |
H6 |
116.249 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
