Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -277.730799 |
| Energy at 298.15K | |
| HF Energy | -277.002647 |
| Nuclear repulsion energy | 125.757954 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3095 |
3095 |
0.00 |
|
|
|
| 2 |
Ag |
1548 |
1548 |
0.00 |
|
|
|
| 3 |
Ag |
1505 |
1505 |
0.00 |
|
|
|
| 4 |
Ag |
1125 |
1125 |
0.00 |
|
|
|
| 5 |
Ag |
1107 |
1107 |
0.00 |
|
|
|
| 6 |
Ag |
470 |
470 |
0.00 |
|
|
|
| 7 |
Au |
3166 |
3166 |
61.77 |
|
|
|
| 8 |
Au |
1273 |
1273 |
5.71 |
|
|
|
| 9 |
Au |
834 |
834 |
0.56 |
|
|
|
| 10 |
Au |
135 |
135 |
13.89 |
|
|
|
| 11 |
Bg |
3142 |
3142 |
0.00 |
|
|
|
| 12 |
Bg |
1326 |
1326 |
0.00 |
|
|
|
| 13 |
Bg |
1209 |
1209 |
0.00 |
|
|
|
| 14 |
Bu |
3098 |
3098 |
60.17 |
|
|
|
| 15 |
Bu |
1555 |
1555 |
2.35 |
|
|
|
| 16 |
Bu |
1397 |
1397 |
19.31 |
|
|
|
| 17 |
Bu |
1120 |
1120 |
193.46 |
|
|
|
| 18 |
Bu |
288 |
288 |
20.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13695.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13695.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.630 |
0.000 |
| C2 |
-0.425 |
-0.630 |
0.000 |
| F3 |
-0.425 |
1.722 |
0.000 |
| F4 |
0.425 |
-1.722 |
0.000 |
| H5 |
1.054 |
0.669 |
0.894 |
| H6 |
1.054 |
0.669 |
-0.894 |
| H7 |
-1.054 |
-0.669 |
0.894 |
| H8 |
-1.054 |
-0.669 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5189 | 1.3834 | 2.3512 | 1.0939 | 1.0939 | 2.1612 | 2.1612 |
C2 | 1.5189 | | 2.3512 | 1.3834 | 2.1612 | 2.1612 | 1.0939 | 1.0939 | F3 | 1.3834 | 2.3512 | | 3.5464 | 2.0232 | 2.0232 | 2.6284 | 2.6284 | F4 | 2.3512 | 1.3834 | 3.5464 | | 2.6284 | 2.6284 | 2.0232 | 2.0232 | H5 | 1.0939 | 2.1612 | 2.0232 | 2.6284 | | 1.7883 | 2.4959 | 3.0704 | H6 | 1.0939 | 2.1612 | 2.0232 | 2.6284 | 1.7883 | | 3.0704 | 2.4959 | H7 | 2.1612 | 1.0939 | 2.6284 | 2.0232 | 2.4959 | 3.0704 | | 1.7883 | H8 | 2.1612 | 1.0939 | 2.6284 | 2.0232 | 3.0704 | 2.4959 | 1.7883 | |
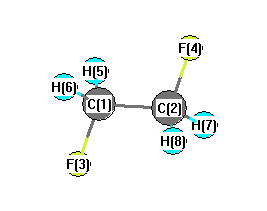 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.120 |
|
C1 |
C2 |
H7 |
110.562 |
| C1 |
C2 |
H8 |
110.562 |
|
C2 |
C1 |
F3 |
108.120 |
| C2 |
C1 |
H5 |
110.562 |
|
C2 |
C1 |
H6 |
110.562 |
| F3 |
C1 |
H5 |
108.951 |
|
F3 |
C1 |
H6 |
108.951 |
| F4 |
C2 |
H7 |
108.951 |
|
F4 |
C2 |
H8 |
108.951 |
| H5 |
C1 |
H6 |
109.648 |
|
H7 |
C2 |
H8 |
109.648 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -277.730999 |
| Energy at 298.15K | |
| HF Energy | -277.002379 |
| Nuclear repulsion energy | 127.945170 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3135 |
19.14 |
|
|
|
| 2 |
A |
3087 |
3087 |
37.35 |
|
|
|
| 3 |
A |
1527 |
1527 |
0.60 |
|
|
|
| 4 |
A |
1489 |
1489 |
18.13 |
|
|
|
| 5 |
A |
1335 |
1335 |
2.12 |
|
|
|
| 6 |
A |
1163 |
1163 |
60.00 |
|
|
|
| 7 |
A |
1160 |
1160 |
20.90 |
|
|
|
| 8 |
A |
898 |
898 |
27.82 |
|
|
|
| 9 |
A |
330 |
330 |
0.19 |
|
|
|
| 10 |
A |
155 |
155 |
3.82 |
|
|
|
| 11 |
B |
3148 |
3148 |
46.78 |
|
|
|
| 12 |
B |
3075 |
3075 |
15.13 |
|
|
|
| 13 |
B |
1524 |
1524 |
7.57 |
|
|
|
| 14 |
B |
1441 |
1441 |
12.79 |
|
|
|
| 15 |
B |
1297 |
1297 |
6.39 |
|
|
|
| 16 |
B |
1131 |
1131 |
41.89 |
|
|
|
| 17 |
B |
930 |
930 |
40.84 |
|
|
|
| 18 |
B |
508 |
508 |
17.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13666.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13666.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.265 |
0.706 |
0.513 |
| C2 |
-0.265 |
-0.706 |
0.513 |
| F3 |
-0.265 |
1.414 |
-0.549 |
| F4 |
0.265 |
-1.414 |
-0.549 |
| H5 |
-0.025 |
1.208 |
1.444 |
| H6 |
1.355 |
0.701 |
0.417 |
| H7 |
0.025 |
-1.208 |
1.444 |
| H8 |
-1.355 |
-0.701 |
0.417 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5087 | 1.3815 | 2.3714 | 1.0964 | 1.0945 | 2.1420 | 2.1477 |
C2 | 1.5087 | | 2.3714 | 1.3815 | 2.1420 | 2.1477 | 1.0964 | 1.0945 | F3 | 1.3815 | 2.3714 | | 2.8771 | 2.0179 | 2.0161 | 3.3059 | 2.5678 | F4 | 2.3714 | 1.3815 | 2.8771 | | 3.3059 | 2.5678 | 2.0179 | 2.0161 | H5 | 1.0964 | 2.1420 | 2.0179 | 3.3059 | | 1.7931 | 2.4160 | 2.5430 | H6 | 1.0945 | 2.1477 | 2.0161 | 2.5678 | 1.7931 | | 2.5430 | 3.0509 | H7 | 2.1420 | 1.0964 | 3.3059 | 2.0179 | 2.4160 | 2.5430 | | 1.7931 | H8 | 2.1477 | 1.0945 | 2.5678 | 2.0161 | 2.5430 | 3.0509 | 1.7931 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.195 |
|
C1 |
C2 |
H7 |
109.611 |
| C1 |
C2 |
H8 |
110.167 |
|
C2 |
C1 |
F3 |
110.195 |
| C2 |
C1 |
H5 |
109.611 |
|
C2 |
C1 |
H6 |
110.167 |
| F3 |
C1 |
H5 |
108.502 |
|
F3 |
C1 |
H6 |
108.474 |
| F4 |
C2 |
H7 |
108.502 |
|
F4 |
C2 |
H8 |
108.474 |
| H5 |
C1 |
H6 |
109.862 |
|
H7 |
C2 |
H8 |
109.862 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
