Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -498.790286 |
| Energy at 298.15K | |
| HF Energy | -498.496945 |
| Nuclear repulsion energy | 45.222514 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3202 |
3202 |
8.53 |
|
|
|
| 2 |
A' |
1460 |
1460 |
14.56 |
|
|
|
| 3 |
A' |
862 |
862 |
34.75 |
|
|
|
| 4 |
A' |
192 |
192 |
75.79 |
|
|
|
| 5 |
A" |
3351 |
3351 |
0.56 |
|
|
|
| 6 |
A" |
1033 |
1033 |
0.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5050.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5050.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.007 |
1.122 |
0.000 |
| Cl2 |
-0.007 |
-0.587 |
0.000 |
| H3 |
0.077 |
1.622 |
0.953 |
| H4 |
0.077 |
1.622 |
-0.953 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7087 | 1.0798 | 1.0798 |
Cl2 | 1.7087 | | 2.4070 | 2.4070 | H3 | 1.0798 | 2.4070 | | 1.9068 | H4 | 1.0798 | 2.4070 | 1.9068 | |
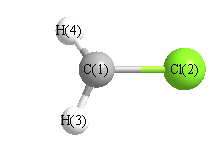 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.577 |
|
Br2 |
C1 |
H4 |
117.577 |
| H3 |
C1 |
H4 |
124.005 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -498.790279 |
| Energy at 298.15K | |
| HF Energy | -498.496864 |
| Nuclear repulsion energy | 45.233165 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3205 |
3205 |
7.98 |
|
|
|
| 2 |
A1 |
1460 |
1460 |
14.75 |
|
|
|
| 3 |
A1 |
863 |
863 |
34.15 |
|
|
|
| 4 |
B1 |
136i |
136i |
80.53 |
|
|
|
| 5 |
B2 |
3355 |
3355 |
0.40 |
|
|
|
| 6 |
B2 |
1031 |
1031 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4889.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4889.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.121 |
| Cl2 |
0.000 |
0.000 |
0.587 |
| H3 |
0.000 |
0.955 |
-1.625 |
| H4 |
0.000 |
-0.955 |
-1.625 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7079 | 1.0795 | 1.0795 |
Cl2 | 1.7079 | | 2.4089 | 2.4089 | H3 | 1.0795 | 2.4089 | | 1.9094 | H4 | 1.0795 | 2.4089 | 1.9094 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.817 |
|
Br2 |
C1 |
H4 |
117.817 |
| H3 |
C1 |
H4 |
124.367 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
