Jump to
S1C2
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -169.869479 |
| Energy at 298.15K | -169.873187 |
| HF Energy | -169.869479 |
| Nuclear repulsion energy | 71.451856 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3774 |
3774 |
57.36 |
|
|
|
| 2 |
A' |
3629 |
3629 |
49.67 |
|
|
|
| 3 |
A' |
3023 |
3023 |
91.49 |
|
|
|
| 4 |
A' |
1871 |
1871 |
428.91 |
|
|
|
| 5 |
A' |
1622 |
1622 |
85.44 |
|
|
|
| 6 |
A' |
1437 |
1437 |
5.38 |
|
|
|
| 7 |
A' |
1280 |
1280 |
117.67 |
|
|
|
| 8 |
A' |
1065 |
1065 |
4.23 |
|
|
|
| 9 |
A' |
574 |
574 |
12.90 |
|
|
|
| 10 |
A" |
1065 |
1065 |
0.54 |
|
|
|
| 11 |
A" |
654 |
654 |
26.24 |
|
|
|
| 12 |
A" |
218 |
218 |
241.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10106.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10106.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| O2 |
1.190 |
0.241 |
0.000 |
| N3 |
-0.932 |
-0.568 |
0.000 |
| H4 |
-0.456 |
1.426 |
0.000 |
| H5 |
-0.624 |
-1.528 |
0.000 |
| H6 |
-1.916 |
-0.366 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2034 | 1.3585 | 1.1050 | 2.0453 | 2.0710 |
O2 | 1.2034 | | 2.2711 | 2.0286 | 2.5337 | 3.1648 | N3 | 1.3585 | 2.2711 | | 2.0508 | 1.0076 | 1.0044 | H4 | 1.1050 | 2.0286 | 2.0508 | | 2.9591 | 2.3121 | H5 | 2.0453 | 2.5337 | 1.0076 | 2.9591 | | 1.7371 | H6 | 2.0710 | 3.1648 | 1.0044 | 2.3121 | 1.7371 | |
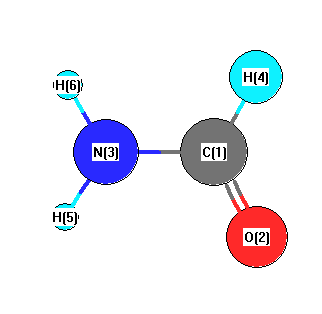 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.889 |
|
C1 |
N3 |
H6 |
121.720 |
| O2 |
C1 |
N3 |
124.760 |
|
O2 |
C1 |
H4 |
122.935 |
| N3 |
C1 |
H4 |
112.305 |
|
H5 |
N3 |
H6 |
119.392 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.302 |
|
|
|
| 2 |
O |
-0.383 |
|
|
|
| 3 |
N |
-0.508 |
|
|
|
| 4 |
H |
0.081 |
|
|
|
| 5 |
H |
0.257 |
|
|
|
| 6 |
H |
0.252 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.827 |
-0.649 |
0.000 |
3.882 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.070 |
-0.188 |
0.000 |
| y |
-0.188 |
-14.637 |
0.000 |
| z |
0.000 |
0.000 |
-18.661 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.421 |
-0.188 |
0.000 |
| y |
-0.188 |
3.729 |
0.000 |
| z |
0.000 |
0.000 |
-2.307 |
|
| Polar |
|---|
| 3z2-r2 | -4.614 |
|---|
| x2-y2 | -3.433 |
|---|
| xy | -0.188 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.163 |
0.270 |
0.000 |
| y |
0.270 |
3.273 |
0.000 |
| z |
0.000 |
0.000 |
1.783 |
<r2> (average value of r
2) Å
2
| <r2> |
40.660 |
| (<r2>)1/2 |
6.377 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G**
| | hartrees |
|---|
| Energy at 0K | -169.869457 |
| Energy at 298.15K | -169.873071 |
| HF Energy | -169.869457 |
| Nuclear repulsion energy | 71.446536 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3771 |
3771 |
58.29 |
|
|
|
| 2 |
A |
3628 |
3628 |
50.45 |
|
|
|
| 3 |
A |
3010 |
3010 |
93.56 |
|
|
|
| 4 |
A |
1868 |
1868 |
432.57 |
|
|
|
| 5 |
A |
1620 |
1620 |
85.56 |
|
|
|
| 6 |
A |
1434 |
1434 |
5.48 |
|
|
|
| 7 |
A |
1280 |
1280 |
117.04 |
|
|
|
| 8 |
A |
1068 |
1068 |
0.54 |
|
|
|
| 9 |
A |
1063 |
1063 |
3.87 |
|
|
|
| 10 |
A |
641 |
641 |
22.84 |
|
|
|
| 11 |
A |
572 |
572 |
12.84 |
|
|
|
| 12 |
A |
161 |
161 |
245.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10058.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10058.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.165 |
0.386 |
0.000 |
| O2 |
1.189 |
-0.246 |
0.000 |
| N3 |
-1.081 |
-0.156 |
-0.000 |
| H4 |
0.142 |
1.491 |
0.000 |
| H5 |
-1.173 |
-1.159 |
0.000 |
| H6 |
-1.906 |
0.416 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2037 | 1.3585 | 1.1047 | 2.0445 | 2.0715 |
O2 | 1.2037 | | 2.2716 | 2.0282 | 2.5329 | 3.1655 | N3 | 1.3585 | 2.2716 | | 2.0509 | 1.0075 | 1.0045 | H4 | 1.1047 | 2.0282 | 2.0509 | | 2.9584 | 2.3130 | H5 | 2.0445 | 2.5329 | 1.0075 | 2.9584 | | 1.7374 | H6 | 2.0715 | 3.1655 | 1.0045 | 2.3130 | 1.7374 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.809 |
|
C1 |
N3 |
H6 |
121.760 |
| O2 |
C1 |
N3 |
124.779 |
|
O2 |
C1 |
H4 |
122.895 |
| N3 |
C1 |
H4 |
112.326 |
|
H5 |
N3 |
H6 |
119.431 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.302 |
|
|
|
| 2 |
O |
-0.383 |
|
|
|
| 3 |
N |
-0.508 |
|
|
|
| 4 |
H |
0.081 |
|
|
|
| 5 |
H |
0.256 |
|
|
|
| 6 |
H |
0.252 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.776 |
0.909 |
0.000 |
3.883 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.675 |
1.109 |
-0.001 |
| y |
1.109 |
-15.033 |
-0.000 |
| z |
-0.001 |
-0.000 |
-18.661 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.828 |
1.109 |
-0.001 |
| y |
1.109 |
3.135 |
-0.000 |
| z |
-0.001 |
-0.000 |
-2.307 |
|
| Polar |
|---|
| 3z2-r2 | -4.614 |
|---|
| x2-y2 | -2.642 |
|---|
| xy | 1.109 |
|---|
| xz | -0.001 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.217 |
-0.137 |
0.000 |
| y |
-0.137 |
3.229 |
0.000 |
| z |
0.000 |
0.000 |
1.783 |
<r2> (average value of r
2) Å
2
| <r2> |
40.665 |
| (<r2>)1/2 |
6.377 |
