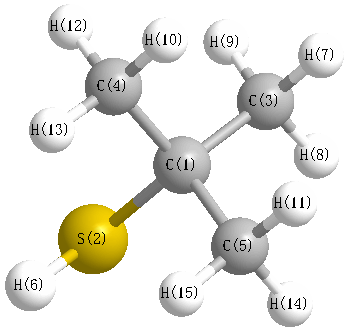
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3140 |
3013 |
24.04 |
|
|
|
| 2 |
A' |
3124 |
2998 |
26.76 |
|
|
|
| 3 |
A' |
3114 |
2987 |
40.36 |
|
|
|
| 4 |
A' |
3048 |
2924 |
21.27 |
|
|
|
| 5 |
A' |
3040 |
2916 |
32.35 |
|
|
|
| 6 |
A' |
2708 |
2598 |
6.16 |
|
|
|
| 7 |
A' |
1508 |
1447 |
9.04 |
|
|
|
| 8 |
A' |
1494 |
1433 |
11.49 |
|
|
|
| 9 |
A' |
1479 |
1419 |
0.63 |
|
|
|
| 10 |
A' |
1423 |
1365 |
1.60 |
|
|
|
| 11 |
A' |
1394 |
1338 |
15.16 |
|
|
|
| 12 |
A' |
1264 |
1213 |
0.37 |
|
|
|
| 13 |
A' |
1198 |
1150 |
47.52 |
|
|
|
| 14 |
A' |
1057 |
1014 |
2.61 |
|
|
|
| 15 |
A' |
948 |
910 |
0.66 |
|
|
|
| 16 |
A' |
882 |
846 |
4.79 |
|
|
|
| 17 |
A' |
840 |
806 |
1.38 |
|
|
|
| 18 |
A' |
599 |
575 |
5.16 |
|
|
|
| 19 |
A' |
387 |
372 |
1.12 |
|
|
|
| 20 |
A' |
365 |
350 |
0.30 |
|
|
|
| 21 |
A' |
295 |
283 |
0.30 |
|
|
|
| 22 |
A' |
281 |
270 |
0.71 |
|
|
|
| 23 |
A" |
3139 |
3011 |
22.96 |
|
|
|
| 24 |
A" |
3133 |
3006 |
5.62 |
|
|
|
| 25 |
A" |
3109 |
2983 |
0.65 |
|
|
|
| 26 |
A" |
3036 |
2913 |
17.12 |
|
|
|
| 27 |
A" |
1496 |
1435 |
9.08 |
|
|
|
| 28 |
A" |
1475 |
1415 |
0.40 |
|
|
|
| 29 |
A" |
1467 |
1407 |
0.05 |
|
|
|
| 30 |
A" |
1395 |
1338 |
14.18 |
|
|
|
| 31 |
A" |
1252 |
1201 |
3.37 |
|
|
|
| 32 |
A" |
1047 |
1005 |
0.35 |
|
|
|
| 33 |
A" |
962 |
923 |
0.01 |
|
|
|
| 34 |
A" |
941 |
903 |
0.44 |
|
|
|
| 35 |
A" |
396 |
380 |
0.34 |
|
|
|
| 36 |
A" |
302 |
290 |
2.93 |
|
|
|
| 37 |
A" |
279 |
267 |
0.07 |
|
|
|
| 38 |
A" |
251 |
241 |
8.66 |
|
|
|
| 39 |
A" |
211 |
202 |
8.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28739.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27572.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.459 |
|
|
|
| 2 |
S |
-0.030 |
|
|
|
| 3 |
C |
-0.293 |
|
|
|
| 4 |
C |
-0.253 |
|
|
|
| 5 |
C |
-0.253 |
|
|
|
| 6 |
H |
0.077 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.148 |
|
|
|
| 15 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.679 |
-0.844 |
0.000 |
1.879 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.738 |
2.351 |
0.000 |
| y |
2.351 |
-39.321 |
0.000 |
| z |
0.000 |
0.000 |
-42.450 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.853 |
2.351 |
0.000 |
| y |
2.351 |
3.273 |
0.000 |
| z |
0.000 |
0.000 |
-1.420 |
|
| Polar |
|---|
| 3z2-r2 | -2.841 |
|---|
| x2-y2 | -3.418 |
|---|
| xy | 2.351 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.937 |
0.050 |
0.000 |
| y |
0.050 |
9.387 |
0.000 |
| z |
0.000 |
0.000 |
8.747 |
<r2> (average value of r
2) Å
2
| <r2> |
164.624 |
| (<r2>)1/2 |
12.831 |