Jump to
S1C2
Energy calculated at wB97X-D/6-311G**
| | hartrees |
|---|
| Energy at 0K | -624.997852 |
| Energy at 298.15K | |
| HF Energy | -624.997852 |
| Nuclear repulsion energy | 192.574454 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3875 |
3706 |
109.81 |
|
|
|
| 2 |
A' |
1312 |
1255 |
141.37 |
|
|
|
| 3 |
A' |
1105 |
1057 |
153.55 |
|
|
|
| 4 |
A' |
773 |
739 |
110.53 |
|
|
|
| 5 |
A' |
505 |
483 |
9.96 |
|
|
|
| 6 |
A' |
370 |
354 |
0.87 |
|
|
|
| 7 |
A' |
215 |
206 |
135.14 |
|
|
|
| 8 |
A" |
3870 |
3702 |
65.18 |
|
|
|
| 9 |
A" |
1047 |
1001 |
161.07 |
|
|
|
| 10 |
A" |
749 |
716 |
252.16 |
|
|
|
| 11 |
A" |
427 |
408 |
1.52 |
|
|
|
| 12 |
A" |
214i |
204i |
89.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7017.0 cm
-1
Scaled (by 0.9565) Zero Point Vibrational Energy (zpe) 6711.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.212 |
0.368 |
0.000 |
| O2 |
-1.129 |
0.911 |
0.000 |
| O3 |
0.212 |
-0.719 |
1.253 |
| O4 |
0.212 |
-0.719 |
-1.253 |
| H5 |
1.121 |
-0.835 |
1.549 |
| H6 |
1.121 |
-0.835 |
-1.549 |
Atom - Atom Distances (Å)
| |
S1 |
O2 |
O3 |
O4 |
H5 |
H6 |
| S1 | | 1.4468 | 1.6591 | 1.6591 | 2.1618 | 2.1618 |
O2 | 1.4468 | | 2.4551 | 2.4551 | 3.2421 | 3.2421 | O3 | 1.6591 | 2.4551 | | 2.5060 | 0.9630 | 2.9482 | O4 | 1.6591 | 2.4551 | 2.5060 | | 2.9482 | 0.9630 | H5 | 2.1618 | 3.2421 | 0.9630 | 2.9482 | | 3.0983 | H6 | 2.1618 | 3.2421 | 2.9482 | 0.9630 | 3.0983 | |
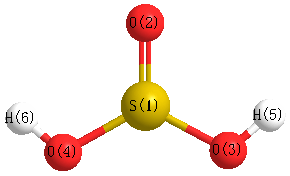 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
O3 |
H5 |
108.112 |
|
S1 |
O4 |
H6 |
108.112 |
| O2 |
S1 |
O3 |
104.248 |
|
O2 |
S1 |
O4 |
104.248 |
| O3 |
S1 |
O4 |
98.091 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.870 |
|
|
|
| 2 |
O |
-0.486 |
|
|
|
| 3 |
O |
-0.481 |
|
|
|
| 4 |
O |
-0.481 |
|
|
|
| 5 |
H |
0.288 |
|
|
|
| 6 |
H |
0.288 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.438 |
-0.694 |
0.000 |
5.482 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.233 |
-0.813 |
0.000 |
| y |
-0.813 |
-31.188 |
0.000 |
| z |
0.000 |
0.000 |
-27.211 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.967 |
-0.813 |
0.000 |
| y |
-0.813 |
-3.966 |
0.000 |
| z |
0.000 |
0.000 |
1.999 |
|
| Polar |
|---|
| 3z2-r2 | 3.998 |
|---|
| x2-y2 | 3.956 |
|---|
| xy | -0.813 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.082 |
-0.940 |
0.000 |
| y |
-0.940 |
3.776 |
0.000 |
| z |
0.000 |
0.000 |
4.404 |
<r2> (average value of r
2) Å
2
| <r2> |
79.985 |
| (<r2>)1/2 |
8.943 |
Jump to
S1C1
Energy calculated at wB97X-D/6-311G**
| | hartrees |
|---|
| Energy at 0K | -625.009009 |
| Energy at 298.15K | -625.012824 |
| HF Energy | -625.009009 |
| Nuclear repulsion energy | 193.052377 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3799 |
3633 |
119.10 |
|
|
|
| 2 |
A' |
1257 |
1202 |
184.02 |
|
|
|
| 3 |
A' |
1116 |
1067 |
11.33 |
|
|
|
| 4 |
A' |
784 |
750 |
173.90 |
|
|
|
| 5 |
A' |
486 |
465 |
20.78 |
|
|
|
| 6 |
A' |
442 |
423 |
161.44 |
|
|
|
| 7 |
A' |
325 |
311 |
31.71 |
|
|
|
| 8 |
A" |
3796 |
3631 |
36.64 |
|
|
|
| 9 |
A" |
1079 |
1033 |
64.64 |
|
|
|
| 10 |
A" |
772 |
738 |
371.27 |
|
|
|
| 11 |
A" |
444 |
425 |
72.54 |
|
|
|
| 12 |
A" |
164 |
157 |
29.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7231.0 cm
-1
Scaled (by 0.9565) Zero Point Vibrational Energy (zpe) 6916.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.292 |
0.329 |
0.000 |
| O2 |
-1.014 |
0.976 |
0.000 |
| O3 |
0.292 |
-0.702 |
1.256 |
| O4 |
0.292 |
-0.702 |
-1.256 |
| H5 |
-0.617 |
-0.921 |
1.501 |
| H6 |
-0.617 |
-0.921 |
-1.501 |
Atom - Atom Distances (Å)
| |
S1 |
O2 |
O3 |
O4 |
H5 |
H6 |
| S1 | | 1.4578 | 1.6254 | 1.6254 | 2.1545 | 2.1545 |
O2 | 1.4578 | | 2.4698 | 2.4698 | 2.4508 | 2.4508 | O3 | 1.6254 | 2.4698 | | 2.5130 | 0.9669 | 2.9115 | O4 | 1.6254 | 2.4698 | 2.5130 | | 2.9115 | 0.9669 | H5 | 2.1545 | 2.4508 | 0.9669 | 2.9115 | | 3.0012 | H6 | 2.1545 | 2.4508 | 2.9115 | 0.9669 | 3.0012 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
O3 |
H5 |
109.810 |
|
S1 |
O4 |
H6 |
109.810 |
| O2 |
S1 |
O3 |
106.336 |
|
O2 |
S1 |
O4 |
106.336 |
| O3 |
S1 |
O4 |
101.257 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.972 |
|
|
|
| 2 |
O |
-0.570 |
|
|
|
| 3 |
O |
-0.489 |
|
|
|
| 4 |
O |
-0.489 |
|
|
|
| 5 |
H |
0.288 |
|
|
|
| 6 |
H |
0.288 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.925 |
-1.436 |
0.000 |
1.708 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.542 |
6.104 |
0.000 |
| y |
6.104 |
-30.308 |
0.000 |
| z |
0.000 |
0.000 |
-28.757 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.009 |
6.104 |
0.000 |
| y |
6.104 |
-0.659 |
0.000 |
| z |
0.000 |
0.000 |
1.668 |
|
| Polar |
|---|
| 3z2-r2 | 3.335 |
|---|
| x2-y2 | -0.233 |
|---|
| xy | 6.104 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.830 |
-0.296 |
0.000 |
| y |
-0.296 |
3.841 |
0.000 |
| z |
0.000 |
0.000 |
4.261 |
<r2> (average value of r
2) Å
2
| <r2> |
79.368 |
| (<r2>)1/2 |
8.909 |
