return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: QCISD/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-311G**
 hartrees
Energy at 0K-871.843494
Energy at 298.15K-871.851343
HF Energy-871.459835
Nuclear repulsion energy192.650898
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2282 2178 139.60      
2 A1 2276 2171 32.29      
3 A1 2254 2150 83.73      
4 A1 985 940 82.59      
5 A1 964 920 30.08      
6 A1 930 887 214.42      
7 A1 595 567 11.05      
8 A1 397 378 0.94      
9 A1 102 98 2.02      
10 A2 2278 2173 0.00      
11 A2 975 930 0.00      
12 A2 736 702 0.00      
13 A2 446 426 0.00      
14 A2 82 78 0.00      
15 B1 2282 2178 267.72      
16 B1 2261 2157 32.57      
17 B1 978 934 99.65      
18 B1 625 596 13.76      
19 B1 339 323 32.22      
20 B1 103 99 0.14      
21 B2 2281 2176 98.90      
22 B2 2271 2166 90.42      
23 B2 976 931 37.05      
24 B2 924 881 369.30      
25 B2 761 726 352.35      
26 B2 480 458 15.74      
27 B2 460 439 19.38      

Unscaled Zero Point Vibrational Energy (zpe) 15020.5 cm-1
Scaled (by 0.9541) Zero Point Vibrational Energy (zpe) 14331.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-311G**
ABC
0.31290 0.06578 0.05759

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-311G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.889
Si2 0.000 1.945 -0.418
Si3 0.000 -1.945 -0.418
H4 1.199 0.000 1.762
H5 -1.199 0.000 1.762
H6 0.000 3.153 0.437
H7 0.000 -3.153 0.437
H8 1.201 1.976 -1.283
H9 -1.201 1.976 -1.283
H10 -1.201 -1.976 -1.283
H11 1.201 -1.976 -1.283

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.34372.34371.48331.48333.18553.18553.17223.17223.17223.1722
Si22.34373.89093.15863.15861.47995.16991.48051.48054.19124.1912
Si32.34373.89093.15863.15865.16991.47994.19124.19121.48051.4805
H41.48333.15863.15862.39763.62453.62453.63014.35184.35183.6301
H51.48333.15863.15862.39763.62453.62454.35183.63013.63014.3518
H63.18551.47995.16993.62453.62456.30652.40582.40585.54145.5414
H73.18555.16991.47993.62453.62456.30655.54145.54142.40582.4058
H83.17221.48054.19123.63014.35182.40585.54142.40264.62453.9514
H93.17221.48054.19124.35183.63012.40585.54142.40263.95144.6245
H103.17224.19121.48054.35183.63015.54142.40584.62453.95142.4026
H113.17224.19121.48053.63014.35185.54142.40583.95144.62452.4026

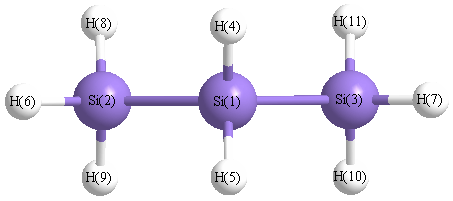
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 110.808 S1 S2 H8 110.041
S1 S2 H9 110.041 S1 S3 H7 110.808
S1 S3 H10 110.041 S1 S3 H11 110.041
S2 S1 S3 112.212 S2 S1 H4 109.173
S2 S1 H5 109.173 S3 S1 H4 109.173
S3 S1 H5 109.173 H4 S1 H5 107.839
H6 S2 H8 108.714 H6 S2 H9 108.714
H7 S3 H10 108.714 H7 S3 H11 108.714
H8 S2 H9 108.468 H10 S3 H11 108.468
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability