Jump to
S1C2
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -216.645475 |
| Energy at 298.15K | |
| HF Energy | -215.977531 |
| Nuclear repulsion energy | 116.302709 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3271 |
3271 |
6.85 |
|
|
|
| 2 |
A' |
3185 |
3185 |
6.53 |
|
|
|
| 3 |
A' |
3169 |
3169 |
10.06 |
|
|
|
| 4 |
A' |
3062 |
3062 |
39.76 |
|
|
|
| 5 |
A' |
1729 |
1729 |
2.14 |
|
|
|
| 6 |
A' |
1536 |
1536 |
2.41 |
|
|
|
| 7 |
A' |
1468 |
1468 |
11.94 |
|
|
|
| 8 |
A' |
1450 |
1450 |
11.24 |
|
|
|
| 9 |
A' |
1320 |
1320 |
0.32 |
|
|
|
| 10 |
A' |
1164 |
1164 |
53.44 |
|
|
|
| 11 |
A' |
1027 |
1027 |
28.31 |
|
|
|
| 12 |
A' |
927 |
927 |
2.37 |
|
|
|
| 13 |
A' |
615 |
615 |
6.75 |
|
|
|
| 14 |
A' |
277 |
277 |
2.69 |
|
|
|
| 15 |
A" |
3104 |
3104 |
35.41 |
|
|
|
| 16 |
A" |
1294 |
1294 |
0.31 |
|
|
|
| 17 |
A" |
1061 |
1061 |
16.89 |
|
|
|
| 18 |
A" |
1018 |
1018 |
13.90 |
|
|
|
| 19 |
A" |
936 |
936 |
36.10 |
|
|
|
| 20 |
A" |
558 |
558 |
8.26 |
|
|
|
| 21 |
A" |
169 |
169 |
3.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16169.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16169.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.949 |
-0.208 |
0.000 |
| C2 |
0.000 |
0.957 |
0.000 |
| C3 |
1.332 |
0.840 |
0.000 |
| F4 |
-0.269 |
-1.411 |
0.000 |
| H5 |
1.970 |
1.718 |
0.000 |
| H6 |
1.809 |
-0.135 |
0.000 |
| H7 |
-0.474 |
1.939 |
0.000 |
| H8 |
-1.589 |
-0.180 |
0.891 |
| H9 |
-1.589 |
-0.180 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5027 | 2.5098 | 1.3821 | 3.4975 | 2.7587 | 2.1986 | 1.0972 | 1.0972 |
C2 | 1.5027 | | 1.3366 | 2.3835 | 2.1122 | 2.1127 | 1.0900 | 2.1476 | 2.1476 | C3 | 2.5098 | 1.3366 | | 2.7623 | 1.0858 | 1.0853 | 2.1135 | 3.2195 | 3.2195 | F4 | 1.3821 | 2.3835 | 2.7623 | | 3.8480 | 2.4385 | 3.3561 | 2.0130 | 2.0130 | H5 | 3.4975 | 2.1122 | 1.0858 | 3.8480 | | 1.8598 | 2.4543 | 4.1313 | 4.1313 | H6 | 2.7587 | 2.1127 | 1.0853 | 2.4385 | 1.8598 | | 3.0838 | 3.5131 | 3.5131 | H7 | 2.1986 | 1.0900 | 2.1135 | 3.3561 | 2.4543 | 3.0838 | | 2.5546 | 2.5546 | H8 | 1.0972 | 2.1476 | 3.2195 | 2.0130 | 4.1313 | 3.5131 | 2.5546 | | 1.7810 | H9 | 1.0972 | 2.1476 | 3.2195 | 2.0130 | 4.1313 | 3.5131 | 2.5546 | 1.7810 | |
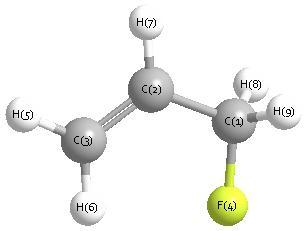 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.145 |
|
C1 |
C2 |
H7 |
115.060 |
| C2 |
C1 |
F4 |
111.362 |
|
C2 |
C1 |
H8 |
110.416 |
| C2 |
C1 |
H9 |
110.416 |
|
C2 |
C3 |
H5 |
121.020 |
| C2 |
C3 |
H6 |
121.105 |
|
C3 |
C2 |
H7 |
120.795 |
| F4 |
C1 |
H8 |
108.020 |
|
F4 |
C1 |
H9 |
108.020 |
| H5 |
C3 |
H6 |
117.876 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-311G**
| | hartrees |
|---|
| Energy at 0K | -216.644531 |
| Energy at 298.15K | |
| HF Energy | -215.976746 |
| Nuclear repulsion energy | 113.964362 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3252 |
3252 |
14.16 |
|
|
|
| 2 |
A |
3187 |
3187 |
6.75 |
|
|
|
| 3 |
A |
3161 |
3161 |
10.13 |
|
|
|
| 4 |
A |
3126 |
3126 |
28.60 |
|
|
|
| 5 |
A |
3072 |
3072 |
37.48 |
|
|
|
| 6 |
A |
1726 |
1726 |
0.36 |
|
|
|
| 7 |
A |
1533 |
1533 |
0.78 |
|
|
|
| 8 |
A |
1486 |
1486 |
26.37 |
|
|
|
| 9 |
A |
1428 |
1428 |
14.12 |
|
|
|
| 10 |
A |
1318 |
1318 |
0.14 |
|
|
|
| 11 |
A |
1300 |
1300 |
3.16 |
|
|
|
| 12 |
A |
1197 |
1197 |
3.45 |
|
|
|
| 13 |
A |
1100 |
1100 |
110.09 |
|
|
|
| 14 |
A |
1024 |
1024 |
33.09 |
|
|
|
| 15 |
A |
999 |
999 |
3.62 |
|
|
|
| 16 |
A |
945 |
945 |
34.74 |
|
|
|
| 17 |
A |
936 |
936 |
1.99 |
|
|
|
| 18 |
A |
651 |
651 |
6.60 |
|
|
|
| 19 |
A |
438 |
438 |
2.29 |
|
|
|
| 20 |
A |
334 |
334 |
6.44 |
|
|
|
| 21 |
A |
112 |
112 |
1.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16162.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16162.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-311G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.612 |
0.436 |
0.309 |
| C2 |
0.643 |
-0.384 |
0.240 |
| C3 |
1.810 |
0.092 |
-0.208 |
| F4 |
-1.648 |
-0.227 |
-0.336 |
| H5 |
2.709 |
-0.518 |
-0.217 |
| H6 |
1.902 |
1.111 |
-0.579 |
| H7 |
0.561 |
-1.408 |
0.600 |
| H8 |
-0.922 |
0.591 |
1.349 |
| H9 |
-0.471 |
1.409 |
-0.175 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5011 | 2.5007 | 1.3885 | 3.4950 | 2.7503 | 2.2050 | 1.0963 | 1.0952 |
C2 | 1.5011 | | 1.3378 | 2.3676 | 2.1197 | 2.1191 | 1.0890 | 2.1515 | 2.1511 | C3 | 2.5007 | 1.3378 | | 3.4752 | 1.0861 | 1.0882 | 2.1136 | 3.1841 | 2.6340 | F4 | 1.3885 | 2.3676 | 3.4752 | | 4.3679 | 3.8016 | 2.6735 | 2.0083 | 2.0218 | H5 | 3.4950 | 2.1197 | 1.0861 | 4.3679 | | 1.8537 | 2.4649 | 4.1066 | 3.7181 | H6 | 2.7503 | 2.1191 | 1.0882 | 3.8016 | 1.8537 | | 3.0884 | 3.4585 | 2.4253 | H7 | 2.2050 | 1.0890 | 2.1136 | 2.6735 | 2.4649 | 3.0884 | | 2.5988 | 3.0984 | H8 | 1.0963 | 2.1515 | 3.1841 | 2.0083 | 4.1066 | 3.4585 | 2.5988 | | 1.7872 | H9 | 1.0952 | 2.1511 | 2.6340 | 2.0218 | 3.7181 | 2.4253 | 3.0984 | 1.7872 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.393 |
|
C1 |
C2 |
H7 |
115.798 |
| C2 |
C1 |
F4 |
109.979 |
|
C2 |
C1 |
H8 |
110.896 |
| C2 |
C1 |
H9 |
110.924 |
|
C2 |
C3 |
H5 |
121.625 |
| C2 |
C3 |
H6 |
121.398 |
|
C3 |
C2 |
H7 |
120.800 |
| F4 |
C1 |
H8 |
107.262 |
|
F4 |
C1 |
H9 |
108.401 |
| H5 |
C3 |
H6 |
116.977 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
