return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(CH3)CONH(CH3) (Urea, N,N'-dimethyl-)

using model chemistry: B2PLYP=FULLultrafine/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B2PLYP=FULLultrafine/6-311G**
 hartrees
Energy at 0K-303.701723
Energy at 298.15K 
HF Energy-303.342856
Nuclear repulsion energy248.182089
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3631 3631 18.13      
2 A 3177 3177 10.31      
3 A 3121 3121 9.36      
4 A 3040 3040 4.14      
5 A 1785 1785 235.33      
6 A 1544 1544 18.85      
7 A 1511 1511 0.27      
8 A 1492 1492 5.26      
9 A 1461 1461 9.22      
10 A 1207 1207 0.02      
11 A 1185 1185 1.01      
12 A 1154 1154 8.24      
13 A 930 930 0.93      
14 A 551 551 88.37      
15 A 445 445 42.02      
16 A 229 229 2.86      
17 A 191 191 0.14      
18 A 97 97 0.46      
19 B 3627 3627 9.45      
20 B 3177 3177 4.57      
21 B 3122 3122 59.81      
22 B 3039 3039 125.94      
23 B 1577 1577 309.32      
24 B 1525 1525 11.70      
25 B 1503 1503 86.09      
26 B 1465 1465 7.05      
27 B 1267 1267 246.27      
28 B 1166 1166 30.83      
29 B 1154 1154 7.29      
30 B 1052 1052 1.11      
31 B 769 769 26.48      
32 B 739 739 21.12      
33 B 508 508 108.33      
34 B 329 329 44.83      
35 B 124 124 8.10      
36 B 103 103 0.57      

Unscaled Zero Point Vibrational Energy (zpe) 26498.6 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 26498.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/6-311G**
ABC
0.32667 0.07237 0.06082

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.099
O2 0.000 0.000 1.319
N3 0.000 1.158 -0.663
N4 0.000 -1.158 -0.663
C5 -0.241 2.425 0.012
C6 0.241 -2.425 0.012
H7 -0.412 1.068 -1.578
H8 0.412 -1.068 -1.578
H9 0.076 3.241 -0.638
H10 -0.076 -3.241 -0.638
H11 0.350 2.446 0.925
H12 -0.350 -2.446 0.925
H13 -1.292 2.569 0.286
H14 1.292 -2.569 0.286

Atom - Atom Distances (Å)
  C1 O2 N3 N4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.21951.38641.38642.43892.43892.03112.03113.32483.32482.60542.60542.88162.8816
O21.21952.29512.29512.76562.76563.11523.11523.78693.78692.50252.50253.05563.0556
N31.38642.29512.31641.45553.65441.00842.44212.08434.39992.07383.95402.13494.0572
N41.38642.29512.31643.65441.45552.44211.00844.39992.08433.95402.07384.05722.1349
C52.43892.76561.45553.65444.87462.09793.89331.09025.70591.08734.95771.09535.2315
C62.43892.76563.65441.45554.87463.89332.09795.70591.09024.95771.08735.23151.0953
H72.03113.11521.00842.44212.09793.89332.28932.41774.42292.95744.31482.54984.4278
H82.03113.11522.44211.00843.89332.09792.28934.42292.41774.31482.95744.42782.5498
H93.32483.78692.08434.39991.09025.70592.41774.42296.48381.77465.91361.78216.0074
H103.32483.78694.39992.08435.70591.09024.42292.41776.48385.91361.77466.00741.7821
H112.60542.50252.07383.95401.08734.95772.95744.31481.77465.91364.94251.76575.1431
H122.60542.50253.95402.07384.95771.08734.31482.95745.91361.77464.94255.14311.7657
H132.88163.05562.13494.05721.09535.23152.54984.42781.78216.00741.76575.14315.7511
H142.88163.05564.05722.13495.23151.09534.42782.54986.00741.78215.14311.76575.7511

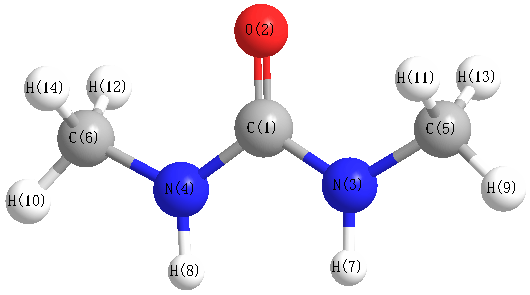
picture of Urea, N,N'-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C5 118.207 C1 N3 H7 115.103
C1 N4 C6 118.207 C1 N4 H8 115.103
O2 C1 N3 123.341 O2 C1 N4 123.341
N3 C1 N4 113.318 N3 C5 H9 109.079
N3 C5 H11 108.416 N3 C5 H13 112.878
N4 C6 H10 109.079 N4 C6 H12 108.416
N4 C6 H14 112.878 C5 N3 H7 115.546
C6 N4 H8 115.546 H9 C5 H11 109.161
H9 C5 H13 109.256 H10 C6 H12 109.161
H10 C6 H14 109.256 H11 C5 H13 107.987
H12 C6 H14 107.987
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability