return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O2 (γ–Butyrolactone)

using model chemistry: CCSD/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCSD/6-311G**
 hartrees
Energy at 0K-305.773088
Energy at 298.15K 
HF Energy-304.778707
Nuclear repulsion energy240.980721
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3169 3169 11.26      
2 A 3158 3158 27.85      
3 A 3147 3147 15.65      
4 A 3089 3089 21.51      
5 A 3084 3084 5.84      
6 A 3072 3072 41.36      
7 A 1914 1914 378.53      
8 A 1555 1555 0.61      
9 A 1523 1523 6.49      
10 A 1492 1492 7.83      
11 A 1434 1434 19.66      
12 A 1375 1375 3.00      
13 A 1337 1337 18.38      
14 A 1290 1290 14.77      
15 A 1243 1243 28.79      
16 A 1223 1223 2.57      
17 A 1210 1210 197.02      
18 A 1119 1119 38.17      
19 A 1108 1108 43.76      
20 A 1027 1027 12.87      
21 A 959 959 3.75      
22 A 916 916 11.81      
23 A 897 897 15.39      
24 A 824 824 5.37      
25 A 691 691 5.72      
26 A 644 644 3.31      
27 A 537 537 3.85      
28 A 499 499 4.34      
29 A 229 229 2.82      
30 A 152 152 0.28      

Unscaled Zero Point Vibrational Energy (zpe) 21958.4 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21958.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD/6-311G**
ABC
0.24285 0.11998 0.08586

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD/6-311G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.884 -0.004 0.005
C2 -0.031 1.204 0.186
C3 -1.395 0.664 -0.246
C4 -1.260 -0.813 0.146
O5 0.125 -1.134 -0.054
O6 2.076 -0.024 -0.076
H7 0.344 2.052 -0.390
H8 -0.021 1.475 1.249
H9 -1.510 0.751 -1.332
H10 -2.245 1.150 0.239
H11 -1.851 -1.494 -0.469
H12 -1.509 -0.971 1.203

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12
C11.52682.38842.29621.36221.19462.16222.13422.84443.34383.15022.8455
C21.52681.52972.36292.35582.45291.09121.09722.16742.21563.31992.8199
C32.38841.52971.53412.36203.54252.23022.18721.09551.09312.21642.1877
C42.29622.36291.53411.43603.43533.32742.82632.16702.19841.09141.0971
O51.36222.35582.36201.43602.24443.21102.91972.80373.30472.05092.0680
O61.19462.45293.54253.43532.24442.72162.89803.87784.48894.21103.9221
H72.16221.09122.23023.32743.21102.72161.77562.45332.81344.17113.8872
H82.13421.09722.18722.82632.91972.89801.77563.06692.46493.88802.8634
H92.84442.16741.09552.16702.80373.87782.45333.06691.77992.42923.0650
H103.34382.21561.09312.19843.30474.48892.81342.46491.77992.76552.4436
H113.15023.31992.21641.09142.05094.21104.17113.88802.42922.76551.7854
H122.84552.81992.18771.09712.06803.92213.88722.86343.06502.44361.7854

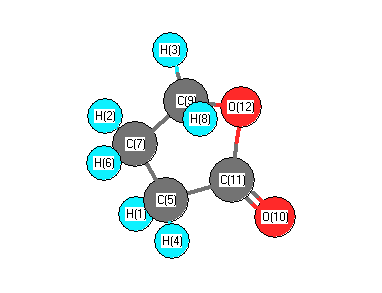
picture of γ–Butyrolactone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 102.777 C1 C2 H7 110.254
C1 C2 H8 107.716 C1 O5 C4 110.263
C2 C1 O5 109.123 C2 C1 O6 128.254
C2 C3 C4 100.929 C2 C3 H9 110.202
C2 C3 H10 114.263 C3 C2 H7 115.613
C3 C2 H8 111.674 C3 C4 O5 105.315
C3 C4 H11 114.110 C3 C4 H12 111.417
C4 C3 H9 109.868 C4 C3 H10 112.529
O5 C1 O6 122.623 O5 C4 H11 107.695
O5 C4 H12 108.716 H7 C2 H8 108.458
H9 C3 H10 108.836 H11 C4 H12 109.332
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability