Jump to
S1C2
Energy calculated at B3LYPultrafine/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -399.871273 |
| Energy at 298.15K | -399.872358 |
| HF Energy | -399.871273 |
| Nuclear repulsion energy | 62.949745 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3LYPultrafine/cc-pCVTZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
1.988 |
| Na3 |
0.000 |
0.000 |
-1.988 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.9881 | 1.9881 |
Na2 | 1.9881 | | 3.9762 | Na3 | 1.9881 | 3.9762 | |
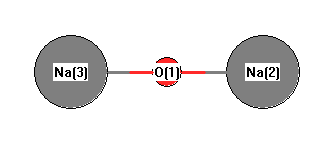 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.828 |
|
|
|
| 2 |
Na |
0.414 |
|
|
|
| 3 |
Na |
0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.277 |
0.000 |
0.000 |
| y |
0.000 |
-20.277 |
0.000 |
| z |
0.000 |
0.000 |
11.697 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.987 |
0.000 |
0.000 |
| y |
0.000 |
-15.987 |
0.000 |
| z |
0.000 |
0.000 |
31.974 |
|
| Polar |
|---|
| 3z2-r2 | 63.948 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.921 |
0.000 |
0.000 |
| y |
0.000 |
10.921 |
0.000 |
| z |
0.000 |
0.000 |
22.364 |
<r2> (average value of r
2) Å
2
| <r2> |
92.964 |
| (<r2>)1/2 |
9.642 |
Jump to
S1C1
Energy calculated at B3LYPultrafine/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -399.871273 |
| Energy at 298.15K | -399.872707 |
| HF Energy | -399.871273 |
| Nuclear repulsion energy | 62.957181 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3LYPultrafine/cc-pCVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.001 |
| Na2 |
0.000 |
1.988 |
-0.000 |
| Na3 |
0.000 |
-1.988 |
-0.000 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.9879 | 1.9879 |
Na2 | 1.9879 | | 3.9757 | Na3 | 1.9879 | 3.9757 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
179.915 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.828 |
|
|
|
| 2 |
Na |
0.414 |
|
|
|
| 3 |
Na |
0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.008 |
0.008 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.276 |
0.000 |
0.000 |
| y |
0.000 |
11.691 |
0.000 |
| z |
0.000 |
0.000 |
-20.276 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.983 |
0.000 |
0.000 |
| y |
0.000 |
31.967 |
0.000 |
| z |
0.000 |
0.000 |
-15.983 |
|
| Polar |
|---|
| 3z2-r2 | -31.967 |
|---|
| x2-y2 | -31.967 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.918 |
0.000 |
0.000 |
| y |
0.000 |
22.357 |
0.000 |
| z |
0.000 |
0.000 |
10.918 |
<r2> (average value of r
2) Å
2
| <r2> |
92.944 |
| (<r2>)1/2 |
9.641 |
