Jump to
S1C2
S1C3
Energy calculated at LSDA/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1588.719714 |
| Energy at 298.15K | |
| HF Energy | -1588.719714 |
| Nuclear repulsion energy | 354.423519 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at LSDA/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
532 |
532 |
0.00 |
|
|
|
| 2 |
A1 |
219 |
219 |
0.00 |
|
|
|
| 3 |
B1 |
441 |
441 |
0.00 |
|
|
|
| 4 |
B2 |
285 |
285 |
0.00 |
|
|
|
| 5 |
E |
419 |
419 |
0.12 |
|
|
|
| 5 |
E |
419 |
419 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1156.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1156.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at LSDA/cc-pCVTZ
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.409 |
0.329 |
| S2 |
0.000 |
-1.409 |
0.329 |
| S3 |
-1.409 |
0.000 |
-0.329 |
| S4 |
1.409 |
0.000 |
-0.329 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8176 | 2.0981 | 2.0981 |
S2 | 2.8176 | | 2.0981 | 2.0981 | S3 | 2.0981 | 2.0981 | | 2.8176 | S4 | 2.0981 | 2.0981 | 2.8176 | |
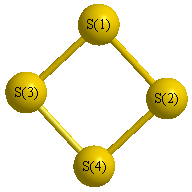 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.821 |
|
S1 |
S3 |
S4 |
47.821 |
| S2 |
S1 |
S3 |
47.821 |
|
S2 |
S4 |
S3 |
47.821 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.766 |
0.000 |
0.000 |
| y |
0.000 |
-48.766 |
0.000 |
| z |
0.000 |
0.000 |
-53.884 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.559 |
0.000 |
0.000 |
| y |
0.000 |
2.559 |
0.000 |
| z |
0.000 |
0.000 |
-5.118 |
|
| Polar |
|---|
| 3z2-r2 | -10.236 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.350 |
0.000 |
0.000 |
| y |
0.000 |
11.350 |
0.000 |
| z |
0.000 |
0.000 |
6.888 |
<r2> (average value of r
2) Å
2
| <r2> |
165.471 |
| (<r2>)1/2 |
12.864 |
Jump to
S1C1
S1C3
Energy calculated at LSDA/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1588.761413 |
| Energy at 298.15K | |
| HF Energy | -1588.761413 |
| Nuclear repulsion energy | 338.220036 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at LSDA/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
717 |
717 |
2.58 |
|
|
|
| 2 |
A1 |
328 |
328 |
4.02 |
|
|
|
| 3 |
A1 |
98 |
98 |
4.58 |
|
|
|
| 4 |
A2 |
241 |
241 |
0.00 |
|
|
|
| 5 |
B2 |
681 |
681 |
82.77 |
|
|
|
| 6 |
B2 |
340 |
340 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1202.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1202.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at LSDA/cc-pCVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.103 |
0.932 |
| S2 |
0.000 |
-1.103 |
0.932 |
| S3 |
0.000 |
1.428 |
-0.932 |
| S4 |
0.000 |
-1.428 |
-0.932 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.2063 | 1.8922 | 3.1434 |
S2 | 2.2063 | | 3.1434 | 1.8922 | S3 | 1.8922 | 3.1434 | | 2.8558 | S4 | 3.1434 | 1.8922 | 2.8558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
99.883 |
|
S1 |
S3 |
S4 |
80.117 |
| S2 |
S1 |
S3 |
99.883 |
|
S2 |
S4 |
S3 |
80.117 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.079 |
|
|
|
| 2 |
S |
0.079 |
|
|
|
| 3 |
S |
-0.079 |
|
|
|
| 4 |
S |
-0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.886 |
0.886 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.167 |
0.000 |
0.000 |
| y |
0.000 |
-52.773 |
0.000 |
| z |
0.000 |
0.000 |
-47.885 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.839 |
0.000 |
0.000 |
| y |
0.000 |
-3.246 |
0.000 |
| z |
0.000 |
0.000 |
4.085 |
|
| Polar |
|---|
| 3z2-r2 | 8.169 |
|---|
| x2-y2 | 1.605 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.911 |
0.000 |
0.000 |
| y |
0.000 |
16.872 |
0.000 |
| z |
0.000 |
0.000 |
12.218 |
<r2> (average value of r
2) Å
2
| <r2> |
191.393 |
| (<r2>)1/2 |
13.834 |
Jump to
S1C1
S1C2
Energy calculated at LSDA/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1588.760664 |
| Energy at 298.15K | |
| HF Energy | -1588.760664 |
| Nuclear repulsion energy | 339.640568 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at LSDA/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
736 |
736 |
0.00 |
|
|
|
| 2 |
Ag |
306 |
306 |
0.00 |
|
|
|
| 3 |
Au |
256 |
256 |
0.00 |
|
|
|
| 4 |
B1u |
687 |
687 |
83.15 |
|
|
|
| 5 |
B2u |
84i |
84i |
12.47 |
|
|
|
| 6 |
B3g |
343 |
343 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1122.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1122.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at LSDA/cc-pCVTZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.943 |
1.242 |
| S2 |
0.000 |
0.943 |
-1.242 |
| S3 |
0.000 |
-0.943 |
1.242 |
| S4 |
0.000 |
-0.943 |
-1.242 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.4835 | 1.8859 | 3.1184 |
S2 | 2.4835 | | 3.1184 | 1.8859 | S3 | 1.8859 | 3.1184 | | 2.4835 | S4 | 3.1184 | 1.8859 | 2.4835 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.305 |
0.000 |
0.000 |
| y |
0.000 |
-47.616 |
0.000 |
| z |
0.000 |
0.000 |
-52.694 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.150 |
0.000 |
0.000 |
| y |
0.000 |
4.383 |
0.000 |
| z |
0.000 |
0.000 |
-3.233 |
|
| Polar |
|---|
| 3z2-r2 | -6.466 |
|---|
| x2-y2 | -3.689 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.904 |
0.000 |
0.000 |
| y |
0.000 |
12.398 |
0.000 |
| z |
0.000 |
0.000 |
16.576 |
<r2> (average value of r
2) Å
2
| <r2> |
187.159 |
| (<r2>)1/2 |
13.681 |
