Jump to
S1C2
S1C3
Energy calculated at B3LYPultrafine/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.886562 |
| Energy at 298.15K | |
| HF Energy | -1592.886562 |
| Nuclear repulsion energy | 348.327620 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
508 |
508 |
0.00 |
|
|
|
| 2 |
A1 |
197 |
197 |
0.00 |
|
|
|
| 3 |
B1 |
445 |
445 |
0.00 |
|
|
|
| 4 |
B2 |
299 |
299 |
0.00 |
|
|
|
| 5 |
E |
417 |
417 |
0.03 |
|
|
|
| 5 |
E |
417 |
417 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1141.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1141.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/cc-pCVTZ
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.441 |
0.310 |
| S2 |
0.000 |
-1.441 |
0.310 |
| S3 |
-1.441 |
0.000 |
-0.310 |
| S4 |
1.441 |
0.000 |
-0.310 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8826 | 2.1306 | 2.1306 |
S2 | 2.8826 | | 2.1306 | 2.1306 | S3 | 2.1306 | 2.1306 | | 2.8826 | S4 | 2.1306 | 2.1306 | 2.8826 | |
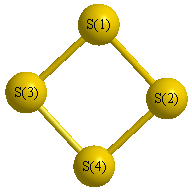 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.431 |
|
S1 |
S3 |
S4 |
47.431 |
| S2 |
S1 |
S3 |
47.431 |
|
S2 |
S4 |
S3 |
47.431 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.817 |
0.000 |
0.000 |
| y |
0.000 |
-48.817 |
0.000 |
| z |
0.000 |
0.000 |
-54.321 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.752 |
0.000 |
0.000 |
| y |
0.000 |
2.752 |
0.000 |
| z |
0.000 |
0.000 |
-5.503 |
|
| Polar |
|---|
| 3z2-r2 | -11.007 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.584 |
0.000 |
0.000 |
| y |
0.000 |
11.584 |
0.000 |
| z |
0.000 |
0.000 |
6.763 |
<r2> (average value of r
2) Å
2
| <r2> |
170.738 |
| (<r2>)1/2 |
13.067 |
Jump to
S1C1
S1C3
Energy calculated at B3LYPultrafine/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.916833 |
| Energy at 298.15K | |
| HF Energy | -1592.916833 |
| Nuclear repulsion energy | 330.028723 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
675 |
675 |
6.21 |
|
|
|
| 2 |
A1 |
372 |
372 |
4.89 |
|
|
|
| 3 |
A1 |
107 |
107 |
0.24 |
|
|
|
| 4 |
A2 |
210 |
210 |
0.00 |
|
|
|
| 5 |
B2 |
648 |
648 |
118.62 |
|
|
|
| 6 |
B2 |
333 |
333 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1172.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1172.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/cc-pCVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.058 |
0.912 |
| S2 |
0.000 |
-1.058 |
0.912 |
| S3 |
0.000 |
1.639 |
-0.912 |
| S4 |
0.000 |
-1.639 |
-0.912 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.1170 | 1.9146 | 3.2566 |
S2 | 2.1170 | | 3.2566 | 1.9146 | S3 | 1.9146 | 3.2566 | | 3.2780 | S4 | 3.2566 | 1.9146 | 3.2780 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
107.650 |
|
S1 |
S3 |
S4 |
72.350 |
| S2 |
S1 |
S3 |
107.650 |
|
S2 |
S4 |
S3 |
72.350 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.129 |
|
|
|
| 2 |
S |
0.129 |
|
|
|
| 3 |
S |
-0.129 |
|
|
|
| 4 |
S |
-0.129 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.397 |
1.397 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.196 |
0.000 |
0.000 |
| y |
0.000 |
-53.185 |
0.000 |
| z |
0.000 |
0.000 |
-48.667 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.270 |
0.000 |
0.000 |
| y |
0.000 |
-3.254 |
0.000 |
| z |
0.000 |
0.000 |
3.524 |
|
| Polar |
|---|
| 3z2-r2 | 7.047 |
|---|
| x2-y2 | 1.989 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.940 |
0.000 |
0.000 |
| y |
0.000 |
18.757 |
0.000 |
| z |
0.000 |
0.000 |
11.932 |
<r2> (average value of r
2) Å
2
| <r2> |
206.938 |
| (<r2>)1/2 |
14.385 |
Jump to
S1C1
S1C2
Energy calculated at B3LYPultrafine/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -1592.910657 |
| Energy at 298.15K | |
| HF Energy | -1592.910657 |
| Nuclear repulsion energy | 333.985122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
725 |
725 |
0.00 |
|
|
|
| 2 |
Ag |
283 |
283 |
0.00 |
|
|
|
| 3 |
Au |
246 |
246 |
0.00 |
|
|
|
| 4 |
B1u |
668 |
668 |
122.71 |
|
|
|
| 5 |
B2u |
146i |
146i |
15.29 |
|
|
|
| 6 |
B3g |
336 |
336 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1055.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1055.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/cc-pCVTZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.948 |
1.278 |
| S2 |
0.000 |
0.948 |
-1.278 |
| S3 |
0.000 |
-0.948 |
1.278 |
| S4 |
0.000 |
-0.948 |
-1.278 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5559 | 1.8966 | 3.1827 |
S2 | 2.5559 | | 3.1827 | 1.8966 | S3 | 1.8966 | 3.1827 | | 2.5559 | S4 | 3.1827 | 1.8966 | 2.5559 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.543 |
0.000 |
0.000 |
| y |
0.000 |
-47.897 |
0.000 |
| z |
0.000 |
0.000 |
-52.754 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.218 |
0.000 |
0.000 |
| y |
0.000 |
4.251 |
0.000 |
| z |
0.000 |
0.000 |
-3.034 |
|
| Polar |
|---|
| 3z2-r2 | -6.067 |
|---|
| x2-y2 | -3.646 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.926 |
0.000 |
0.000 |
| y |
0.000 |
12.496 |
0.000 |
| z |
0.000 |
0.000 |
17.625 |
<r2> (average value of r
2) Å
2
| <r2> |
193.763 |
| (<r2>)1/2 |
13.920 |
