Jump to
S1C2
Energy calculated at QCISD(T)=FULL/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -100.322674 |
| Energy at 298.15K | -100.321731 |
| HF Energy | -99.809295 |
| Nuclear repulsion energy | 27.547422 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)=FULL/cc-pCVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.070 |
| C2 |
0.000 |
0.000 |
-0.153 |
| N3 |
0.000 |
0.000 |
1.018 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9173 | 3.0879 |
C2 | 1.9173 | | 1.1706 | N3 | 3.0879 | 1.1706 | |
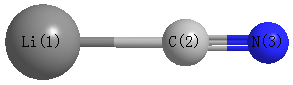 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -100.326828 |
| Energy at 298.15K | -100.326088 |
| HF Energy | -99.815814 |
| Nuclear repulsion energy | 29.259760 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)=FULL/cc-pCVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.410 |
-0.621 |
0.000 |
| C2 |
-0.705 |
-0.368 |
0.000 |
| N3 |
0.000 |
0.582 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1296 | 1.8530 |
C2 | 2.1296 | | 1.1830 | N3 | 1.8530 | 1.1830 | |
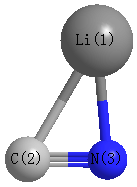 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
60.238 |
|
Li1 |
N3 |
C2 |
86.105 |
| C2 |
Li1 |
N3 |
33.657 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
