Jump to
S1C2
Energy calculated at wB97X-D/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -478.041586 |
| Energy at 298.15K | |
| HF Energy | -478.041586 |
| Nuclear repulsion energy | 107.701789 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
3136 |
23.29 |
|
|
|
| 2 |
A' |
3084 |
3084 |
20.68 |
|
|
|
| 3 |
A' |
3057 |
3057 |
20.01 |
|
|
|
| 4 |
A' |
2758 |
2758 |
4.01 |
|
|
|
| 5 |
A' |
1512 |
1512 |
2.94 |
|
|
|
| 6 |
A' |
1499 |
1499 |
3.06 |
|
|
|
| 7 |
A' |
1426 |
1426 |
4.24 |
|
|
|
| 8 |
A' |
1305 |
1305 |
27.35 |
|
|
|
| 9 |
A' |
1121 |
1121 |
1.75 |
|
|
|
| 10 |
A' |
1007 |
1007 |
3.46 |
|
|
|
| 11 |
A' |
867 |
867 |
1.08 |
|
|
|
| 12 |
A' |
690 |
690 |
1.01 |
|
|
|
| 13 |
A' |
308 |
308 |
2.09 |
|
|
|
| 14 |
A" |
3146 |
3146 |
25.94 |
|
|
|
| 15 |
A" |
3123 |
3123 |
1.46 |
|
|
|
| 16 |
A" |
1495 |
1495 |
9.40 |
|
|
|
| 17 |
A" |
1273 |
1273 |
0.50 |
|
|
|
| 18 |
A" |
1051 |
1051 |
0.28 |
|
|
|
| 19 |
A" |
798 |
798 |
3.47 |
|
|
|
| 20 |
A" |
230 |
230 |
2.63 |
|
|
|
| 21 |
A" |
173 |
173 |
14.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16528.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16528.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/cc-pCVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.514 |
0.684 |
0.000 |
| C2 |
0.000 |
0.827 |
0.000 |
| S3 |
-0.755 |
-0.833 |
0.000 |
| H4 |
1.982 |
1.668 |
0.000 |
| H5 |
1.858 |
0.145 |
0.883 |
| H6 |
1.858 |
0.145 |
-0.883 |
| H7 |
-0.332 |
1.367 |
0.884 |
| H8 |
-0.332 |
1.367 |
-0.884 |
| H9 |
-2.035 |
-0.438 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5207 | 2.7295 | 1.0897 | 1.0899 | 1.0899 | 2.1576 | 2.1576 | 3.7220 |
C2 | 1.5207 | | 1.8231 | 2.1536 | 2.1666 | 2.1666 | 1.0881 | 1.0881 | 2.3958 | S3 | 2.7295 | 1.8231 | | 3.7079 | 2.9263 | 2.9263 | 2.4085 | 2.4085 | 1.3390 | H4 | 1.0897 | 2.1536 | 3.7079 | | 1.7646 | 1.7646 | 2.4954 | 2.4954 | 4.5358 | H5 | 1.0899 | 2.1666 | 2.9263 | 1.7646 | | 1.7659 | 2.5072 | 3.0673 | 4.0336 | H6 | 1.0899 | 2.1666 | 2.9263 | 1.7646 | 1.7659 | | 3.0673 | 2.5072 | 4.0336 | H7 | 2.1576 | 1.0881 | 2.4085 | 2.4954 | 2.5072 | 3.0673 | | 1.7681 | 2.6346 | H8 | 2.1576 | 1.0881 | 2.4085 | 2.4954 | 3.0673 | 2.5072 | 1.7681 | | 2.6346 | H9 | 3.7220 | 2.3958 | 1.3390 | 4.5358 | 4.0336 | 4.0336 | 2.6346 | 2.6346 | |
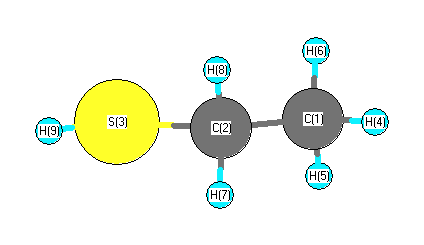 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.095 |
|
C1 |
C2 |
H7 |
110.494 |
| C1 |
C2 |
H8 |
110.494 |
|
C2 |
C1 |
H4 |
110.078 |
| C2 |
C1 |
H5 |
111.110 |
|
C2 |
C1 |
H6 |
111.110 |
| C2 |
S3 |
H9 |
97.334 |
|
S3 |
C2 |
H7 |
109.021 |
| S3 |
C2 |
H8 |
109.021 |
|
H4 |
C1 |
H5 |
108.111 |
| H4 |
C1 |
H6 |
108.111 |
|
H5 |
C1 |
H6 |
108.212 |
| H7 |
C2 |
H8 |
108.684 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.285 |
|
|
|
| 2 |
C |
-0.180 |
|
|
|
| 3 |
S |
-0.187 |
|
|
|
| 4 |
H |
0.099 |
|
|
|
| 5 |
H |
0.106 |
|
|
|
| 6 |
H |
0.106 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.047 |
1.671 |
0.000 |
1.671 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.786 |
-0.227 |
0.000 |
| y |
-0.227 |
-28.025 |
0.000 |
| z |
0.000 |
0.000 |
-28.700 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.577 |
-0.227 |
0.000 |
| y |
-0.227 |
-1.282 |
0.000 |
| z |
0.000 |
0.000 |
-2.294 |
|
| Polar |
|---|
| 3z2-r2 | -4.589 |
|---|
| x2-y2 | 3.239 |
|---|
| xy | -0.227 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.229 |
0.877 |
0.000 |
| y |
0.877 |
6.828 |
0.000 |
| z |
0.000 |
0.000 |
5.262 |
<r2> (average value of r
2) Å
2
| <r2> |
82.922 |
| (<r2>)1/2 |
9.106 |
Jump to
S1C1
Energy calculated at wB97X-D/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -478.042530 |
| Energy at 298.15K | |
| HF Energy | -478.042530 |
| Nuclear repulsion energy | 107.446120 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3142 |
3142 |
26.47 |
|
|
|
| 2 |
A |
3122 |
3122 |
11.67 |
|
|
|
| 3 |
A |
3114 |
3114 |
18.44 |
|
|
|
| 4 |
A |
3079 |
3079 |
13.18 |
|
|
|
| 5 |
A |
3045 |
3045 |
24.70 |
|
|
|
| 6 |
A |
2737 |
2737 |
4.04 |
|
|
|
| 7 |
A |
1505 |
1505 |
3.19 |
|
|
|
| 8 |
A |
1498 |
1498 |
10.16 |
|
|
|
| 9 |
A |
1487 |
1487 |
1.90 |
|
|
|
| 10 |
A |
1421 |
1421 |
4.58 |
|
|
|
| 11 |
A |
1318 |
1318 |
17.24 |
|
|
|
| 12 |
A |
1289 |
1289 |
2.27 |
|
|
|
| 13 |
A |
1130 |
1130 |
6.31 |
|
|
|
| 14 |
A |
1076 |
1076 |
0.20 |
|
|
|
| 15 |
A |
999 |
999 |
6.23 |
|
|
|
| 16 |
A |
886 |
886 |
6.25 |
|
|
|
| 17 |
A |
744 |
744 |
1.80 |
|
|
|
| 18 |
A |
674 |
674 |
2.80 |
|
|
|
| 19 |
A |
334 |
334 |
1.34 |
|
|
|
| 20 |
A |
257 |
257 |
1.92 |
|
|
|
| 21 |
A |
205 |
205 |
14.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16530.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16530.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/cc-pCVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.635 |
-0.349 |
-0.053 |
| C2 |
0.493 |
0.642 |
0.090 |
| S3 |
-1.160 |
-0.100 |
-0.080 |
| H4 |
2.595 |
0.165 |
0.022 |
| H5 |
1.597 |
-1.104 |
0.732 |
| H6 |
1.590 |
-0.860 |
-1.013 |
| H7 |
0.545 |
1.172 |
1.040 |
| H8 |
0.531 |
1.392 |
-0.698 |
| H9 |
-1.073 |
-0.925 |
0.972 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5190 | 2.8062 | 1.0913 | 1.0903 | 1.0890 | 2.1666 | 2.1601 | 2.9525 |
C2 | 1.5190 | | 1.8190 | 2.1574 | 2.1635 | 2.1625 | 1.0884 | 1.0896 | 2.3837 | S3 | 2.8062 | 1.8190 | | 3.7658 | 3.0444 | 3.0014 | 2.4034 | 2.3386 | 1.3396 | H4 | 1.0913 | 2.1574 | 3.7658 | | 1.7640 | 1.7701 | 2.5007 | 2.5072 | 3.9430 | H5 | 1.0903 | 2.1635 | 3.0444 | 1.7640 | | 1.7625 | 2.5264 | 3.0684 | 2.6868 | H6 | 1.0890 | 2.1625 | 3.0014 | 1.7701 | 1.7625 | | 3.0714 | 2.5082 | 3.3220 | H7 | 2.1666 | 1.0884 | 2.4034 | 2.5007 | 2.5264 | 3.0714 | | 1.7519 | 2.6493 | H8 | 2.1601 | 1.0896 | 2.3386 | 2.5072 | 3.0684 | 2.5082 | 1.7519 | | 3.2762 | H9 | 2.9525 | 2.3837 | 1.3396 | 3.9430 | 2.6868 | 3.3220 | 2.6493 | 3.2762 | |
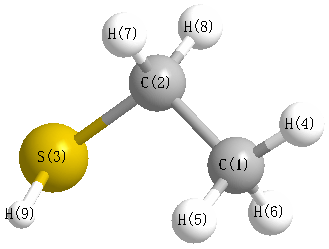 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.129 |
|
C1 |
C2 |
H7 |
111.318 |
| C1 |
C2 |
H8 |
110.732 |
|
C2 |
C1 |
H4 |
110.408 |
| C2 |
C1 |
H5 |
110.955 |
|
C2 |
C1 |
H6 |
110.956 |
| C2 |
S3 |
H9 |
96.823 |
|
S3 |
C2 |
H7 |
108.903 |
| S3 |
C2 |
H8 |
104.212 |
|
H4 |
C1 |
H5 |
107.913 |
| H4 |
C1 |
H6 |
108.554 |
|
H5 |
C1 |
H6 |
107.948 |
| H7 |
C2 |
H8 |
107.100 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.277 |
|
|
|
| 2 |
C |
-0.184 |
|
|
|
| 3 |
S |
-0.182 |
|
|
|
| 4 |
H |
0.097 |
|
|
|
| 5 |
H |
0.097 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
| 9 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.535 |
0.138 |
0.691 |
1.689 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.926 |
1.304 |
-0.685 |
| y |
1.304 |
-26.952 |
-1.645 |
| z |
-0.685 |
-1.645 |
-26.776 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.062 |
1.304 |
-0.685 |
| y |
1.304 |
0.899 |
-1.645 |
| z |
-0.685 |
-1.645 |
1.163 |
|
| Polar |
|---|
| 3z2-r2 | 2.327 |
|---|
| x2-y2 | -1.974 |
|---|
| xy | 1.304 |
|---|
| xz | -0.685 |
|---|
| yz | -1.645 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.778 |
0.377 |
0.028 |
| y |
0.377 |
5.990 |
-0.250 |
| z |
0.028 |
-0.250 |
5.682 |
<r2> (average value of r
2) Å
2
| <r2> |
83.530 |
| (<r2>)1/2 |
9.139 |
