Jump to
S1C2
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -190.470080 |
| Energy at 298.15K | |
| HF Energy | -189.620184 |
| Nuclear repulsion energy | 88.901379 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.865 |
| C2 |
0.000 |
0.000 |
-0.554 |
| C3 |
0.000 |
0.000 |
0.719 |
| O4 |
0.000 |
0.000 |
1.884 |
| H5 |
0.000 |
0.920 |
-2.434 |
| H6 |
0.000 |
-0.920 |
-2.434 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3116 | 2.5839 | 3.7494 | 1.0815 | 1.0815 |
C2 | 1.3116 | | 1.2722 | 2.4378 | 2.0936 | 2.0936 | C3 | 2.5839 | 1.2722 | | 1.1655 | 3.2844 | 3.2844 | O4 | 3.7494 | 2.4378 | 1.1655 | | 4.4153 | 4.4153 | H5 | 1.0815 | 2.0936 | 3.2844 | 4.4153 | | 1.8394 | H6 | 1.0815 | 2.0936 | 3.2844 | 4.4153 | 1.8394 | |
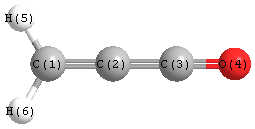 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.749 |
| C2 |
C1 |
H6 |
121.749 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.502 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -190.470613 |
| Energy at 298.15K | |
| HF Energy | -189.620123 |
| Nuclear repulsion energy | 89.109510 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.898 |
-1.590 |
0.000 |
| C2 |
0.000 |
-0.626 |
0.000 |
| C3 |
-0.307 |
0.627 |
0.000 |
| O4 |
-0.764 |
1.694 |
0.000 |
| H5 |
1.961 |
-1.389 |
0.000 |
| H6 |
0.604 |
-2.629 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3173 | 2.5238 | 3.6801 | 1.0816 | 1.0802 |
C2 | 1.3173 | | 1.2907 | 2.4423 | 2.1038 | 2.0924 | C3 | 2.5238 | 1.2907 | | 1.1598 | 3.0346 | 3.3820 | O4 | 3.6801 | 2.4423 | 1.1598 | | 4.1138 | 4.5344 | H5 | 1.0816 | 2.1038 | 3.0346 | 4.1138 | | 1.8382 | H6 | 1.0802 | 2.0924 | 3.3820 | 4.5344 | 1.8382 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
150.792 |
|
C2 |
C1 |
H5 |
122.259 |
| C2 |
C1 |
H6 |
121.244 |
|
C2 |
C3 |
O4 |
170.599 |
| H5 |
C1 |
H6 |
116.497 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
