Jump to
S1C2
Energy calculated at MP2/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -277.969512 |
| Energy at 298.15K | |
| HF Energy | -277.056701 |
| Nuclear repulsion energy | 126.116415 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3112 |
2952 |
0.00 |
224.82 |
0.06 |
0.11 |
| 2 |
Ag |
1538 |
1459 |
0.00 |
8.21 |
0.74 |
0.85 |
| 3 |
Ag |
1466 |
1390 |
0.00 |
2.40 |
0.33 |
0.49 |
| 4 |
Ag |
1121 |
1063 |
0.00 |
6.93 |
0.47 |
0.64 |
| 5 |
Ag |
1087 |
1031 |
0.00 |
8.40 |
0.37 |
0.54 |
| 6 |
Ag |
464 |
440 |
0.00 |
2.46 |
0.39 |
0.56 |
| 7 |
Au |
3196 |
3031 |
32.85 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1252 |
1188 |
4.49 |
0.00 |
0.75 |
0.86 |
| 9 |
Au |
823 |
780 |
0.01 |
0.00 |
0.75 |
0.86 |
| 10 |
Au |
121 |
115 |
13.13 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3172 |
3008 |
0.00 |
99.39 |
0.75 |
0.86 |
| 12 |
Bg |
1319 |
1251 |
0.00 |
7.42 |
0.75 |
0.86 |
| 13 |
Bg |
1193 |
1131 |
0.00 |
1.75 |
0.75 |
0.86 |
| 14 |
Bu |
3119 |
2958 |
47.44 |
0.00 |
0.05 |
0.10 |
| 15 |
Bu |
1548 |
1468 |
3.76 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1376 |
1305 |
7.14 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1098 |
1041 |
198.46 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
279 |
265 |
19.49 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 13641.6 cm
-1
Scaled (by 0.9484) Zero Point Vibrational Energy (zpe) 12937.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/cc-pVQZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.424 |
0.624 |
0.000 |
| C2 |
-0.424 |
-0.624 |
0.000 |
| F3 |
-0.424 |
1.718 |
0.000 |
| F4 |
0.424 |
-1.718 |
0.000 |
| H5 |
1.049 |
0.666 |
0.888 |
| H6 |
1.049 |
0.666 |
-0.888 |
| H7 |
-1.049 |
-0.666 |
0.888 |
| H8 |
-1.049 |
-0.666 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5086 | 1.3842 | 2.3419 | 1.0866 | 1.0866 | 2.1499 | 2.1499 |
C2 | 1.5086 | | 2.3419 | 1.3842 | 2.1499 | 2.1499 | 1.0866 | 1.0866 | F3 | 1.3842 | 2.3419 | | 3.5391 | 2.0157 | 2.0157 | 2.6200 | 2.6200 | F4 | 2.3419 | 1.3842 | 3.5391 | | 2.6200 | 2.6200 | 2.0157 | 2.0157 | H5 | 1.0866 | 2.1499 | 2.0157 | 2.6200 | | 1.7763 | 2.4848 | 3.0545 | H6 | 1.0866 | 2.1499 | 2.0157 | 2.6200 | 1.7763 | | 3.0545 | 2.4848 | H7 | 2.1499 | 1.0866 | 2.6200 | 2.0157 | 2.4848 | 3.0545 | | 1.7763 | H8 | 2.1499 | 1.0866 | 2.6200 | 2.0157 | 3.0545 | 2.4848 | 1.7763 | |
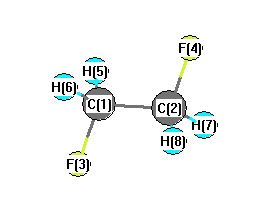 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.027 |
|
C1 |
C2 |
H7 |
110.821 |
| C1 |
C2 |
H8 |
110.821 |
|
C2 |
C1 |
F3 |
108.027 |
| C2 |
C1 |
H5 |
110.821 |
|
C2 |
C1 |
H6 |
110.821 |
| F3 |
C1 |
H5 |
108.733 |
|
F3 |
C1 |
H6 |
108.733 |
| F4 |
C2 |
H7 |
108.733 |
|
F4 |
C2 |
H8 |
108.733 |
| H5 |
C1 |
H6 |
109.642 |
|
H7 |
C2 |
H8 |
109.642 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -277.970911 |
| Energy at 298.15K | |
| HF Energy | -277.057222 |
| Nuclear repulsion energy | 128.235778 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3000 |
10.38 |
79.09 |
0.74 |
0.85 |
| 2 |
A |
3103 |
2943 |
34.48 |
208.39 |
0.02 |
0.03 |
| 3 |
A |
1512 |
1434 |
0.66 |
1.33 |
0.71 |
0.83 |
| 4 |
A |
1458 |
1383 |
11.88 |
1.90 |
0.17 |
0.29 |
| 5 |
A |
1324 |
1255 |
2.24 |
9.13 |
0.67 |
0.80 |
| 6 |
A |
1150 |
1091 |
15.60 |
1.68 |
0.48 |
0.65 |
| 7 |
A |
1144 |
1085 |
70.01 |
3.06 |
0.41 |
0.58 |
| 8 |
A |
890 |
844 |
23.38 |
5.45 |
0.16 |
0.27 |
| 9 |
A |
327 |
310 |
0.41 |
0.35 |
0.36 |
0.53 |
| 10 |
A |
155 |
147 |
3.64 |
0.13 |
0.75 |
0.86 |
| 11 |
B |
3175 |
3011 |
24.64 |
12.21 |
0.75 |
0.86 |
| 12 |
B |
3096 |
2937 |
8.01 |
29.95 |
0.75 |
0.86 |
| 13 |
B |
1510 |
1432 |
8.19 |
6.97 |
0.75 |
0.86 |
| 14 |
B |
1419 |
1346 |
6.86 |
0.04 |
0.75 |
0.86 |
| 15 |
B |
1281 |
1215 |
6.42 |
2.45 |
0.75 |
0.86 |
| 16 |
B |
1111 |
1054 |
50.09 |
3.38 |
0.75 |
0.86 |
| 17 |
B |
920 |
872 |
39.18 |
2.44 |
0.75 |
0.86 |
| 18 |
B |
503 |
477 |
16.29 |
0.51 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13619.9 cm
-1
Scaled (by 0.9484) Zero Point Vibrational Energy (zpe) 12917.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/cc-pVQZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.265 |
0.700 |
0.511 |
| C2 |
-0.265 |
-0.700 |
0.511 |
| F3 |
-0.265 |
1.417 |
-0.546 |
| F4 |
0.265 |
-1.417 |
-0.546 |
| H5 |
-0.017 |
1.197 |
1.438 |
| H6 |
1.348 |
0.697 |
0.410 |
| H7 |
0.017 |
-1.197 |
1.438 |
| H8 |
-1.348 |
-0.697 |
0.410 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4967 | 1.3822 | 2.3656 | 1.0891 | 1.0874 | 2.1255 | 2.1362 |
C2 | 1.4967 | | 2.3656 | 1.3822 | 2.1255 | 2.1362 | 1.0891 | 1.0874 | F3 | 1.3822 | 2.3656 | | 2.8830 | 2.0107 | 2.0079 | 3.2931 | 2.5604 | F4 | 2.3656 | 1.3822 | 2.8830 | | 3.2931 | 2.5604 | 2.0107 | 2.0079 | H5 | 1.0891 | 2.1255 | 2.0107 | 3.2931 | | 1.7802 | 2.3938 | 2.5325 | H6 | 1.0874 | 2.1362 | 2.0079 | 2.5604 | 1.7802 | | 2.5325 | 3.0350 | H7 | 2.1255 | 1.0891 | 3.2931 | 2.0107 | 2.3938 | 2.5325 | | 1.7802 | H8 | 2.1362 | 1.0874 | 2.5604 | 2.0079 | 2.5325 | 3.0350 | 1.7802 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.452 |
|
C1 |
C2 |
H7 |
109.565 |
| C1 |
C2 |
H8 |
110.518 |
|
C2 |
C1 |
F3 |
110.452 |
| C2 |
C1 |
H5 |
109.565 |
|
C2 |
C1 |
H6 |
110.518 |
| F3 |
C1 |
H5 |
108.316 |
|
F3 |
C1 |
H6 |
108.195 |
| F4 |
C2 |
H7 |
108.316 |
|
F4 |
C2 |
H8 |
108.195 |
| H5 |
C1 |
H6 |
109.754 |
|
H7 |
C2 |
H8 |
109.754 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
