Jump to
S1C2
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -189.622863 |
| Energy at 298.15K | |
| HF Energy | -188.857408 |
| Nuclear repulsion energy | 70.759832 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
-1.023 |
-0.438 |
0.000 |
| O3 |
1.150 |
0.109 |
0.000 |
| H4 |
-0.369 |
1.443 |
0.000 |
| H5 |
-0.650 |
-1.324 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3338 | 1.1911 | 1.0892 | 1.8600 |
O2 | 1.3338 | | 2.2403 | 1.9913 | 0.9615 | O3 | 1.1911 | 2.2403 | | 2.0223 | 2.3006 | H4 | 1.0892 | 1.9913 | 2.0223 | | 2.7817 | H5 | 1.8600 | 0.9615 | 2.3006 | 2.7817 | |
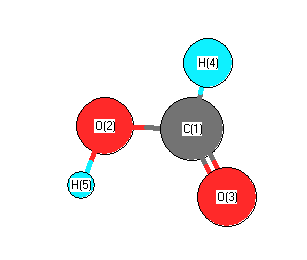 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
107.141 |
|
O2 |
C1 |
O3 |
124.972 |
| O2 |
C1 |
H4 |
110.127 |
|
O3 |
C1 |
H4 |
124.901 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -189.615930 |
| Energy at 298.15K | |
| HF Energy | -188.849790 |
| Nuclear repulsion energy | 70.522335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.382 |
0.000 |
| O2 |
-0.891 |
-0.621 |
0.000 |
| O3 |
1.170 |
0.195 |
0.000 |
| H4 |
-0.457 |
1.377 |
0.000 |
| H5 |
-1.777 |
-0.262 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3407 | 1.1846 | 1.0954 | 1.8901 |
O2 | 1.3407 | | 2.2160 | 2.0439 | 0.9564 | O3 | 1.1846 | 2.2160 | | 2.0112 | 2.9822 | H4 | 1.0954 | 2.0439 | 2.0112 | | 2.1041 | H5 | 1.8901 | 0.9564 | 2.9822 | 2.1041 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.598 |
|
O2 |
C1 |
O3 |
122.560 |
| O2 |
C1 |
H4 |
113.690 |
|
O3 |
C1 |
H4 |
123.750 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
