Jump to
S1C2
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -79.743484 |
| Energy at 298.15K | -79.749416 |
| HF Energy | -79.265266 |
| Nuclear repulsion energy | 42.455480 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3072 |
3072 |
0.00 |
|
|
|
| 2 |
A1g |
1452 |
1452 |
0.00 |
|
|
|
| 3 |
A1g |
1031 |
1031 |
0.00 |
|
|
|
| 4 |
A1u |
313 |
313 |
0.00 |
|
|
|
| 5 |
A2u |
3071 |
3071 |
50.73 |
|
|
|
| 6 |
A2u |
1429 |
1429 |
1.38 |
|
|
|
| 7 |
Eg |
3130 |
3130 |
0.00 |
|
|
|
| 7 |
Eg |
3130 |
3130 |
0.00 |
|
|
|
| 8 |
Eg |
1530 |
1530 |
0.00 |
|
|
|
| 8 |
Eg |
1530 |
1530 |
0.00 |
|
|
|
| 9 |
Eg |
1244 |
1244 |
0.00 |
|
|
|
| 9 |
Eg |
1244 |
1244 |
0.00 |
|
|
|
| 10 |
Eu |
3152 |
3152 |
57.17 |
|
|
|
| 10 |
Eu |
3152 |
3152 |
57.17 |
|
|
|
| 11 |
Eu |
1531 |
1531 |
8.47 |
|
|
|
| 11 |
Eu |
1531 |
1531 |
8.47 |
|
|
|
| 12 |
Eu |
835 |
835 |
3.03 |
|
|
|
| 12 |
Eu |
835 |
835 |
3.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16605.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16605.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.760 |
| C2 |
0.000 |
0.000 |
-0.760 |
| H3 |
0.000 |
1.013 |
1.153 |
| H4 |
-0.877 |
-0.506 |
1.153 |
| H5 |
0.877 |
-0.506 |
1.153 |
| H6 |
0.000 |
-1.013 |
-1.153 |
| H7 |
-0.877 |
0.506 |
-1.153 |
| H8 |
0.877 |
0.506 |
-1.153 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5197 | 1.0866 | 1.0866 | 1.0866 | 2.1648 | 2.1648 | 2.1648 |
C2 | 1.5197 | | 2.1648 | 2.1648 | 2.1648 | 1.0866 | 1.0866 | 1.0866 | H3 | 1.0866 | 2.1648 | | 1.7543 | 1.7543 | 3.0699 | 2.5193 | 2.5193 | H4 | 1.0866 | 2.1648 | 1.7543 | | 1.7543 | 2.5193 | 2.5193 | 3.0699 | H5 | 1.0866 | 2.1648 | 1.7543 | 1.7543 | | 2.5193 | 3.0699 | 2.5193 | H6 | 2.1648 | 1.0866 | 3.0699 | 2.5193 | 2.5193 | | 1.7543 | 1.7543 | H7 | 2.1648 | 1.0866 | 2.5193 | 2.5193 | 3.0699 | 1.7543 | | 1.7543 | H8 | 2.1648 | 1.0866 | 2.5193 | 3.0699 | 2.5193 | 1.7543 | 1.7543 | |
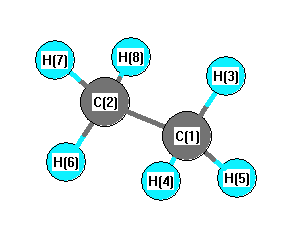 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.232 |
|
C1 |
C2 |
H7 |
111.232 |
| C1 |
C2 |
H8 |
111.232 |
|
C2 |
C1 |
H3 |
111.232 |
| C2 |
C1 |
H4 |
111.232 |
|
C2 |
C1 |
H5 |
111.232 |
| H3 |
C1 |
H4 |
107.655 |
|
H3 |
C1 |
H5 |
107.655 |
| H4 |
C1 |
H5 |
107.655 |
|
H6 |
C2 |
H7 |
107.655 |
| H6 |
C2 |
H8 |
107.655 |
|
H7 |
C2 |
H8 |
107.655 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -79.739063 |
| Energy at 298.15K | |
| HF Energy | -79.260424 |
| Nuclear repulsion energy | 42.290365 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3086 |
3086 |
0.00 |
|
|
|
| 2 |
A1' |
1465 |
1465 |
0.00 |
|
|
|
| 3 |
A1' |
1028 |
1028 |
0.00 |
|
|
|
| 4 |
A1" |
305i |
305i |
0.00 |
|
|
|
| 5 |
A2" |
3078 |
3078 |
52.77 |
|
|
|
| 6 |
A2" |
1426 |
1426 |
0.94 |
|
|
|
| 7 |
E' |
3160 |
3160 |
49.01 |
|
|
|
| 7 |
E' |
3160 |
3160 |
49.01 |
|
|
|
| 8 |
E' |
1538 |
1538 |
9.92 |
|
|
|
| 8 |
E' |
1538 |
1538 |
9.92 |
|
|
|
| 9 |
E' |
910 |
910 |
2.30 |
|
|
|
| 9 |
E' |
910 |
910 |
2.30 |
|
|
|
| 10 |
E" |
3138 |
3138 |
0.00 |
|
|
|
| 10 |
E" |
3138 |
3138 |
0.00 |
|
|
|
| 11 |
E" |
1527 |
1527 |
0.00 |
|
|
|
| 11 |
E" |
1527 |
1527 |
0.00 |
|
|
|
| 12 |
E" |
1196 |
1196 |
0.00 |
|
|
|
| 12 |
E" |
1196 |
1196 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16357.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16357.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.767 |
| C2 |
0.000 |
0.000 |
-0.767 |
| H3 |
0.000 |
1.009 |
1.167 |
| H4 |
-0.874 |
-0.504 |
1.167 |
| H5 |
0.874 |
-0.504 |
1.167 |
| H6 |
0.000 |
1.009 |
-1.167 |
| H7 |
0.874 |
-0.504 |
-1.167 |
| H8 |
-0.874 |
-0.504 |
-1.167 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5335 | 1.0855 | 1.0855 | 1.0855 | 2.1815 | 2.1815 | 2.1815 |
C2 | 1.5335 | | 2.1815 | 2.1815 | 2.1815 | 1.0855 | 1.0855 | 1.0855 | H3 | 1.0855 | 2.1815 | | 1.7474 | 1.7474 | 2.3349 | 2.9164 | 2.9164 | H4 | 1.0855 | 2.1815 | 1.7474 | | 1.7474 | 2.9164 | 2.9164 | 2.3349 | H5 | 1.0855 | 2.1815 | 1.7474 | 1.7474 | | 2.9164 | 2.3349 | 2.9164 | H6 | 2.1815 | 1.0855 | 2.3349 | 2.9164 | 2.9164 | | 1.7474 | 1.7474 | H7 | 2.1815 | 1.0855 | 2.9164 | 2.9164 | 2.3349 | 1.7474 | | 1.7474 | H8 | 2.1815 | 1.0855 | 2.9164 | 2.3349 | 2.9164 | 1.7474 | 1.7474 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.662 |
|
C1 |
C2 |
H7 |
111.662 |
| C1 |
C2 |
H8 |
111.662 |
|
C2 |
C1 |
H3 |
111.662 |
| C2 |
C1 |
H4 |
111.662 |
|
C2 |
C1 |
H5 |
111.662 |
| H3 |
C1 |
H4 |
107.194 |
|
H3 |
C1 |
H5 |
107.194 |
| H4 |
C1 |
H5 |
107.194 |
|
H6 |
C2 |
H7 |
107.194 |
| H6 |
C2 |
H8 |
107.194 |
|
H7 |
C2 |
H8 |
107.194 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
