All results from a given calculation for CH3NH2 (methyl amine)
using model chemistry: CCSD=FULL/cc-pVQZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -95.771731 |
| Energy at 298.15K | |
| HF Energy | -95.261107 |
| Nuclear repulsion energy | 42.243182 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.050 |
0.701 |
0.000 |
| N2 |
0.050 |
-0.754 |
0.000 |
| H3 |
-0.940 |
1.162 |
0.000 |
| H4 |
0.586 |
1.059 |
0.874 |
| H5 |
0.586 |
1.059 |
-0.874 |
| H6 |
-0.442 |
-1.103 |
-0.806 |
| H7 |
-0.442 |
-1.103 |
0.806 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4557 | 1.0916 | 1.0857 | 1.0857 | 2.0368 | 2.0368 |
N2 | 1.4557 | | 2.1565 | 2.0829 | 2.0829 | 1.0067 | 1.0067 | H3 | 1.0916 | 2.1565 | | 1.7616 | 1.7616 | 2.4550 | 2.4550 | H4 | 1.0857 | 2.0829 | 1.7616 | | 1.7473 | 2.9247 | 2.3952 | H5 | 1.0857 | 2.0829 | 1.7616 | 1.7473 | | 2.3952 | 2.9247 | H6 | 2.0368 | 1.0067 | 2.4550 | 2.9247 | 2.3952 | | 1.6121 | H7 | 2.0368 | 1.0067 | 2.4550 | 2.3952 | 2.9247 | 1.6121 | |
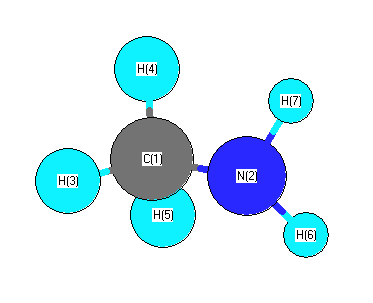 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H6 |
110.281 |
|
C1 |
N2 |
H7 |
110.281 |
| N2 |
C1 |
H3 |
114.935 |
|
N2 |
C1 |
H4 |
109.222 |
| N2 |
C1 |
H5 |
109.222 |
|
H3 |
C1 |
H4 |
108.011 |
| H3 |
C1 |
H5 |
108.011 |
|
H4 |
C1 |
H5 |
107.159 |
| H6 |
N2 |
H7 |
106.384 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
