All results from a given calculation for CH3SH (Methanethiol)
using model chemistry: CCSD=FULL/cc-pVQZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -438.271125 |
| Energy at 298.15K | |
| HF Energy | -437.764118 |
| Nuclear repulsion energy | 56.589066 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.048 |
1.148 |
0.000 |
| S2 |
-0.048 |
-0.662 |
0.000 |
| H3 |
1.277 |
-0.825 |
0.000 |
| H4 |
-1.088 |
1.452 |
0.000 |
| H5 |
0.430 |
1.540 |
0.889 |
| H6 |
0.430 |
1.540 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
S2 |
H3 |
H4 |
H5 |
H6 |
| C1 | | 1.8096 | 2.3763 | 1.0839 | 1.0826 | 1.0826 |
S2 | 1.8096 | | 1.3348 | 2.3563 | 2.4222 | 2.4222 | H3 | 2.3763 | 1.3348 | | 3.2833 | 2.6649 | 2.6649 | H4 | 1.0839 | 2.3563 | 3.2833 | | 1.7611 | 1.7611 | H5 | 1.0826 | 2.4222 | 2.6649 | 1.7611 | | 1.7774 | H6 | 1.0826 | 2.4222 | 2.6649 | 1.7611 | 1.7774 | |
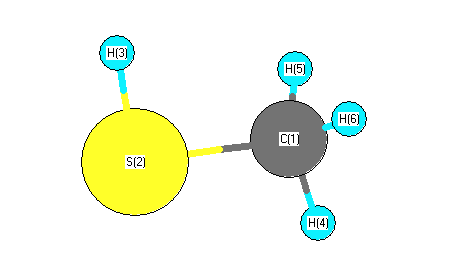 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
S2 |
H3 |
97.023 |
|
S2 |
C1 |
H4 |
106.324 |
| S2 |
C1 |
H5 |
111.255 |
|
S2 |
C1 |
H6 |
111.255 |
| H4 |
C1 |
H5 |
108.758 |
|
H4 |
C1 |
H6 |
108.758 |
| H5 |
C1 |
H6 |
110.351 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
