All results from a given calculation for CH2CHF (Ethene, fluoro-)
using model chemistry: CCSD=FULL/cc-pVQZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -177.690379 |
| Energy at 298.15K | |
| HF Energy | -176.967918 |
| Nuclear repulsion energy | 67.934910 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.432 |
0.000 |
| C2 |
1.179 |
-0.152 |
0.000 |
| F3 |
-1.137 |
-0.267 |
0.000 |
| H4 |
-0.175 |
1.494 |
0.000 |
| H5 |
1.274 |
-1.224 |
0.000 |
| H6 |
2.063 |
0.458 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
H4 |
H5 |
H6 |
| C1 | | 1.3160 | 1.3352 | 1.0762 | 2.0890 | 2.0633 |
C2 | 1.3160 | | 2.3194 | 2.1314 | 1.0756 | 1.0745 | F3 | 1.3352 | 2.3194 | | 2.0072 | 2.5940 | 3.2818 | H4 | 1.0762 | 2.1314 | 2.0072 | | 3.0796 | 2.4656 | H5 | 2.0890 | 1.0756 | 2.5940 | 3.0796 | | 1.8584 | H6 | 2.0633 | 1.0745 | 3.2818 | 2.4656 | 1.8584 | |
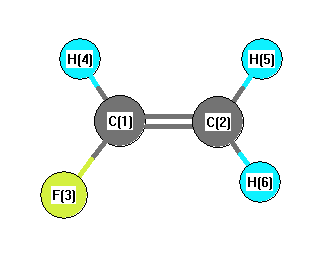 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.407 |
|
C1 |
C2 |
H6 |
118.989 |
| C2 |
C1 |
F3 |
122.058 |
|
C2 |
C1 |
H4 |
125.698 |
| F3 |
C1 |
H4 |
112.244 |
|
H5 |
C2 |
H6 |
119.604 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
