Jump to
S1C2
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -169.757400 |
| Energy at 298.15K | |
| HF Energy | -169.015991 |
| Nuclear repulsion energy | 71.728913 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.417 |
0.000 |
| O2 |
1.189 |
0.233 |
0.000 |
| N3 |
-0.932 |
-0.560 |
0.000 |
| H4 |
-0.442 |
1.421 |
0.000 |
| H5 |
-0.639 |
-1.516 |
0.000 |
| H6 |
-1.905 |
-0.344 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2030 | 1.3506 | 1.0966 | 2.0360 | 2.0520 |
O2 | 1.2030 | | 2.2642 | 2.0177 | 2.5294 | 3.1476 | N3 | 1.3506 | 2.2642 | | 2.0408 | 0.9999 | 0.9971 | H4 | 1.0966 | 2.0177 | 2.0408 | | 2.9435 | 2.2928 | H5 | 2.0360 | 2.5294 | 0.9999 | 2.9435 | | 1.7258 | H6 | 2.0520 | 3.1476 | 0.9971 | 2.2928 | 1.7258 | |
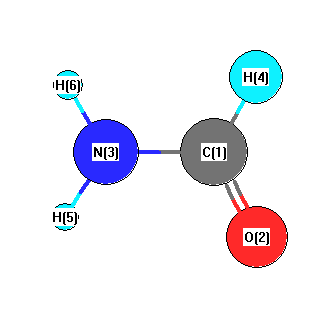 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.296 |
|
C1 |
N3 |
H6 |
121.126 |
| O2 |
C1 |
N3 |
124.819 |
|
O2 |
C1 |
H4 |
122.589 |
| N3 |
C1 |
H4 |
112.592 |
|
H5 |
N3 |
H6 |
119.577 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -169.757400 |
| Energy at 298.15K | -169.761001 |
| HF Energy | -169.015990 |
| Nuclear repulsion energy | 71.730060 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3802 |
3802 |
53.42 |
|
|
|
| 2 |
A |
3661 |
3661 |
51.63 |
|
|
|
| 3 |
A |
3032 |
3032 |
75.22 |
|
|
|
| 4 |
A |
1849 |
1849 |
423.14 |
|
|
|
| 5 |
A |
1649 |
1649 |
64.51 |
|
|
|
| 6 |
A |
1453 |
1453 |
5.21 |
|
|
|
| 7 |
A |
1301 |
1301 |
120.72 |
|
|
|
| 8 |
A |
1079 |
1079 |
5.81 |
|
|
|
| 9 |
A |
1079 |
1079 |
1.68 |
|
|
|
| 10 |
A |
641 |
641 |
15.02 |
|
|
|
| 11 |
A |
580 |
580 |
12.12 |
|
|
|
| 12 |
A |
146 |
146 |
216.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10136.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10136.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.161 |
0.385 |
-0.000 |
| O2 |
-1.186 |
-0.245 |
0.000 |
| N3 |
1.076 |
-0.157 |
-0.000 |
| H4 |
-0.141 |
1.481 |
0.000 |
| H5 |
1.175 |
-1.152 |
0.000 |
| H6 |
1.891 |
0.418 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2030 | 1.3505 | 1.0966 | 2.0360 | 2.0521 |
O2 | 1.2030 | | 2.2640 | 2.0177 | 2.5291 | 3.1476 | N3 | 1.3505 | 2.2640 | | 2.0409 | 0.9999 | 0.9971 | H4 | 1.0966 | 2.0177 | 2.0409 | | 2.9436 | 2.2934 | H5 | 2.0360 | 2.5291 | 0.9999 | 2.9436 | | 1.7257 | H6 | 2.0521 | 3.1476 | 0.9971 | 2.2934 | 1.7257 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.295 |
|
C1 |
N3 |
H6 |
121.144 |
| O2 |
C1 |
N3 |
124.802 |
|
O2 |
C1 |
H4 |
122.587 |
| N3 |
C1 |
H4 |
112.610 |
|
H5 |
N3 |
H6 |
119.561 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
